• By Kevin E. Noonan –

    Supreme Court Building #1The Supreme Court's consideration of the standards for satisfying the enablement provisions of 35 U.S.C. § 112(a) has been occasioned for the first time in over a century by the Court's granting certiorari in Amgen v. Sanofi.  This has not surprisingly generated a great deal of interest and amicus briefing.  While some of these briefs, for both Petitioners (see "Patent Law Academics File Amicus Brief in Amgen v. Sanofi"; "AbbVie Files Amicus Brief in Amgen v. Sanofi"; "GlaxoSmithKline Files Amicus Brief in Amgen v. Sanofi") and Respondents (see "Esteemed Scientists File Amicus Brief in Amgen v. Sanofi on Respondents' Behalf"; "Another Group of Law Professors File Amicus Brief in Amgen v. Sanofi"; "U.S. Government Files Amicus Brief in Amgen v. Sanofi") have been the subject of earlier posts prior to oral argument last month, in the interest of completeness and because there were interesting positions taken and arguments raised in the remaining briefs, a more succinct but not cursory review of these briefs is warranted.  Amicus briefs in support of neither Party are the subject of this post.

    Amicus briefs in support of neither party

    The amicus brief filed by the Intellectual Property Owners (IPO) argued that the enablement requirement "strikes a balance" on claim breadth with regard to the quid pro quo, from the perspective of the person of ordinary skill in the relevant art, and that it is important that the inquiry be based on the language of the statute.  But even with these policy goals in mind, "an applicant is not required to describe in the specification every conceivable possible and future embodiment of his invention" they argue, citing Cordis Corp. v. Medtronic AVE, Inc., 339 F.3d 1352, 1365 (Fed. Cir. 2003), and In re Angstadt, 537 F.2d 498, 503 (C.C.P.A. 1976).

    Undue experimentation is the proper standard which is a fact-intensive inquiry that is helped but not limited to the Wands factors.  However the inquiry is made, it must be applied in a predictable and fair manner that is consistent with the statutory language and longstanding precedent and should not "disrupt the settled expectations of the inventing community."  See Festo Corp. v. Shoketsu Kinzoku Kogyo Kabushiki Co., 535 U.S. 722, 739 (2002), these amici argue.  Also emphasized in their brief is the statutory presumption of validity that can only be overcome by clear and convincing evidence.  When enablement does arise as an invalidity defense in federal district court, the burden always rests on the patent challenger to prove lack of enablement by clear and convincing evidence, under Microsoft Corp. v. i4i Limited P'ship, 564 U.S. 91, 100-03 (2011).  On appellate review, courts " give due deference to the fact-finder's credibility determinations in weighing competing expert evidence," citing Graver Tank & Mfg. Co. v. Linde Air Prods. Co., 339 U.S. 605, 611 (1950).  All this to say is that the IPO's brief hews the course of its neutrality regarding either party in the dispute by sticking to these eternal verities of enablement law.

    The Regenexbio, IGM Biosciences, and Adapative Phage Therapeutics brief was filed by a self-described group of "small to mid-sized biotech companies currently pursuing innovation across multiple platforms," wherein Regenexbio is developing gene therapy approaches for treating "retinal, metabolic, and neurodegenerative diseases"; IGM Biosciences is developing IgM-based immunotherapies against "cancer, infectious diseases, and autoimmune and inflammatory diseases"; and Adapative Phage Therapeutics is developing bacteriophage banks that "provide evergreen, broad spectrum, and polymicrobial coverage."

    Their expressly asserted reason for filing their amicus brief is a concern that the Federal Circuit's application of the enablement standard "threatens innovation in the life sciences" by "shift[ing] the burden from the challenger to the patentee to prove enablement of its issued patents" and "makes the burden of proving enablement nearly insurmountable for innovative start-ups, small-to-medium-sized companies, universities, and other innovators with limited resources."  This test will make innovators "waste precious resources by forcing them to reduce to practice not just instructive and illustrative examples but an undefined number of individual species that embody every 'corner' of the claims" they argue, which will be prejudicial to smaller companies.  Accordingly, amici address the Court seeking "reasonable disclosure standards" for enablement.

    Their arguments rely on the presumption of validity:

    Amici rely on the statutory presumption of validity that adheres to every patent claim after the Patent Office examines and issues the patent.  The presumption ensures that the innovative value captured by the claims will not be wiped away on a dubious or speculative case.  It does so by placing the burden of proof on the patent challenger.  And by requiring the challenger to meet that burden with clear-and-convincing evidence.

    The reach-the-full-scope standard undermines that presumption.  The Federal Circuit acknowledges this when it "raises the bar" or sets "high hurdles" for particular claims.  These phrases lay the burden of proof on the patentee as soon as the court reads the claims.  Further, they show that the court is adjusting the standard of proof to be higher for the patentee (or, more properly, lower for the patent challenger) than the standard codified in the statute.

    Amici also argue that the Federal Circuit's full scope test is inconsistent with constructive reduction to practice.

    The Intellectual Property Law Association of Chicago (IPLAC — amici asserting that they are "the country's oldest bar association devoted exclusively to intellectual property matters") in their brief ask the Court to vacate and remand because the case can be settled by the application of the statute and stare decisis.  "Enablement is not, because it never has been and never should be—but for decisions such as the reviewed one—a matter of whether the specification has catalogued all possible embodiments of invention and taught each of them; that is not the law" they assert.  The brief relies heavily on the Karshtedt et al. law review article, and one by Jason Rantanen, The Doctrinal Structure of Patent Law's Enablement Requirement, 69 Vanderbilt L. Rev. 1679, 1680 (2019).  Both stand for the proposition that the Federal Circuit has recently "varied greatly from deciding enablement in a proper manner and muddled their own jurisprudence on the issue."  Their brief also criticizes In re Wands for leaving the definition of undue experimentation "amorphous"

    The brief suggests that the Court's decision in Consol. Elec. Light Co. v. McKeesport Light Co. (Incandescent Lamp), 159 U.S. 465 (1895), sets the standard for undue experimentation (albeit it should be recognized that this takes the definition from the extreme, where >6,000 failed attempts constitute undue experimentation; while that is certainly the case it seems unlikely to be encountered routinely).  The brief distinguishes the Consol. Elec. standard with the facts below, where Respondent did not identify any antibody that would/could not be obtained using Amgen's disclosure (the "roadmap").

    And In a sad or amusing footnote (depending on your prejudices) the brief sets forth a "list of areas of patent law in which this Court has overturned the Court of Appeals [that] is long, and the frequency of the overrulings notorious"; with two exceptions these cases were all decided in this century and over 80% since 2010.

    Another local intellectual property law group, the New York Intellectual Property Law Association (NY IPLA) argued in its brief that the Federal Circuit has gone beyond the requirements of the plain language of the statute, which has been the standard applied by the court since In re Hyatt, 708 F.2d 712, 714 (Fed. Cir. 1983).  But as applied to biotechnology inventions amicus argues that genus claims have required "disclosure of every species in the genus," citing Wyeth v. Abbott Laboratories, 720 F.3d 1380, 1385 (2013), Enzo Life Sciences, Inc. v. Roche Molecular Systems, Inc., 928 F.3d 1340, 1345–48 (Fed. Cir. 2019), and Idenix Pharmaceuticals LLC v. Gilead Sciences Inc., 941 F.3d 1149, 1160–63, 1165 (Fed. Cir. 2019), based on the court's perceptions regarding "the uncertainty of the science and the required functionality."  The NYIPLA contend that "the requirement for enablement of the full scope of the claimed invention is not supported by the plain language of the statute and is contrary to th[e] Court's holding as Minerals Separation v. Hyde, 242 U.S. 261, 270-71 (1916)."

    Although the brief recognizes that there is a valid concern at issue, that an inventor not be "granted patent claim coverage that exceeds what the inventor has invented," amici predict a severe adverse impact that would arise should the Court affirm the Federal Circuit test, specifically that "could lead inventors to disclose less than what they discovered, such as a functional relationship at the heart of why the invention works.  Although the amicus appreciates the potential inequities that could be occasioned by claims like Amgen's, on balance the consequences of affirming the Federal Circuit's "full scope" test are worse due to the inherent unpredictability of the chemical and biotechnological arts, amicus argues, citing In re Fisher, 427 F.2d 833, 839 (CCPA 1970), for the principle that "the scope of enablement obviously varies inversely with the degree of unpredictability of the factors involved.")  Affirming the Federal Circuit would create a situation wherein "the inventor will have no incentive (or duty) to disclose the genus, which perhaps comprises the most important part of the discovery" and "[t]he rest will be kept from the public as a trade secret," until such time that the patentee develops another species that can be subject to its own 20-year term.

    While amicus argues that the Federal Circuit's decisions "goes beyond the literal requirements of the statute's plain language," the brief also gives little credence to Amgen's arguments that the Federal Circuit had only recently imposed an unnecessarily stringent enablement standard, citing earlier precedent to the same effect, including In re Fisher, 427 F.2d 833, 839 (CCPA 1970); In re Hyatt, 708 F.2d 712, 714 (Fed. Cir. 1983); Amgen, Inc. v. Chugai Pharm. Co., 927 F.2d 1200, 1212, (Fed. Cir. 1991); In re Wright, 999 F.2d 1557, 1561 (Fed. Cir. 1993); and Genentech Inc. v. Novo Nordisk A/S, 108 F.3d 1361, 1365 (Fed. Cir. 1997).

    Their solution:  requiring disclosure of a "reasonable number of species" test for prosecution before the USPTO that could be limited at trial by claim construction and having undue experimentation be a defense against infringement species by species, the patentee being protected by the clear and convincing evidence standard (to the extent it is respected by district courts and the Federal Circuit in deciding JMOL motions).

    As for disposition, the brief advises that "[i]n the present case, respectfully the Court should remand the case for a determination of whether the Sanofi antibody was obtainable from the disclosure in the Amgen patents with some — but not undue — experimentation at the time of the filing of the Amgen application."
  • By Kevin E. Noonan –

    Supreme Court Building #2The Supreme Court's consideration of the standards for satisfying the enablement provisions of 35 U.S.C. 112(a) has been occasioned for the first time in over a century by the Court's granting certiorari in Amgen v. Sanofi.  This has not surprisingly generated a great deal of interest and amicus briefing.  While some of these briefs, for both Petitioners (see "Patent Law Academics File Amicus Brief in Amgen v. Sanofi"; "AbbVie Files Amicus Brief in Amgen v. Sanofi"; "GlaxoSmithKline Files Amicus Brief in Amgen v. Sanofi") and Respondents (see "Esteemed Scientists File Amicus Brief in Amgen v. Sanofi on Respondents' Behalf"; "Another Group of Law Professors File Amicus Brief in Amgen v. Sanofi"; "U.S. Government Files Amicus Brief in Amgen v. Sanofi") have been the subject of earlier posts prior to oral argument last month, in the interest of completeness and because there were interesting positions taken and arguments raised in the remaining briefs a more succinct but not cursory review of these briefs is warranted.  The remaining amicus briefs in support of Respondents are the subject of this post.

    Amicus briefs in support of Respondents

    Like the amicus briefs in favor of the Petitioner, the briefs for Respondents recite some themes in common.  These include that permitting patenting of Amgen's claims (and those like them) would upset the "balance" between disclosure and claim scope to the public's detriment.  Many of the briefs discuss the history of the discovery of PCSK9, its biological activity and binding to the live LDL receptor, and how blocking this binding was effective in lowering serum cholesterol; Amgen did not invent or discover any of these relationships or uses for PCSK9 antibodies.  Several briefs mentioned 35 U.S.C. 112(f) as an alternative for functional claiming of PCSK9 antibodies, and Consolidated Electric Light Co v. McKeesport Light Co., 159 U.S. 465, 472-76 (1895), was recognized as a seminal case supporting Respondent's position.

    Fresenius Kabi ISA LLC's amicus brief argues that enablement provides balance to the public, and overbroad patents disrupt the balance to the public's detriment.  The amicus is concerned about gamesmanship by which patentees will withhold disclosure in favor of broad, functionally claimed patents with staggered specific disclosures.  Citing Consolidated Electric Light Co v. McKeesport Light Co., 159 U.S. 465, 472-76 (1895) (the lead case for respondent), amicus argues that there must be a robust enablement standard, and Petitioner's standard will "negatively affect competition" because patentees will use "broad, nonspecific patent disclosures to block entire fields of innovation."

    The brief recites "tensions" between innovators and the public, accusing biopharma companies of disclosing a "drug peptide sequence" and holding back "details that are required by competitors to expand on and further innovate—or even actually use the peptide—based on the work of the initial application invention" (e.g., "details on physicochemical or functional properties of the drug" such as "glycan profile, charge profile, variants profile, impurity profile, immunochemical properties, and functional activities"), which amicus calls "gam[ing] the system."  These allegations of claim shenanigans against biopharma companies extend to including claim language directed towards "therapeutically effective amount" and "excipients, adjuvants, or diluents" that may be capable of overextending patent protection inequitably (although such terminology certainly did not arise with biotechnology patents).  Functional claiming is available, if desired, under 35 U.S.C. 112(f), says this amicus, which does not have the aforementioned negative impacts on innovation because it has consistently been construed to "limit the scope of the claim to only those specific structures described in the specification," citing Traxcell Techs., LLC v. Spring Commc'ns Co., 15 F.4th 1121, 1134 (Fed. Cir. 2021).

    The brief contains a litany of opportunities for patentees to "control" patent prosecution and claim scope, including continuation applications (but warns of these being abused too, stating that "[w]ithout a robust enablement standard, continuation applications can be used to continually expand the scope of exclusionary patent rights unfairly by encompassing embodiments not appreciated or even discovered at the time of filing the original application") and suggests that continuation-in-part applications provide an avenue for any such desired (and deserved) protections (which might be at least a trifle unrealistic).

    Finally, the brief uniquely argues that the change to a "first inventor to file" regime under the AIA has increased the risk of loosened enablement standards, because under the "first to invent" provisions of earlier U.S. patent law broadened claims could be challenged by earlier inventors but no more.

    Eli Lilly & Co., Ipsen Bioscience, Inc., and Innovent Biologics, Inc. filed an amicus brief arguing that Amgen's claims violated the "letter, logic, and purpose" of the enablement requirement because they recited purely functional results.  The patent bargain was raised as a basis for such claims to be invalid and amici accused Amgen of attempting to "control all antibody therapeutics to a particular target" which would be "indisputably detrimental to the public."  The brief provides various calculations for the number of antibodies falling within the claim scope, noting the conventional "millions" in the argument but reciting in a footnote 2060 (1 x 1078) based on 6 CDRs and 10 amino acids/CDR and 20 amino acids.

    Amici asserted that claims are "nothing more than a hunting license" contrary to Brenner v. Manson, 383 U.S. 519, 536 (1966), and that they are also not properly considered to be genus claims because they do not disclose a structural similarity between them, just their function in blocking PCSK9 binding to LDL receptor.

    And amici countered Amgen's (actually GSK's) R&D efficiency argument with the hypothetical that a troll with Amgen's claims could "dramatically undermine resource allocation efficiency" and Amgen's airplane analogy to permit claiming all airplanes using the Bernoulli principle, saying that this illustrates the "perennial preemption problem" created by functional claims.

    These amici also argue that the Federal Circuit's enablement jurisprudence comports with the legal tenets under Consol. Elec. and Holland Furniture Co. v. Perkins Glue Co., 277 U.S. 245, 256-257 (1928), that the boundaries of what is claimed should be clear, and distinguish Minerals Separation v. Hyde, based on Amgen not providing anything to optimize the claimed antibodies but rather encompassing "a virtually limitless universe of other, non-conservative antibodies," stating that "[t]o hold such paltry disclosure sufficient to enable claims preempting an entire technological field would turn the patent quid pro quo on its head," citing Ex parte Sloane, 22 U.S.P.Q. 222 (P.O.B.A. Jan. 18, 1934); In re Angstadt, 537 F.2d 498, 500, 503 (C.C.P.A. 1976); and Atlas Powder Co. v. E.I. du Pont De Nemours & Co., 750 F.2d 1569, 1576 (Fed. Cir. 1984).

    This brief also argues regarding the use of 35 U.S.C. 112(f) for functional claims such as Amgen's, because that portion of the statute "creat[ed] a limited exception to this Court's prohibition against functional claiming," citing Warner-Jenkinson Co., Inc. v. Hilton Davis Chemical Co., 520 U.S. 17, 28 (1997) (but even that doesn't save Amgen's claims amici say because "none of Amgen's claims at issue are a combination of structural and functional elements" because reciting "monoclonal antibody" does not import sufficient structure).

    Turning to patent policy, amici challenge Amgen and amici supporting Amgen's position with regard to the Federal Circuit's test harming the U.S. biotechnology and pharma industries, asserting that U.S. Biotech and pharma will continue global leadership using current interpretations of enablement law, based on assertions that the sector is "booming" and accuses Amgen of trying to "upend the legal regime that has empowered that growth, innovation, and undeniable benefit to the public" by upending the full-scope standard (!) "under the guise of an 'innovation' imperative."  The brief sets forth as an example antibodies to cell surface marker CD20, wherein "four innovators developed six different antibody therapies for three different illnesses while targeting the same protein."  They warn "if Amgen's interpretation of the Patent Act were the law of the land, the first party to obtain a functional claim to all antibodies that inhibit binding to a target could prevent market entry and/or extract rents from all future innovators for twenty years."

    Pfizer filed a brief supporting Respondents, again reciting the quid pro quo patent bargain and that permitting claims such as Amgen's would preempt future research.  While acknowledging that genus claims can provide important patent protection Amgen's claims went too far into purely functional claiming without sufficient structure to support them.  "[I]t is the undue breadth of the claims and the exclusive rights [Amgen] seek['s] to encompass, rather than a heightened standard for enablement of genus claims, which led the district court and the Federal Circuit to conclude that the claims are invalid as a matter of law."  The brief also calls these claims "a naked attempt to preempt future innovation and an unwarranted extension of the patent monopoly."

    Another group of biotech and pharma companies, including Genentech, AstraZeneca Pharma, Bayer AG, Gilead Sciences, and Johnson & Johnson, filed an amicus brief where they argue that the Federal Circuit upholds genus claims that are supported by disclosure having appropriate scope using a flexible standard that is consistent with Supreme Court and its own precedent, including O'Reilly v. Morse, 56 U.S. 62, 120-21 (1853); Holland Furniture Co. v. Perkins Glue Co., 277 U.S. 245, 257-58 (1928); Consol. Elec. Light Co. v. McKeesport Light Co., 159 U.S. 465, 476 (1895).  But that is not the case here, they argue, and the Federal Circuit properly invalidated the claims on enablement grounds.  Their brief particularly decries arguments that merely showing how to make and use one embodiment could be enough, asserting that "[a] patentee who chooses to claim a broad functional genus, unlimited by claim language delineating the specific structures that achieve that function, will necessarily have to disclose more than a patentee claiming certain specified structures or species within that genus" (and that one of the Wands factors is claim breadth).

    The brief states that "Amgen and its amici desire a regime where patentees can jump the gun, rushing to the patent office with broad functional genus claims before they have discovered and can explain how to make and use, without excessive brute-force experimentation, vast swaths of the genus claimed, including structurally dissimilar species which may vary substantially in how they perform the claimed function.  But such unsupported broad claims "discourage rather than promote invention" by allowing a patentee to "foreclose efforts to discover other and better types" within the genus.  Holland Furniture, 277 U.S. at 257; accord Consolidated Elec., 159 U.S. at 476."

    Curiously, most of the scholarly works cited in support of the arguments made by amici are at least 10 years old and some are significantly older.

    Another collection of industries, the Small and Medium Biotechnology Companies (which includes ABL Bio (USP 11,261,259), Kiniksa, OPKO Health, and SK bioscience) say the Court should adopt an "if it ain't broke, don't fix it" principle and emphasized the reliance interest on the Federal Circuit's enablement jurisprudence, stating "[t]he longstanding enablement standard is consistent with text and precedent.  The balance it strikes promotes innovation and saves lives.  And departing from the status quo would unleash harmful consequences for industry participants like Amici, for patients, and for the public."  The Wands factors "are drawn from, and reflect" a century and a half of Supreme Court case law on enablement, and have been applied "evenhandedly" "for decades" they assert.  The brief cites a number of "incumbent and follow-on innovators," illustrated by Humira®, Remicade®, Cimzia®, and Simponi®, all of which are TNF-alpha inhibitors and Skyrizi®, Tremfya®, and Illumya®, which inhibit IL-23, for the purported benefits of permitting patent protection for both types of innovation.

    The same curious reliance on scholarly works cited are at least 10 years old and some significantly older is the case here as in Pfizer's brief.

    In addition to the legal academics' amicus briefs that were the subject of previous posts, a collection of IP Law Professors (including Sean Tu, Arti Rai, Oskar Litvak, Kevin Collins, and Bernard Chao) filed a brief and argued that claims should be limited to what is disclosed because that is what is invented.  In their view, "[t]rial-and-error inventing" (like Amgen's here, in their view) is inherently narrow, as evidenced by Amgen's "roadmap" disclosure, and the professors draw a distinction between trial-and-error to find the invention and trial-and-error in making and using what has been invented.  Countering some of the more heated rhetoric from Amgen and its amici (by asserting "[t]here is no paradox, no death of anything or otherwise a need for alarm"), the professors write that "[t]his case presents a classic example of a narrow invention that is coupled to overbroad claims," which are overbroad because a patent applicant "can claim broadly only when [they have] disclosed structural features that unify different species."  The brief cites several examples of competing antibodies that (putatively) would be precluded by grant of broad genus claims as Amgen advocates and the professors make the case that narrow antibody claims are more conducive to innovation in this area.

    Much of the argument is taken from S. Sean Tu and Christopher M. Holman, Antibody Patents: Use of the Written Description and Enablement Requirements at the Patent & Trademark Office, 38 Berkeley Tech. L. J. (Figure 8) (2023 forthcoming) (available at https://papers.ssrn.com/sol3/papers.cfm?abstract_id=4025167).

    In her brief, Professor Robin Feldman takes the extreme position that "[t]his Court's precedent holds that the correct standard here is not undue experimentation but rather any experimentation," based on Wood v. Underhill, 46 U.S. (5 How.) 1, 4 (1847), which can be read this way, and Consolidated Electric Light Co. v. McKeesport Light Co., 159 U.S. 465, 474-75 (1895).  She writes that overbroad claiming "could bar scientists and competitors from those building blocks of invention" (notice conditional phrase) and that the certiorari grant was improvident because the Supreme Court could validate the undue experimentation standard.

    The AIPLA (whose brief "suggest[s]" affirmance) argues that the Federal Circuit applied established law but that while enablement does not require disclosure of all embodiments Wands is the correct legsal and analytical framework.  The brief cites as examples the seemingly settled principle that one non-enabled embodiment is not enough to be undue experimentation and states that Amgen's argument that the Federal Circuit had developed a new "full scope" test is a "facile strawman."

    The Association for Accessible Medicines filed an amicus brief wherein they argued that the Federal Circuit's decision aligns with statutory text and with the structure and purpose of the Patent Act, and that Amgen's theory will inhibit competition in the pharma industry.  As in other briefs discussed above this brief emphasizes the patent bargain, the dangers of permitting overbroad claiming, and the availability of Section 112(f) for such claims (consistent with its enactment in response to the Supreme Court's Halliburton Oil Well Cementing Co. v. Walker decision, 329 U.S. 1 (1946).  The brief focuses on the interplay between Sections 112(a) and 112(b) (wherein the latter defines the invention) and that the existence of the "using" requirement in Section 112(a) mandates disclosing how to use the invention, i.e., which species are operable.

    The brief cites the usual litany of prior Supreme Court cases, such as O'Reilly v. Morse, Consolidated Electric, and Holland Furniture, and distinguishes cases Amgen cites by stating that "the issue in each case was that an operator seeking to use the invention would need to fill in some gaps in the patent's instructions to achieve the invention's desired outcome.  This Court held in each case that the specification contained sufficient information for a skilled artisan to fill in those gaps."  Which is not the case here, they argue.

    The brief concludes by exhorting the Court to restrict "therapeutically effective amount" language and excipient claims (also argued by Fresenius Kabi ISA LLC, see above), citing biologic drugs with "secondary" characteristics not disclosed or claimed in protein composition of matter-claiming patents based on failure to disclose how to use a claimed biologic drug.

    A sole generic drugmaker, Viatris Inc. filed an amicus brief making many of these same arguments, for example about the quid pro quo and the patent bargain, and asserts that deviating from the Wands precedent would destabilize U.S. patent law and innovation.  Uniquely this brief also argues that changes in the enablement standard would have a "major ripple effect" into other areas of patent law including obviousness.  This brief also ties enablement under 112(a) with the statutory requirements of defining an invention by claims in 112(b) as the basis for the scope of the claim be commensurate with the "full scope" of the invention as claimed.  Asserting the crux of their argument the brief recites:

    Accordingly, this Court has long held that the specification—"[a]ddressed as it is to those skilled in the art"—"may leave something to their skill in applying the invention" (Mowry v. Whitney, 81 U.S. 620, 644 (1872)), provided the requisite experimentation is routine rather than "painstaking" (Consol. Elec. Light Co. v. McKeesport Light Co., 159 U.S. 465, 475 (1895)) or "elaborate" (Holland Furniture Co. v. Perkins Glue Co., 277 U.S. 245, 256-257 (1928)).

    A group of medical doctors and affiliated groups, styled in the caption as Arnold Ventures, National Centers for Health Research, and Certain Medical Doctors, filed an amicus brief whose arguments are directed to perceived negative consequences on innovation and competition in pharma, citing reports about high drug costs, many by the "certain medical doctors" themselves.  They argue (as have others) that functional genus claims would permit pharma companies to use such claims to prevent other companies from developing different drugs to the same target.  The brief cites four ways that adopting Amgen's position would hurt patients:

    1) "multiple treatment options will be delayed in development and approval, or not available at all" (e.g., if the claimed drug fails in the clinic);
    2) "patients will be forced to take medicine that may be either more risky or less efficacious—or both—than an alternative." (when first-to-market drugs are not the most efficacious);
    3) "patients may not have the treatment option that works best for their specific circumstances" (when humans have differential responses to different drugs to the same target); and
    4) "competition between innovators, under the right circumstances, can lower spending on brand-name drugs," stating that "[h]igh drug prices are a major cause of lack of patient access, or of long-term non-adherence to treatments, which contributes to thousands of excess hospitalizations and deaths annually."

    The brief provides an informative schematic showing the timeline of PCSK9 research, which throws shade on Amgen's entitlement to a patent in view of what they contend Amgen did not invent:

    Figure 1
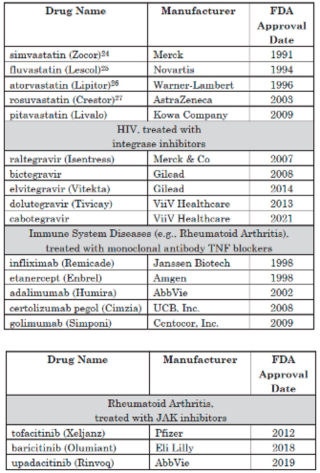
    and provides a series of tables showing "second to market" drugs that could have been precluded if Amgen's enablement position had been the standard:

    Table 1
    Table 2
    The amicus filed by Unified Patents illustrated the concerns the enablement issue raised for patentees outside the biotech and pharma industries.  This brief argues that functional claims impede innovation regardless of technology and that the "full scope" test is needed to prevent this negative outcome.  In addition, the brief argues that the "full scope" test is not new but instead is supported by established precedent which Amgen's proposed test would overturn, citing Morse, Béné v. Jeantet, The Incandescent Lamp Patent, Corona Cord Tire Co. v. Dovan Chem. Corp., Holland Furniture and CCPA/Federal Circuit precedent and stating that:

    The Court has identified three relevant factors: (1) whether the patent involves an area of invention where results are unpredictable; (2) whether the claim breadth vastly outstrips the disclosures in the specification; and (3) whether the patentee has demonstrated that the proven function of disclosed embodiments can be reliably extrapolated to non-disclosed embodiments within the claim scope.

    For this amicus the greatest threat raised by Amgen's argument is the issuance of overbroad "high-tech" patents, devoting an entire section of the brief to this "problem" and stating that  such broad functional claims would have an in terrorem effect on the public.  The brief also provides a footnote asserting that a study of patent litigations estimated that 100% of NPE-asserted software patents and 50% of non-NPE asserted software patents utilized functional patent claiming.  Colleen V. Chien and Aashish R. Karkhanis, Functional Claiming and Software Patents (Santa Clara Univ., Working Paper No. 06-13, 2013), https://ssrn.com/abstract=2215867).

    The Court at oral argument and the parties cited amicus briefs they hoped would resonate with the Justices.  It will be interesting to see if any of those briefs or the ones discussed herein have any influence on how the Court will rule.

  • By Kevin E. Noonan –

    Supreme Court Building #1"Hope springs eternal [in the human breast]" (Alexander Pope) and "Insanity is doing the same thing over and over and expecting different results" (the latter attributed variably to Albert Einstein and Werner Erhart) are two aphorisms that irresistibly come to mind with the recent filing of a petition for certiorari by patentees in CareDX, Inc. v. Natera, Inc. and CareDx, Inc. v. Eurofins Viracor, Inc.

    To recap, the case arose over CareDx's assertion of the claims in U.S. Patent Nos. 8,703,652, 9,845,497, and 10,329,607 directed to "methods to help predict the status or outcomes of transplant recipients through sequencing of cell-free nucleic acids ("cfDNA") found in the bodily fluids of a recipient."  The rationale behind the invention was that rejection of a transplanted organ in a recipient is accompanied by cell death, which releases donor-specific DNA into the recipient's bodily fluids.  Claim 1 of the '652 patent, claim 1 of the '497 patent, and claim 1 of the '607 patent were illustrative:

    Claim 1 of the '652 patent recites:

    1.  A method for detecting transplant rejection, graft dysfunction, or organ failure, the method comprising:
        (a) providing a sample comprising cell-free nucleic acids from a subject who has received a transplant from a donor;
        (b) obtaining a genotype of donor-specific polymorphisms or a genotype of subject-specific polymorphisms, or obtaining both a genotype of donor-specific polymorphisms and subject-specific polymorphisms, to establish a polymorphism profile for detecting donor cell-free nucleic acids, wherein at least one single nucleotide polymorphism (SNP) is homozygous for the subject if the genotype comprises subject-specific polymorphisms comprising SNPs;
        (c) multiplex sequencing of the cell-free nucleic acids in the sample followed by analysis of the sequencing results using the polymorphism profile to detect donor cell-free nucleic acids and subject cell-free nucleic acids; and
        (d) diagnosing, predicting, or monitoring a transplant status or outcome of the subject who has received the transplant by determining a quantity of the donor cell-free nucleic acids based on the detection of the donor cell-free nucleic acids and subject cell-free nucleic acids by the multiplexed sequencing,
        wherein an increase in the quantity of the donor cell-free nucleic acids over time is indicative of transplant rejection, graft dysfunction or organ failure, and wherein sensitivity of the method is greater than 56% compared to sensitivity of current surveillance methods for cardiac allograft vasculopathy (CAV).

    Claim 1 of the '497 patent recites:

    1.  A method of detecting donor-specific circulating cell-free nucleic acids in a solid organ transplant recipient, the method comprising:
        (a) genotyping a solid organ transplant donor to obtain a single nucleotide polymorphism (SNP) profile of the solid organ transplant donor;
        (b) genotyping a solid organ transplant recipient to obtain a SNP profile of the solid organ transplant recipient, wherein the solid organ transplant recipient is selected from the group consisting of: a kidney transplant, a heart transplant, a liver transplant, a pancreas transplant, a lung transplant, a skin transplant, and any combination thereof;
        (c) obtaining a biological sample from the solid organ transplant recipient after the solid organ transplant recipient has received the solid organ transplant from the solid organ transplant donor, wherein the biological sample is selected from the group consisting of blood, serum and plasma, and wherein the biological sample comprises circulating cell-free nucleic acids from the solid organ transplant; and
        (d) determining an amount of donor-specific circulating cell-free nucleic acids from the solid organ transplant in the biological sample by detecting a homozygous or a heterozygous SNP within the donor-specific circulating cell-free nucleic acids from the solid organ transplant in at least one assay,
        wherein the at least one assay comprises high-throughput sequencing or digital polymerase chain reaction (dPCR), and
        wherein the at least one assay detects the donor-specific circulating cell-free nucleic acids from the solid organ transplant when the donor-specific circulating cell-free nucleic acids make up at least 0.03% of the total circulating cell-free nucleic acids in the biological sample.

    Claim 1 of the '607 patent recites:

    1.  A method of quantifying kidney transplant-derived circulating [cfDNA] in a human kidney transplant recipient, said method comprising:
        (a) providing a plasma sample from said human kidney transplant recipient, wherein said human kidney transplant recipient has received a kidney transplant from a kidney transplant donor, wherein said plasma sample from said human kidney transplant recipient comprises kidney transplant-derived circulating [cfDNA] and human kidney transplant recipient-derived circulating [cfDNA];
        (b) extracting circulating [cfDNA] from said plasma sample from said human kidney transplant recipient in order to obtain extracted circulating [cfDNA], wherein said extracted circulating [cfDNA] comprises said kidney transplant-derived circulating [cfDNA] and human kidney transplant recipient-derived circulating [cfDNA];
        (c) performing a selective amplification of target [DNA] sequences, wherein said selective amplification of said target [DNA] sequences is of said extracted circulating [cfDNA], wherein said selective amplification of said target [DNA] sequences amplifies a plurality of genomic regions comprising at least 1,000 single nucleotide polymorphisms, wherein said at least 1,000 single nucleotide polymorphisms comprise homozygous single nucleotide polymorphisms, heterozygous single nucleotide polymorphisms, or both homozygous single nucleotide polymorphisms and heterozygous single nucleotide polymorphisms, and wherein said selective amplification of said target deoxyribonucleic acid sequences is by polymerase chain reaction (PCR);
        (d) performing a high throughput sequencing reaction, wherein said high throughput sequencing reaction comprises performing a sequencing-by-synthesis reaction on said selectively-amplified target [DNA] sequences from said extracted circulating [cfDNA], wherein said sequencing-by-synthesis reaction has a sequencing error rate of less than 1.5%;
        (e) providing sequences from said high throughput sequencing reaction, wherein said provided sequences from said high throughput sequencing reaction comprise said at least 1,000 single nucleotide polymorphisms; and
        (f) quantifying an amount of said kidney transplant-derived circulating [cfDNA] in said plasma sample from said human kidney transplant recipient to obtain a quantified amount, wherein said quantifying said amount of said kidney transplant-derived circulating [cfDNA] in said plasma sample from said human kidney transplant recipient comprises using markers distinguishable between said human kidney transplant recipient and said kidney transplant donor, wherein said markers distinguishable between said human kidney transplant recipient and said kidney transplant donor comprises single nucleotide polymorphisms selected from said at least 1,000 single nucleotide polymorphisms identified in said provided sequences from said high throughput sequencing reaction, and wherein said quantified amount of said kidney transplant-derived circulating [cfDNA] in said plasma sample from said human kidney transplant recipient comprises at least 0.03% of the total circulating [cfDNA] from said plasma sample from said human kidney transplant recipient.

    The District Court granted Natera's motion for summary judgment that the claims were invalid under Section 101 for lack of subject matter eligibility (the petition noting that the District Court characterized the state of Section 101 law as being "fraught, incoherent, unclear, inconsistent, and confusing, and indeterminate and often leading to arbitrary results"), and the Federal Circuit affirmed.  The panel found that the claims failed the first prong of the Alice eligibility test for being directed to a natural phenomenon and failed the second prong of the test by reciting only conventional, well-understood, and routine methods that did not rise to the ineluctable "something more" required for eligibility.  The Federal Circuit relied upon disclosure in the specification that was similar to the disclosure that supported the Court's affirmance of ineligibility in Ariosa v Sequenom, specifically that the disclosed methods applied to detect cfDNA specific for the transplanted organ were conventional, the panel citing the following disclosure from the '652 patent in support of their conclusions:

    • col. 9 ll. 8–14, which stated that "[d]etection, identification and/or quantitation of the donor-specific markers (e.g.[,] polymorphic markers such as SNPs) can be performed using real-time PCR, chips (e.g., SNP chips), high throughput shotgun sequencing of circulating nucleic acids (e.g.[,] [cfDNA]), as well as other methods known in the art");

    • col. 10 ll. 11–12, which stated that, to obtain cfDNA samples, "any technique known in the art may be used, e.g. a syringe or other vacuum suction device");

    • col. 13 ll. 51–53, which stated that step 2 of claimed methods can be performed "using existing genotyping platforms know[n] in the art");

    • col. 15 ll. 6–8, which stated that techniques recited in step 2 of claimed methods "can be accomplished through classic Sanger sequencing methods which are well known in the art");

    • col. 13 ll. 58–61, which stated that "[c]ompanies (such as Applied Biosystems, Inc.) currently offer both standard and custom designed TaqMan probe sets for SNP genotyping that can in principle target any desired SNP position for a PCR based assay");

    • col. 20 ll. 31–34 (stating that genotyping recited in claimed methods "may be performed by any suitable method known in the art including those described herein such as sequencing, nucleic acid array or PCR");

    • col. 15 ll. 22–65 (discussing commercial high throughput sequencing products);

    • col. 14 ll. 58–67 (citing articles from 2006 and 2007 as supporting the statement that "digital PCR is a much more accurate and reliable method to quantitate nucleic acid species");

    • col. 18 l. 55–col. 19 l. 2 (stating that "[m]ethods for quantifying nucleic acids," including high throughput genotyping, "are known in the art"); and

    • col. 21 ll. 5–9 (stating that "[t]he presence or absence of one or more nucleic acids from the transplant donor in the transplant recipient may be determined by any suitable method known in the art including those described herein such as sequencing, nucleic acid arrays or PCR").

    (Despite this litany, the petition faults the District Court for relying on a sole statement in the specification that "the methods 'employ[], unless otherwise indicated, conventional techniques'" and disregarding the qualifier "unless otherwise indicated" in assessing the purported conventionality of the disclosed detection methods.)  The opinion stated summarily that "[t]he claimed methods are indistinguishable from other diagnostic method claims the Supreme Court found ineligible in Mayo and that we found ineligible on multiple occasions," including Athena Diagnostics, Inc. v. Mayo Collaborative Servs., LLC, 915 F.3d 743 (Fed. Cir. 2019); Genetic Veterinary Scis., Inc. v. LABOKLIN GmbH & Co. KG, 933 F.3d 1302 (Fed. Cir. 2018); Roche Molecular Sys., Inc. v. Cepheid, 905 F.3d 1363 (Fed. Cir. 2018); Cleveland Clinic Found. v. True Health Diagnostics LLC, 859 F.3d 1352 (Fed. Cir. 2017); and Ariosa Diagnostics, Inc. v. Sequenom, Inc., 788 F.3d 1371 (Fed. Cir. 2015).

    The Federal Circuit's appreciation of the similarity to the Ariosa decision (which in some ways propelled the Court down this path of per se ineligibility) was express:

    Here, as in Ariosa, the claims boil down to collecting a bodily sample, analyzing the cfDNA using conventional techniques, including PCR, identifying naturally occurring DNA from the donor organ, and then using the natural correlation between heightened cfDNA levels and transplant health to identify a potential rejection, none of which was inventive.  The claims here are equally as ineligible as those in Ariosa.

    Petitioner CareDx and Stanford argue in their certiorari petition that the District Court and the Federal Circuit erred in this conclusion of conventionality to distinguish this case from the earlier precedent, and assert as analytical error in the Federal Circuit's application of the Supreme Court test under Mayo Collaborative Services v. Prometheus Laboratories, Inc., 566 U.S. 66 (2012), and Alice Corp. Pty. Ltd. v. CLS Bank Int'l, 573 U.S. 208, 217 (2014), consideration of conventionality (and thus Section 103 issues) in the first prong of the Mayo/Alice test.

    The focus of Petitioner's argument is that the invention is an improvement on prior art methods that were ineffective, and that it is this improvement as expressly recited in the statute ("or any new and useful improvement thereof") that supports eligibility.  According to this argument, this improvement distinguishes the invalidated claims from the natural phenomenon that cfDNA from a transplanted organ exist in a recipient's blood or that such cfDNA could be detected therein.  The petition sets forth the history of efforts to detect organ rejection in transplant patients, from "invasive and expensive tissue biopsies from the organ" to detecting increased evidence of cfDNA in patients undergoing organ rejection, including cfDNA specific for Y chromosomes or human leukocyte antigen (HLA) gene fragments, none of which were effective "after ten year of unsuccessful attempts" and all of which were admitted prior art in the common specification of the patents-in-suit.

    The petition emphasizes the specific technical solutions of "high-throughput" and "multiplex" sequencing coupled with digital polymerase chain reaction technology using single nucleotide polymorphism detection to identify transplanted organ-specific cfDNA in recipient blood.  (These choices, the petition contends, are significant because the technologies were developed after the failed attempts by others and the targets, while not new, were not the targets used in earlier, failed attempts.  It should not escape the reader's attention that these considerations are also ones relevant to non-obviousness.)  The petition also notes that patentee disclaimed discovery of the natural phenomenon and preexisting methods for measuring it.

    The petition's asserted reasons for the Court to grant certiorari frankly tell the Court that it "needs to take another Section 101 case" based on the Justices calling for the views of the Solicitor General five times in the past five years (citing Hikma Pharm. v. Vanda Pharm., Inc.; HP Inc. v. Berkheimer; Am. Axle & Mfg., Inc. v. Neapco Holdings LLC; and most recently in Tropp v. Travel Sentry, Inc. and Interactive Wearables, LLC v. Polar Electro Oy) and that the SG has counseled the Court that they should grant cert "in an appropriate Section 101 case."  This, according to the petition, is that case, even more so that Interactive Wearables or Tropp.  The reasons petitioner asserts in advocating for the Court to choose this case is that while it raises many of the same issues as in Tropp and Interactive Wearables, in this one "the problems are even worse."  Those cases are abstract-idea cases which, the petition argues "have not been the source of considerable controversy."  Not so application of the natural phenomenon eligibility exception in medical diagnostics claims, where the petition notes both the SG and the Federal Circuit have called for the Court's intervention (citing Athena Diagnostics, Inc. v. Mayo Collaborative Servs., LLC; Am. Axle & Mfg., Inc. v. Neapco Holdings LLC; Interval Licensing LLC v. AOL, Inc.).  In those cases the circuit's judges have termed the subject matter eligibility question to be "baffling," leaving them "at a loss as to how to uniformly apply § 101," that it is "near impossible to know with any certainty whether [an] invention is or is not patent eligible," and that this situation "ha[s had] a serious effect on the innovation incentive in all fields of technology."  Similar sentiments from district courts and the U.S. Patent and Trademark Office and former Directors of the agency are cited in support of this argument.  The objective evidence for this state of affairs is the Federal Circuit's track record, wherein that Court has invalidated "every single diagnostic method patent it has encountered since Mayo[]."  Moreover, the petition notes that the cumulative effect of the Federal Circuit's jurisprudence on subject matter eligibility for medical diagnostics claims is that these claims are virtually per se ineligible (including citations to Cleveland Clinic Found. v. True Health Diagnostics LLC and Roche Molecular Sys., Inc. v. CEPHEID along with those cases cited earlier in the petition).  These decisions have "powerfully undercut[] the incentive to innovate and invest in life-saving medical diagnostics" consequently.

    In addition, the petition argues that this case is one where the claimed invention was "on specific improved methods for measuring the relative proportion of the donor's DNA [that] ensures the absence of any preemption concerns" because "[p]atents that claim specific improvements upon preexisting processes for applying a previously known natural phenomenon cannot monopolize the underlying phenomenon itself, because other methods already exist to apply it and thus remain outside the scope of the patent."  These circumstances, the petition states, provide the Court with an opportunity to "reinvigorate the role of Section 101's statutory text in a manner consonant with the preemption concerns that animate this Court's precedents."  The petition notes in support for this argument that the Court has not considered the application of Section 101 to such improvements "upon a preexisting useful process" and that this case provides such an opportunity.  The petition cites the Court's decision in Tilghman v. Proctor, 102 U.S. 707 (1880), as being consistent with these arguments by being "a particular mode of bringing about [a] desired result" (and perhaps being a basis for the Court to be persuaded by them).  And petitioners emphasize that claims to such an improvement do not claim either a natural phenomenon nor preexisting methods which are expressly in the prior art.

    Finally, the way the District Court and Federal Circuit considered issues of "conventionality" for both the first and second prongs of the Mayo/Alice test (the petition argues) was error needing Supreme Court correction.  Concerns over the (im)proper application of conventionality in the Mayo/Alice test was voiced by the Solicitor General in Interactive Wearables with regard to Step 2 but is arguably more erroneous here according to the petition, where the lower courts relied on conventionality in both Step 1 and Step 2 (the Federal Circuit going so far as to state that it had "repeatedly analyzed conventionality at step one," citing Athena Diagnostics, Inc. v. Mayo Collaborative Servs., LLC and Universal Secure Registry LLC v. Apple Inc. in support of this practice).  (This argument has greater force than the corresponding argument regarding Step 2, where considerations of conventionality go hand in hand with the Supreme Court's instruction for courts to search for an "inventive concept," although the petition bravely argues that such conflation of Section 103 issues in a Section 101 determination is equally improper.)  The petition contends that the Federal Circuit's practice in this regard "collapses the entire Section 101 inquiry into 'a search for an inventive concept'" and "effectively conflat[es] Section 101 with the statutory requirements of novelty, 35 U.S.C. 102, nonobviousness, 35 U.S.C. 103, and enablement, 35 U.S.C. 112" (which while a persuasive argument for much of the patent bar may be less so for the Court, which has tended in earlier decisions to consider patent law with much less granularity; see Mayo).  And petitioners assert that in doing so the Federal Circuit "circumvented important protections guaranteed by this Court's obviousness precedents under Section 103," citing Graham v. John Deere and KSR Int'l Co. v. Teleflex Inc.  The petition characterizes the invention at issue here as being closer to an application of a law of nature as sanctioned by the Court's decision in Diamond v. Diehr than the invention in Mayo and notes that the Court held in Ass'n for Molecular Pathology v. Myriad Genetics, Inc. that "new applications of knowledge about" natural phenomena are patent-eligible.

    Finally, the petition asks the Court "at a minimum" to hold the case until the Court has decided the outcome of the Interactive Wearables and Tropp cases.

    The arguments in the petition thus thread a very narrow needle in giving the Court a reason to grant certiorari and emphasize (as has been emphasized before) the need to do so.  Earlier in the petition the Court's attention is directed albeit somewhat obliquely to the "heav[y] investment CareDx made in bringing this technology to market" and the frankly infringing behavior of both Natera and Eurofin in bringing their own "copycat" products to market.  Those hoping for a certiorari grant (and positive outcome) have reason for such hopes if the Court is listening to these circumstances and the importance and negative consequences to innovation in the medical diagnostic arts that the lower courts' interpretations of their subject matter eligibility jurisprudence has produced over the past decade.

  • By Michael Borella —

    European Union (EU) FlagAfter using a large language model, such as ChatGPT, for a while, it is not hard to image an array of nightmarish scenarios that these generative artificial intelligence (AI) programs could bring about.  While ChatGPT and its emerging rivals currently have "guardrails" — ethical limits on what it will do in response to a prompt — the bounds thereof are not well understood.  Through clever prompting, it is not hard to convince the current iteration of ChatGPT to do away with certain guardrails from time to time.  Further, the companies behind these models have not defined the extent of the guardrails, while the very structures underlying the models are well known to behave in unpredictable ways.  Not to mention what might happen if a "jailbroken" large language model is ever released to the public.

    As an example, a user might ask the model to describe terrorist attack vectors that no human has ever previously conceived of.  Or, a model might generate software code and convince a gullible user to download and execute it on their computer, resulting in personal financial information being sent to a third party.

    Perhaps one of the most relevant risks of large language models is that once they are implemented and deployed, the marginal cost of creating misinformation becomes close to zero.  If a political campaign, interest group, or government wishes to inundate social media with misleading posts about a public figure, a policy, or a law, it will be able to do so at volume without having to employ a roomful of humans.

    In 2021, the European Commission of the European Union (EU) proposed harmonized rules for the regulation of AI.  The Commission recognized both the perils and the benefits of AI and attempted to come up with a framework for regulation that employs oversight in proportion to the specific dangers inherent in certain uses of AI.  The resulting laws enacted by member states would potentially have the Brussels Effect, in that EU regulation of its own markets become a de facto standard for the rest of the world.  This is largely what happened for the EU's General Data Protection Regulation (GDPR) laws.

    But very few people saw generative AI coming or the meteoric rise of ChatGPT at the end of 2022.  Thus, the Commission is in the process of re-evaluating its rules in view of these paradigm-breaking technologies.

    The Commission's proposal places all AI systems into one of three risk levels:  (i) unacceptable risk, (ii) high risk, and (iii) low or minimal risk.  The amount of regulation would be the greatest for category (i) and the least (e.g., none) for category (iii).

    Uses of AI that create an unacceptable risk include those that violate fundamental rights, manipulate individuals subliminally, exploit specific vulnerable groups (e.g., children and persons with disabilities), engage in social scoring (evaluating the trustworthiness of persons based on their social behavior), and facilitate real-time biometric recognition for purposes of law enforcement.  These uses would be prohibited.

    A high risk AI may be classified as such based on its intended purpose and modalities of use.  There are two main types of high risk systems: (i) those intended to be used as safety component of products (e.g., within machinery, toys, radio equipment, recreational vehicles, and medical devices), and (ii) other systems explicitly listed (e.g., involving biometrics, critical infrastructure, education, employment, law enforcement, and immigration).  These categories are quite broad and would impact many diverse industries.  The proposal sets forth detailed legal requirements for such systems relating to data governance, documentation and recording keeping, transparency and provision of information to users, human oversight, robustness, accuracy, and security, as well as conformity assessment procedures.

    Regarding low or minimal risk AI systems, their use would be permitted with no restrictions.  However, the Commission envisions these systems potentially adhering to voluntary codes of conduct relating to transparency concerns.

    To that point, the proposal also states that "[t]ransparency obligations will apply for systems that (i) interact with humans, (ii) are used to detect emotions or determine association with (social) categories based on biometric data, or (iii) generate or manipulate content ('deep fakes')."  In these situations, there is an obligation to disclose that the content has been machine-generated in order to allow the users to make informed choices.

    Currently, the Commission is considering whether to place ChatGPT and its ilk in the high risk category, thus subjecting it to significant regulation.  There has been pushback, however, from parties who believe that the regulations should distinguish between harmful uses of these models (e.g., spreading misinformation) and minimal-risk uses (e.g., coming up with new recipes, composing funny poems).  In other words, the amount of regulation that applied to ChatGPT should vary based on its use — and aesthetically pleasing goal but one that would be difficult to carry out in practice because of the model's broad scope and general applicability.

    Whether this results in the proposed regulations being delayed and/or rewritten remains to be seen.  The Commission will be taking up the issue.

  • By Michael Borella —

    Federal Circuit SealThe patent statute requires that, to be patentable, the subject matter of an invention must be at least one of a process, machine, article of manufacture, or composition of matter.  It is hard to find examples of things that do not fall into these broad categories, though signals in motion and data at rest are two.  The former has been well-litigated at this point and it is accepted that various forms of computer-readable medium (CRM) claims must recite that the medium is non-transitory.  Consider the 2007 Federal Circuit decision of In re Nuijten as the standard bearer in this regard.

    Nonetheless, disputes over the interpretation of CRM language do pop up from time to time, as was the situation in this case.

    Sequoia asserted U.S. Patent No. 6,718,436 against Dell and several other companies (most notably, Red Hat, which is a subsidiary of IBM) in the District of Delaware.  The parties butted heads over claim construction issues in district court with Sequoia coming up on the losing end.  Thus, they stipulated non-infringement under this construction.  The District Court also found that claims 8-10 were ineligible under § 101 due to construction of the term "computer-readable recording medium" to include transitory media.  Sequoia appealed.

    Claim 8 of the '436 patent reads:

    8.    A computer-readable recording medium storing instructions for executing a method for managing a logical volume in order to support dynamic online resizing and minimizing a size of metadata, said method comprising the steps of:
        a) creating the logical volume by gathering disk partitions in response to a request for creating the logical volume in a physical storage space;
        b) generating the metadata including information of the logical volume and the disk partitions forming the logical volume and storing it the metadata to the disk partitions forming the logical volume;
        c) dynamically resizing the logical volume in response to a request for resizing, and modifying the metadata on the disk partitions forming the logical volume; and
        d) calculating and returning a physical address corresponding to a logical address of the logical volume by using mapping information of the metadata containing information of the physical address corresponding to the logical address;
        wherein the metadata includes,
            a disk partition table containing information of a disk partition in which the metadata is stored;
            a logical volume table for maintaining the information of the logical volume by storing duplicated information of the logical volume onto all disk partitions of the logical volume;
            an extent allocation table for indicating whether each extent in the disk partition is used or not used; and
            a mapping table for maintaining a mapping information for a physical address space corresponding to a logical address space which is a continuous address space equal in size of storage space to an entirety of said logical volume.

    As noted, the main part of the § 101 dispute was over the interpretation of the term "computer-readable recording medium."  The Federal Circuit immediately noted that this term explicitly recites a "recording medium storing instructions" and that "a person of ordinary skill would not understand transitory signals, such as carrier waves, to record or store instructions in memory systems."  This understanding is supported by other claim elements, such as "creating the logical volume in a physical storage space" and "storing [sic] the metadata to the disk partitions forming the logical volume."  All of this establishes that the computer-readable recording medium of claim 8 does not encompass non-persistent or transient storage.

    The Court found further support for its position in the specification.  Particularly, the specification provides several examples of hardware-only computer-readable media including RAM, CDROM, and various types of disk drives.

    One of the defendants, Red Hat, argued that the specification does not exclude transitory media.  But the Court pushed back, noting that the claim itself recites a "storage medium" and that Red Hat's proposed interpretation would contradict the teachings of the specification and render the invention inoperable.

    The Court also found that Red Hat's expert's testimony was "inconsistent with the intrinsic evidence and also based on different express definitions of CRM in patent specifications directed to different inventions."  Notably, the expert looked to 34 other patent applications to help define the claim term.  The Court was not amused:

    This evidence merely shows that in thirty-four other specifications, the inventors chose to be their own lexicographers and expressly defined CRM or like terms to include transitory media.  The inventors here chose otherwise.  That other inventors chose to be their own lexicographers and define CRM to include transitory signals does not demonstrate what CRM means in the context of the '436 patent.  Nor does it establish the plain and ordinary meaning of the claim term "computer-readable recording medium for storing."

    In a similar manner, Red Hat also relied on the Court's decision in Mentor Graphics Corp. v. EVE-USA, Inc., where the Court found that a claimed "computer readable medium" included transitory signals.  But this conclusion was based on that specification expressly including carrier waves in its definition of the term.  Accordingly, the Court found that how a term is defined other patents and applications cannot be used to contradict how it is defined in the specification at hand.  Specifically, the Court wrote "[s]imply put, extrinsic evidence of what other inventors chose to do cannot surmount the intrinsic evidence of what the inventors chose here; context is key in claim construction."  Thus, the Court discounted the testimony of Red Hat's expert and the relevance of these extrinsic documents.

    Finally, Red Hat pointed to the USPTO's 2010 memo on computer-readable medium claims, alleging that it establishes that the term in question should be interpreted broadly enough to include transitory media.  But, as the Court pointed out, this memo merely states that the broadest reasonable interpretation of a claim during prosecution may result in claims being interpreted to cover transitory media.  However, this does not provide the plain and ordinary meaning of the term to be used in litigation, nor does it mean that there is a presumption that claim 8 reads on transitory media.

    Given all of this, the Court concluded that the District Court erred, and reversed the finding of invalidity under § 101.

    A practice note from all of this is that you should explicitly recite in your CRM claims language that clearly establishes that the CRM is non-transitory.  This does not need to be the exact words "non-transitory" but however your language is defined in the specification should make that point clear and unambiguous.

    Unlike the claimed invention, this victory for Sequoia turned out to be transitory — it lost on other claim construction issues and the Court ultimately affirmed the determination of non-infringement.

    Sequoia Technology LLC v. Dell Inc. (Fed. Cir. 2023)
    Panel: Circuit Judges Lourie, Dyk, and Stoll
    Opinion by Circuit Judge Stoll

  • CalendarApril 25, 2023 – "Amendments to Patent Claims — Global Updates" (Dannemann Siemsen) – 8:30 am (ET)

    April 25, 2023 – Listening session on current state of artificial intelligence (AI) technologies and inventorship issues (U.S. Patent and Trademark Office) – 10:30 am to 3:30 pm (ET), Alexandria, VA

    April 25, 2023 – "Using Disclaimers at the EPO: A Practical Guide" (J A Kemp) – 16:00 pm (BST)

    April 25, 2023 – "Women and Intellectual Property: Accelerating Innovation and Creativity" (IPWatchdog and IP.com) – 12:00 pm (ET)

    April 26, 2023 – "MentorshIP: The secret sauce for women entrepreneurs" (U.S. Patent and Trademark Office) – 12:00 pm to 1:00 pm (ET), Alexandria, VA

    April 26, 2023 – "Women in IP: Opportunities & Challenges" (Center for Intellectual Property, Information & Privacy Law at the University of Illinois Chicago School of Law) – 12:00 pm to 1:30 pm (CDT)

    April 27, 2023 – "Strategies for Adding Value and Building a Strong Biotech Patent Portfolio" (IPWatchdog and CAS) – 12:00 pm (ET)

  • USPTO SealThe U.S. Patent and Trademark Office will be holding a listening session to seek stakeholder input on the current state of artificial intelligence (AI) technologies and inventorship issues that may arise in view of the advancement of such technologies.  The event is being held from 10:30 am to 3:30 pm (ET) on April 25, 2023 at the National Inventors Hall of Fame Museum at the USPTO Headquarters in Alexandria, VA.  An agenda for the event can be found here.

    Those interested in registering for the event, can do so here.

  • Dannemann SiemsenDannemann Siemsen will be offering a webinar entitled "Amendments to Patent Claims — Global Updates" on April 25, 2023 at 8:30 am (ET).  Ankush Verma of Remfry & Saga; Alexander Wyrwoll of Winter, Brandl; and Monique Rodrigues Teixeira, Gustavo de Freitas Morais, and Patricia Porto of Dannemann Siemsen will discuss issues on amendments to patent claims in Brazil and other jurisdictions, including the following topics:

    • The allowable scope and extent of the amendments to a patent application claim;
    • The time limit for making claim amendments; and
    • The treatment given to this subject among the different jurisdictions.

    Those wishing to register for the webinar can do so here.

  • J A KempJ A Kemp will be offering a webinar entitled "Using Disclaimers at the EPO: A Practical Guide" on April 25, 2023 at 16:00 pm (BST).  Chris Milton and Imogen Parry of J A Kemp will review the current state of the law surrounding the use of disclaimers at the EPO, and then consider examples of how these can be used to your advantage, both when pursuing patent protection and when attacking problematic patents.  The webinar will address the following topics:

    • Considering when disclaimers may be appropriate at the EPO
    • The state of EPO case law concerning allowability of disclaimers
    • Preparing a case where disclaimers may be needed: drafting tips
    • Attacking weak disclaimers
    • Case studies
    • Audience questions

    There is no registration fee for this webinar.  However, those interested in registering for the webinar, should do so here.

  • IPWatchdogIPWatchdog and IP.com will be offering a webinar entitled "Women and Intellectual Property: Accelerating Innovation and Creativity" on April 25, 2023 at 12:00 pm (ET).  Renée Quinn of IPWatchdog, Inc. will moderate a panel consisting of Nina Archie of the Department of Defense Office of Small Business Program, Alison Erickson of Hallmark, Susanne Hollinger of Newell Brands, and Marlene Valderrama of Halliburton, who will share their achievements, insights, and perspectives on the IP industry.  The webinar will explore the challenges that these women have overcome, their leadership and accomplishments, and their outlook on the industry.  The panel will also discuss the importance of encouraging more women to use the IP system to protect and add value to their work and how IP can help woman-led businesses support economic recovery and build a stronger future.

    There is no registration fee for this webinar.  However, those interested in registering for the webinar, should do so here.