• San Francisco #2 American Conference Institute (ACI) will be holding the West Coast Edition of its Advanced Forum on Life Sciences Collaborative Agreements and Acquisitions from July 20-21, 2011 in San Francisco, CA.  The conference will allow attendees to:

    • Position their companies in light of emerging trends in deal structuring and increasing M&A activity;
    • Optimize business development strategies by incorporating the key takeaways from recent life sciences deals;
    • Structure option deals including multiple staged acquisitions and hybrid deals to meet both parties' objectives;
    • Negotiate milestone payments and other essential critical terms to maximize profitability;
    • Raise capital by cultivating lucrative relationships with key industry players including universities, research institutions, non-profits, and government organizations;
    • Strengthen market power and bargaining position through an effective due diligence analysis;
    • Protect assets and minimize risk by including critical termination provisions;
    • Forge strategic alliances to facilitate continued growth and sustainability in emerging markets; and
    • Maintain effective alliance management to ensure mutually beneficial influx of capital and resources.

    Brochure In particular, ACI's faculty will offer presentations on the following topics:

    • Market watch: Understanding the driving forces and changing dynamics impacting the current deal-making landscape;
    • Integrating increasing M&A activity into your business development strategy;
    • Best practices for selecting the deal structure and negotiating your most beneficial collaborative agreement yet;
    • Negotiating options, staged acquisitions, hybrid transactions and straight-forward M&As;
    • Streamlining entry into lucrative emerging markets via strategic partnering;
    • Beyond the shadow of the valley: Alternative pathways and partnerships to facilitate funding drug discovery and development;
    • Preparing for the exit: Proactively drafting critical termination provisions to protect company assets;
    • Protecting current and future collaborations: The do's and don'ts of partnering / negotiations;
    • Aligning divergent interests to negotiate mutually beneficial collaborative research agreements with academic institutions;
    • Utilizing applicable valuation models to set realistic expectations for a deal;
    • Mitigating legal risks in the life sciences M&A setting: A six-step approach; and
    • Licensing strategies for biosimilars: Regulatory, commercial and IP considerations for the new frontier.

    A post-conference master class, entitled "Life Sciences IP Due Diligence Boot Camp: Conducting Effective & Strategic Due Diligence for Life Science Partnering and M&A," will be offered on July 22, 2011.  In this workshop, ACI faculty will offer presentations on the following topics:

    • Spotting red flags that impact the value and success of a life sciences transaction;
    • Factoring recent relevant IP developments into your due diligence analysis: Biosimilars, Myriad, Prometheus, and more  — to be presented in part by Patent Docs author Dr. Donald Zuhn;
    • Properly aligning the business objective of the deal and the IP diligence assessment: An interactive checklist;
    • Evaluating the scope, breadth, validity and enforceability of the target's patents under evolving patent standards and regulatory protocols;
    • Who invented what when? Reducing and resolving inventorship disputes; and
    • Ensuring that the purchaser/licensee has the right to commercialize the IP at issue.

    The agenda for the Life Sciences Collaborative Agreements and Acquisitions conference can be found here.  A complete brochure for this conference, including an agenda, detailed descriptions of conference sessions, list of speakers, and registration form can be obtained here.

    ACI - American Conference Institute The registration fee for the conference is $2,295 (conference alone), $3,095 (conference and master class), or $1,395 (master class alone).  Those registering by May 20, 2011 will receive a $300 discount off the conference alone or conference and master class, and those registering by June 24, 2011 will receive a $200 discount off the conference alone or conference and master class.  Those interested in registering for the conference can do so here, by calling 1-888-224-2480, or by faxing a registration form to 1-877-927-1563.

    Patent Docs is a media partner of ACI's Life Sciences Collaborative Agreements and Acquisitions conference.

  • By Kevin E. Noonan —
    Endo Pharmaceuticals The Federal Circuit continued its explication both of the contours of obviousness for pharmaceutical formulations after KSR Int'l Co. v. Teleflex Inc. as well as how it exercises its supervisory powers over the U.S. Patent and Trademark Office after Dickinson v. Zurko, in a decision last Friday, In re Huai-Hunk Kao et al.  The subject matter was controlled-release oxymorphone formulations, assigned to Endo Pharmaceuticals Inc. in three applications:  U.S. Application Nos. 11/680,432 (Kao, Baichwal, McCall and Lee, inventors), 12/167,859 (Kao, Baichwal, McCall and Lee, inventors), and 11/766,740 (Ahdieh, inventor).  All claims at issue were rejected as being obvious.

    Oxymorphone Controlled-release formulations are important for opioid drugs such as oxymorphone (at left) because these drugs undergo significant "first pass" metabolism by the liver, and repeated administration of immediate release formulations have deleterious side effects.  The three applications rejected by the U.S. Patent and Trademark Office provided controlled-release formulations that overcame these limitations.

    All the applications had been filed under the PTO's Accelerated Examination Program, which limited the claims that were argued on appeal.  Claim 1 of the '432 application reads as follows:

    1.  An analgesically effective controlled release pharmaceutical composition with a twelve hour dosing interval in the form of a tablet, comprising oxymorphone or a pharmaceutically acceptable salt thereof as the sole active ingredient in the tablet and a controlled release delivery system comprising at least one pharmaceutical excipient, wherein upon placement of the composition in an in vitro dissolution test comprising USP Paddle Method at 50 rpm in 500 ml media having a pH of 1.2 to 6.8 at 37oC, about 15% to about 50%, by weight, of the oxymorphone or salt thereof is released from the tablet at about 1 hour in the test.

    This claim was rejected for being obvious over International Publication No. WO 01/08661 ("the Maloney reference"), alone or in combination with U.S. Patent No. 5,047,248.  According to the Examiner, the Maloney reference discloses every element of the claim except the dissolution rate, and the Examiner required Applicants to demonstrate that the practice of the formulations disclosed by Maloney did not (inherently?) produce that dissolution rate.  In response, Applicants submitted declarations distinguishing the Maloney reference on the grounds that it disclosed oxycodone that showed a "markedly different bioavailability profile" than oxymorphone.  In addition, the method used to demonstrate the dissolution rate in the Maloney reference was different from the method used by Applicants, and the dissolution rates were not comparable.  Finally, Applicants asserted that their claimed formulations showed an unexpected result — multiple peaks in blood concentration of the drug using the claimed formulation as well as commercial success of Endo's drug product.  The Examiner, affirmed by the Board, rejected these arguments and distinctions, finding that the claims were obvious because the Maloney reference disclosed oxymorphone formulations as "preferred embodiments" and the dissolution rates in the reference overlapped with the range of rates in the '432 application claims.  The Board also rejected Applicants' profferred evidence of secondary considerations because that evidence was not "commensurate in scope" with the claimed invention.

    The '859 application claimed methods of relieving pain using controlled release delivery of oxymorphone:

    8.  A method for treating pain in a human subject in need of acute or chronic pain relief, comprising the steps of:
        (a)  providing a solid oral dosage form comprising about 5 mg to about 80 mg oxymorphone or a pharmaceutically acceptable salt thereof in a controlled release delivery system with a release rate profile designed to provide an adequate blood plasma level over at least 12 hours to provide sustained pain relief over this same period, the sys- tem comprising a filler and a hydrophilic material, wherein oxymorphone is the sole active ingredient; and,
        (b)  administering the dosage form to the subject, wherein the oxymorphone Cmax is at least about 50% higher when the dosage form is administered to the subject under fed versus fasted conditions.

    The Maloney reference was also asserted by the Examiner against the claims of the '859 application in support of an obviousness determination, substantially consistent with the ground asserted against the '432 application claims.  The Examiner, affirmed by the Board also discounted evidence of secondary considerations because that evidence was not commensurate in scope with the '859 application claims.

    The claims of the '740 application were also rejected for obviousness:

    21.  A method of providing extended pain relief to patients in need thereof, comprising:
        providing information that the average bioavailability of oxymorphone in an oral extended release formulation designed to have a 12 hour dosing cycle is increased by at least about 26% for subjects with renal impairment compared to that for healthy subjects, and
        providing a therapeutically effective amount of such an extended release oral dosage form of oxymorphone or its pharmaceutically acceptable salt thereof.

    The obviousness rejection of these claims was based on the Maloney reference, in light of U.S. Patent Application Publication No. 2002/0032581, which disclosed a clinical evaluation kit.  As with the other two applications, evidence regarding secondary considerations was not persuasive in overcoming the obviousness rejections.
    Federal Circuit Seal The Federal Circuit, in an opinion by Judge Linn, joined by Judge Moore and Chief Judge Rader, vacated and remanded the Board's obviousness rejection of the '432 application claims but affirmed the rejection of the '859 and '740 applications.  The Board's opinion affirming rejection of the '432 application claims on obviousness was vacated as not being supported by substantial evidence.  The PTO had based its obviousness determination on the disclosure in the Maloney reference that the formulation being prepared with oxymorphone as well as the "equivalence" of the two types of dissolution tests used by the art and the Applicants.  The Federal Circuit opinion accepts the Office's contention that it would have been obvious to substitute oxymorphone for oxycodone based on the disclosure of the reference that oxymorphone is a preferred embodiment.  However, where the Office's argument fails according to the opinion is in the determination that the formulation disclosed in the Maloney reference would have the dissolution properties required in the claim.  This is due to the differences in the assay method (the "paddle" method versus the "basket" method) and the failure of the Office to establish this by substantial evidence, particularly in view of the declaration from the Applicant to the contrary.  The deficiency in the Board's decision is that "there is a lack of direct factual support in the record for the view that the claimed range of dissolution rates actually overlaps with the dissolution rate disclosed in [the] Maloney [reference], a premise upon which the Board's reasoning is founded."

    The Office tried to use evidence to support the correlation from a declaration submitted by Applicants, but erred according to the opinion because it:

    [R]elied on four data points from an exhibit correlating the dissolution rates of Opana® ER, when tested by both methods, and Maloney's Formula 6, tested only by the Basket Method, and concluded, without any reasoning, that because the Basket Method dissolution in the first hour for Opana® ER was 1.3 times faster than the dissolution rate of the Paddle Method for Opana® ER, this correlation would also hold for Maloney's Formula 6.

    This conclusion was formed in the face of express statements from the declarant that "there is no general correlation between the Basket and Paddle Methods," a statement supported by scientific literature citations.  The opinion states that the record is devoid of any evidence or reasoning why the declarant's statements are not accurate.  Since "substantial evidence" is "such evidence as a reasonable mind might accept as adequate to support a conclusion" (citing Consol. Edison Co. v. Nat'l Labor Relations Bd., 305 U.S. 197, 229 (1938)), the Office's unsupported conclusion does not meet the standard according to the Federal Circuit.

    However, the opinion also states that the importance of the dissolution range in the claim to the obviousness issue (i.e., whether this distinction would be sufficient to render the claims non-obvious) was not before the Court, and the panel directed the Board to consider this question on remand, citing to Ormco Corp. v. Align Technology, Inc., 463 F.3d 1299, 1311 (Fed. Cir. 2006), for the principle that "overlap between [a] claimed range and [the] prior art gives rise to a presumption of obviousness where claimed range and prior art value are insubstantially different."  The panel also cited to one of the Supreme Court's most cogent statements of its obviousness jurisprudence in this regard:

    When there is . . . market pressure to solve a problem and there are a finite number of identified, predictable solutions, a person of ordinary skill in the art has good reason to pursue the known options within his or her technical grasp.  If this leads to the anticipated success, it is likely the product not of innovation but of ordinary skill and common sense.  KSR Intern. Co. v. Teleflex Inc., 550 U.S. 398, 402-403 (2007).

    With regard to secondary considerations, the Board was not persuaded that Applicants' evidence of secondary considerations raised in support of the non-obviousness of any of the claims at issue, on the grounds that the evidence of unexpected results and commercial success were "not commensurate with the scope of the claims."  The panel opinion stated that the Office must consider evidence of secondary considerations when asserted, and that the evidence thereof "must be reasonably commensurate with the scope of the claims," citing In re Tiffin, 448 F.2d 791, 792 (CCPA 1971), and In re Hiniker, 150 F.3d 1362, 1369 (Fed. Cir. 1998).  While this does not require an applicant to "test every embodiment" within the scope of the claim to demonstrate secondary considerations, there must be sufficient evidence presented that would "show a correlation" throughout the scope of the claims according to the Court, citing In re Greenfield, 571 F.2d 1185, 1189 (CCPA 1978), and In re Cescon, 474 F.2d 1331, 1334 (CCPA 1973).

    However, the Court characterized as "a more fundamental requirement" that there be a nexus between the evidence asserted and the "merits of the claimed invention."  Wyers v. Master Lock Co., 616 F.3d 1231, 1246 (Fed. Cir. 2010) (emphasis in original).  Thus, the opinion states that where the proffered "secondary consideration" is the consequence of something "other than what is both claimed and novel in the claim," a nexus is not established, citing Tokai Corp. v. Easton Enters., Inc., 632 F.3d 1358, 1369 (Fed. Cir. 2011), Ormco Corp., and In re Woodruff, 919 F.2d 1575, 1578 (Fed. Cir. 1990).

    For the '432 patent, the panel found that there was not substantial evidence (indeed, there was no evidence) supporting the Board's conclusion that the asserted unexpected result (that the formulation caused "multiple peaks of oxymorphone blood concentration") "must be caused by some component of that particular disclosed controlled-release formulation."  This formed another basis for remand to the Board to "determine whether there is a nexus between the unexpected in vivo concentration profile and aspects of the claimed invention not already present in the prior art."  "[F]or the unexpected in vivo concentration profile of the applicant's product to have substantial weight, there must be a nexus to some aspect of the claim not already in the prior art, such as the claimed range of dissolution rates, as against other unclaimed prior-art dissolution rates."

    Regarding the evidence of commercial success, the opinion reasserted the principle that not every embodiment of a claimed invention must be sold to demonstrate unexpected results ("'[i]t seems unlikely that a company would sell a product containing multiple, redundant embodiments of the patented invention . . . . Under the [Office's] logic, there would never be commercial success evidence for a claim that covers more than one embodiment," citing In re Glatt Air Techniques, Inc., 630 F.3d. 1026, 1030 (Fed. Cir 2011)), but found that "the record is nearly silent on whether the commercial success was caused by the merits of the invention as distinct from the prior art," forming yet another factual issue to be determined by the Office on remand.

    Turning to the '859 and '740 applications, here the Court was not persuaded that the Board had erred in determining that these claims were obvious.  With regard to the '859 application, the panel agreed that the increased efficacy of administering the claimed oxymorphone formulations with food was an inherent property of the drug not limited to the claimed controlled-release formulations, relying on King Pharmaceuticals, Inc. v. Eon Labs, Inc., 616 F.3d 1267, 1275-76 (Fed. Cir. 2010), to support this proposition.  The Court also agreed that the 12-hour effectiveness limitation was disclosed in the Maloney reference.  Finally, the Court agreed with the Board, saying that "[s]ubstantial evidence supports the Board's finding that Maloney teaches a controlled release formulation using both hydrophobic and hydrophilic materials" as required by the claims at issue.

    While the Court faulted the Board for failing to give proper weight to the secondary considerations asserted in support of the non-obviousness of the claims of the '859 patent, here the panel affirmed the obviousness rejection because there was "a strong showing of obviousness," consistent with the Court's post-KSR practice of discounting secondary considerations when the evidence supporting an obviousness rejection seems compelling (see "Pharmastem Therapeutics, Inc. v. Viacell, Inc. (Fed. Cir. 2007)"; This is an ironic treatment of the secondary, or "objective" indicia of non-obviousness, which were intended by the Supreme Court under Graham v. John Deere Co. to rebut an obviousness determination under just those circumstances where unavoidable hindsight might lead to an erroneous conclusion.)

    Finally, the opinion addresses the obviousness rejection of the '740 application, which was also affirmed.  The Applicant asserted that the "information" regarding an "undiscovered correlation between renal failure and bioavailability" rendered the claims non-obvious.  The panel considered this argument to be the same argument it rejected in King Pharmaceuticals, which also involved increasing oral bioavailability using information unappreciated in the prior art (in King, administering metaxalone with food).  This case is not "meaningfully distinct," the panel states, because "informing someone of the correlation cannot confer patentability absent a functional relationship between the informing and administering steps," citing In re Ngai, 367 F.3d 1336, 1338 (Fed. Cir. 2004), and General Elec. Co. v. Jewel Incandescent Lamp Co., 326 U.S. 242, 249 (1945).  The opinion states that at least one basis for its decision regarding the obviousness of the '740 application's claim at issue is that "there is no requirement in the claim that the dosage be adjusted in response to the informing step."  In addition, "because there is no indication of who is to be informed or to whom the drug is to be administered, the claim would presumably cover a situation where a doctor informs patient A of the correlation and administers a therapeutically effective dose of controlled release oxymorphone to patient B," i.e., "there is no functional relationship between the two steps in the method."  Since merely administering controlled release oxymorphone is "squarely present in the prior art," the Board properly determined that the claims of the '740 application were obvious.

    While the Court believed that the Board had erroneously failed to give proper weight to evidence of secondary considerations with regard to the '740 application claims, this error was harmless because the Applicant failed to establish the required nexus between the claimed invention and the asserted unexpected results or commercial success.

    These results, while not inconsistent with prior precedent illustrate nicely how the Court is applying the rubrics set forth by the Supreme Court in KSR and how the outcome has changed due to that decision.  Arguably, for at least the '859 and '740 applications there was no teaching, suggestion or motivation to provide formulations having the specifically claimed components (as claimed in the '859 application) or with the benefit of understanding the correlation between bioavailability and renal failure (as claimed in the '740 application), and these claims might otherwise have issued without the renewed focus on the predictability of the claimed method (or the common sense of the skilled artisan at arriving at these methods).  Even with regard to the '432 application, the panel appears to believe it necessary to remind the Board of KSR's emphasis on predictability in deciding whether a claim is obvious.  This opinion reinforces the appearance that the Federal Circuit has understood the Supreme Court's interpretation of the appellate court's mandate and is doing what it can to comply.

    In re Kao (Fed. Cir. 2011)
    Panel:  Chief Judge Rader and Circuit Judges Linn and Moore
    Opinion by Circuit Judge Linn

  • By Donald Zuhn —

    USPTO Seal Last week, in Ex parte DeGrado, an appeal before the Board of Patent Appeals and Interferences concerning U.S. Application No. 10/801,951 (U.S. Patent Application Publication No. US 2006/0041023), the Board issued an order calling for further briefing on two issues related to a claim that the Board "conservatively estimate[s] . . . encompasses in excess of 400 billion oligomers."  In particular, the Board is requiring Applicants to brief the following questions:

    1.  Whether Applicants may be required to restrict their claims to a single invention under the provisions of 35 U.S.C. § 121.

    2.  Whether Claim 16 is a proper "Markush Claim."

    Aside from the issues for which the Board seeks further briefing, the appeal is notable in that the panel consists of the Commissioner of Patents.

    In its order, the Board notes that claim 16 relates to a method for treating microbial infections by administering a compositions including at least one amphiphilic oligomer, and that the amphiphilic oligomers falling within the scope of the claims are defined by the formula R1-[-x-A1-x-y-A2-y-]m-R2 (wherein the definitions of R1, R2, x, y, A1, A2, and m take up three pages of the 12-page order).  Reviewing the prosecution history, the Board stated that the application as originally filed contained seven independent claims, five directed to methods of treating microbial infections by administering oligomers and two directed to groups of oligomers.  None of the originally filed claims required that the oligomers be amphiphilic.

    The Examiner opened prosecution by issuing a restriction requirement that identified fifteen inventions and required Applicants to select one of the inventions as well as a single species of oligomer from the elected group.  Following Applicants' selection of the invention of Group III and a specific oligomer, the Examiner issued an additional restriction requirement for the invention of Group III that identified 28 classes of oligomers within Group III and required Applicants to select one of the classes as well as a single species of oligomer from the elected class.  Applicants responded by amending claim 16 to require that the oligomers be amphiphilic, and traversing the restriction and election requirement, in part, on the ground that it ignored the amphiphilicity feature of the invention, which Applicants explained gave the oligomers their anti-microbial property.  Applicants also argued that they could not elect one of the 28 classes of oligomers because none of the classes "permit the property of amphiphilicity, which is the central feature of Applicants' invention."  Applicants further argued that requiring a restriction between independent inventions present in a single claim is improper under In re Weber, 580 F.2d 455 (C.C.P.A. 1978) and In re Haas, 580 F.2d 461 (C.C.P.A. 1978), and that M.P.E.P. § 803.02 requires the Office to examine all claims exactly as presented by Applicants unless the claims lack "unity of invention."  Applicants contended that the recited amphiphilic oligomers demonstrated unity of invention because they share the common utility of antimicrobial activity and the structural feature of amphiphilicity.  Following Applicants' amendment of claim 16, the Examiner withdrew the earlier restriction requirements and issued a new restriction requirement that identified 21 inventions.  Applicants selected the same group of claims (16-48) and oligomer as selected in response to the initial restriction requirement, and again argued that the property of amphiphilicity common to all the claimed compounds and their effectiveness for treating microbial infections precluded restriction.  Applicants also argued the impropriety of restricting between multiple inventions in a single claim, noting that 35 U.S.C. § 121 "does not grant the PTO the authority to refuse to examine a single claimed invention."

    The Board notes in its order that in Weber:

    The Court held that in requiring that "[t]he specification shall conclude with one or more claims particularly pointing out and distinctly claiming the subject matter which the applicant regards as his invention," 35 U.S.C. § 112, 2nd paragraph, allows the inventor to claim the invention in the way he chooses (subject to the other requirements of law), including claiming multiple independent and distinct inventions in a single claim.

    However, the order "require[s] that Applicants brief the apparent conflict between the plain language of § 121 [which appears to empower the Director to require any "application to be restricted to one of the inventions"] and the Weber and Haas opinions.

    Citing In re Harnisch, 631 F.2d 716, 722 (C.C.P.A. 1980), and Ex parte Hozumi, 3 USPQ2d 1059, 1060 (Bd. Pat. App. & Int. 1984), the Board states that "[a] Markush claim is improper if the inventions (1) do not share a common use; or (2) do not share a 'single structural similarity,' that is, a substantial structural feature disclosed as being essential to the common utility."  Noting that the Examiner restricted claims 16-48 into 21 inventions on the basis of "the markedly different chemical structures between the oligomers in the different groups," the Board also requires additional briefing from Applicants on whether claims 16-48 are proper Markush Claims.  In particular, the Board states that:

    Applicants are required to brief whether the recitation of a broad general formula covering a very large group of compounds, the recitation of a general chemical property (amphiphilicity) that may be possessed by those compounds, and the recitation of the single broad step of "administering an effective amount" is per se sufficient to create "unity of invention" as that concept was used by the Harnisch court.

    Applicants must submit their brief by June 6.

  • By Kevin E. Noonan —

    Supreme Court Building #1 The Supreme Court refused today to grant certiorari in Eli Lilly & Co. v. Sun Pharmaceutical Industries, Ltd. on the question of obviousness-type double patenting.  In doing so, the Court let stand precedent that seriously expands the scope of the obviousness-type double patenting standard, turning a legal principle that was "clear" to one "distorted by divergent statements[] leading to [a] flawed ruling," according to Judge Newman (joined by Judges Lourie and Linn and Chief Judge Rader) in dissent from denial of Eli Lilly's petition for rehearing en banc.

    To recap, a Federal Circuit panel (opinion by Judge Prost, joined by Judges Gajarsa and Bryson) affirmed the District Court judgment that claims of Eli Lilly's U.S. Patent No. 5,464,826 ("the '826 patent) were invalid for obviousness-type double patenting over another Lilly patent, U.S. Patent No. 4,808,614 ("the '614 patent") (see Patent Docs report).  The '614 patent claims gemcitabine, and methods of using gemcitabine to treat viral infections, while the '826 patent claims methods of using gemcitabine for treating cancer.  Eli Lilly markets the drug GEMZAR®, which contains gemcitabine as the active ingredient, for the treatment of various forms of cancer.  Accordingly, both the '614 and '826 patents were listed in the Orange Book with respect to GEMZAR®.  Sun filed an ANDA in 2006 to market a generic version of GEMZAR®, and filed a declaratory judgment action in 2007, asserting, among other things, that the '826 patent was invalid.  The U.S. District Court for the Eastern District of Michigan granted Sun's motion for partial summary judgment that the asserted claims of the '826 were invalid based on obviousness-type double patenting.

    The history of these patents and their interrelationship were important factors leading to the panel decision.  The '614 patent was a continuation-in-part of a priority patent that disclosed gemcitabine and its antiviral activity.  The application that eventually gave rise to the '614 patent contained additional disclosure regarding the anticancer activities of gemcitabine but contained no claims to this activity.  The '826 patent, in an application filed on the same day as the application that resulted in the '614 patent, both disclosed and claimed this activity and methods for exploiting the anticancer properties of gemcitabine in methods for treating cancer with the drug.  Thus, the '614 patent was entitled to the priority claim to the original patent while the '826 patent was not.

    The panel opinion relied on two earlier cases, Geneva Pharm., Inc. v. GlaxoSmithKline PLC, 349 F.3d 1373 (Fed. Cir. 2003), and Pfizer, Inc. v. Teva Pharm. USA, Inc., 518 F.3d 1353 (Fed. Cir. 2008) in deciding that the '826 claims were invalid over the '614 patent in obviousness-type double patenting grounds.  These cases can be distinguished, however, since in both cases the specifications disclosed one and only one activity (or use) for the claimed compound, not (as here) an additional, new use.  Also, the panel used what was disclosed, rather than just what was claimed in the '614 patent in making its obviousness-type double patenting rejection of the '826 patent, and this is where the opinion veered significantly from prior precedent.  In doing so, the panel created a per se rule of obviousness-type double patenting that, while promoting certainty also "upsets the settled expectations of the patenting community" by significantly expanding the scope of the obviousness-type double patenting doctrine.

    Specifically, under the rule established by the panel opinion, if the earlier of two co-owned patents includes claims to a compound and discloses one or more uses for the compound in the specification, under the Federal Circuit's ruling any claim(s) in a later patent to any method of use disclosed (but not claimed) in the specification of the earlier patent are per se invalid based on obviousness-type double patenting.  This rule is contrary to prior case law, both in removing flexibility in considering the effect of the later patent on "improperly extending 'protection, beyond the date of expiration of the [earlier] patent, of the very same invention claimed therein . . . or of a mere variation of that invention which would have been obvious to those of ordinary skill in the relevant art,'" In re Kaplan, 789 F.2d 1574, 1579-80 (Fed. Cir. 1986), but also by collapsing the obviousness-type double patenting analysis into obviousness under 35 U.S.C. § 103 (using the patentee's own prior patent as if it were prior art under 35 U.S.C. § 102).  This constitutes a new, non-statutory type of prior art, a patentee's own prior patent application that was not "by another" and did not constitute a statutory bar (i.e., the patentee's own earlier work disclosed more than a year before the later application's filing date).  This is expressly contrary to the Federal Circuit's earlier precedent, e.g., General Foods Corp. v. Studiengesellschaft Kohle mbH, 972 F.2d 1272 (Fed. Cir. 1992), that a commonly owned, prior patent disclosure cited in support of a double patenting rejection cannot be used as though it were prior art.  Specifically, the earlier-filed patent disclosure is not to be considered in making an obviousness-type double patenting rejection.  In re Kaplan, 789 F.2d 1574 (Fed. Cir. 1986).

    Despite this disjunction between prior precedent and the panel opinion, the Federal Circuit declined to rehear the case en banc, in a 5-4 ruling from a diminished number of active judges.  Thus, Eli Lilly's only recourse was to appeal to the Supreme Court; however, consideration by the Court was unlikely, for several reasons.  First, the Supreme Court is a generalist court interested in the more significant, "high profile" cases that have characterized the Court's patent jurisprudence for the past decade (including KSR Int'l Co. v. Teleflex Inc., eBay Inc. v. MercExchange, LLC, Quanta Computer, Inc. v. LG Electronics, Inc., Microsoft Corp. v. AT&T Corp., Dickinson v. Zurko).  The obviousness-type double patenting doctrine lies deep in the minutiae of patent law and was unlikely to attract the Court's attention.  The Court also takes interest in patent law questions from the Federal Circuit where the court seems divided (such as Warner-Jenkinson Co. v. Hilton Davis Chemical Co. and Festo Corp. v. Shoketsu Kinzoku Kogyo Kabushiki Co.).  This question was important (for the pharmaceutical industry in general) but it will take the Federal Circuit in its entirety to choose a case where the irregularities introduced into this important patent law doctrine will be set straight.

    Patent Docs filed an amicus case in support of certiorari in this case.

  • By Donald Zuhn —

    USPTO Seal The U.S. Patent and Trademark Office announced today that it is expanding the Enhanced First Action Interview pilot program to include all utility applications in all technology areas and filing dates.  Like its predecessor programs, under the Full First Action Interview pilot program, an applicant is entitled to a first action interview, upon request, prior to the first Office action on the merits.  The new pilot program is scheduled to run through May 16, 2012.

    The Office initiated the initial First Action Interview pilot program (see "PTO Announces New Program to Reduce Pendency and Improve Patent Quality") in April 2008.  Under the original pilot program, an examiner assigned to a participating application conducted a prior art search and provided the applicant with a pre-interview communication containing a condensed preview of objections or rejections against the claims of the participating application.  Within 30 days from the issuance of the pre-interview communication, the applicant either scheduled a first action interview or decided not to have the interview.  If the applicant chose the latter option, the examiner issued a First Action Interview Office action giving the applicant the longer of one month or 30 days to reply.  If agreement could not be reached during the first action interview, the First Action Interview Office action was issued (with the reply period specified above).

    In October 2009, the Office expanded the First Action Interview pilot program to encompass additional technology areas (see "USPTO Expands First Action Interview Pilot Program").  One year after expanding the program, the Office extended the ending date of the pilot program from April 1, 2010 to October 1, 2010 (see "USPTO Announces Extension of Enhanced First Action Interview Pilot Program").  More recently, the Office revised the eligibility requirements to include utility nonprovisional applications that were filed three months later than the previously listed dates (see "USPTO Extends Enhanced First Action Interview Pilot Program").

    Until today's expansion of the program, biotech and pharma patent practitioners were limited to seeking participation for applications assigned to the 161X (drug, bio-affecting and body treating compositions) and 1795 (chemistry: electrical current producing apparatus, product, and process) art groups.  Under the Full First Action Interview pilot program, however, subject matter and filing date restrictions have been removed.  Other than expanding the program to encompass all technology areas and filing dates, the process remains unchanged from the process used during the Enhanced First Action Interview Pilot Program that ended April 1, 2011.

    In the Office's announcement, it noted that 34% of applications participating in prior iterations of the program were allowed on the first action on the merits as compared with the approximately 11% of applications that are allowed on the first action on the merits on average across all technologies for new non-continuing applications.

    Additional details regarding the Full First Action Interview pilot program can be found in a pre-OG notice issued by the Office.  This notice indicates that applicants wishing to secure a non-discretionary interview before the first action on the merits must electronically file a request (preferably using Form PTO/SB/413C).  Following this request, the Full First Action Interview process would proceed as outlined above for the original program.  To participate in the program, the following requirements must be met:

    (1)  The application must be a non-reissue, non-provisional utility application filed under 35 U.S.C. § 111(a), or an international application that has entered the national stage in compliance with 35 U.S.C. § 371(c).

    (2)  The application must contain three or fewer independent claims and twenty or fewer total claims, and no multiple dependent claims (applicants may file a preliminary amendment in order to comply with this requirement).

    (3)  The claims must be directed to a single invention (if the Office issues a restriction requirement, the application will be removed from the program; if an oral election without traverse is made during a telephone interview, the application will remain in the program).

    (4)  The request for a first action interview must be filed electronically, preferably using Form PTO/SB/413C.

    (5)  The request must be filed at least one day before a first Office action on the merits of the application appears in the Patent Application Information Retrieval (PAIR) system.

    (6)  The request for a first action interview must include a statement that applicant agrees not to file a request for a refund of the search fee and any excess claim fees paid in the application after the mailing or notification date of the Pre-Interview Communication (the Office notes that any petition for express abandonment under 37 C.F.R. § 1.138(d), request for a refund of search fee, and any excess claim fees filed after the mailing or notification date of a Pre-Interview Communication will not be granted).

    The pre-OG notice also indicates that:

    • The time period for scheduling an interview following the issuance of the Pre-Interview Communication may be extended for one additional month pursuant to 37 C.F.R. § 136(a).

    • Interviews must be scheduled electronically using the Applicant Initiated Interview Request Form (PTOL-413A).  The applicant must propose an interview date within the longer of two months or 60 days from the filing of the Applicant Initiated Interview Request Form.

    • Amendments filed between the issuance of the Pre-Interview Communication and the interview will not be entered unless approved by the examiner.  At the examiner's discretion, however, an amendment may be entered if it is clearly limited to the cancellation of claims, adoption of examiner suggestions, placement of the application in prima facie condition for allowance, or correction of informalities.

    Additional details regarding the program can be found in the pre-OG notice.

  • By Sherri Oslick —

    Gavel About Court Report:  Each week we will report briefly on recently filed biotech and pharma cases.

    Elan Corp. et al. v. Teva Pharmaceuticals USA Inc.
    1:11-cv-00367; filed April 25, 2011 in the District Court of Delaware

    • Plaintiffs:  Elan Corp. plc; Elan Pharma International Ltd.
    • Defendant:  Teva Pharmaceuticals USA Inc.

    Infringement of U.S. Patent Nos. 6,228,398 ("Multiparticulate Modified Release Composition," issued May 8, 2001) and 6,730,325 (same title, issued May 4, 2004), licensed to Novartis, following a Paragraph IV certification as part of Teva's filing of an ANDA to manufacture a generic version of Novartis' Focalin® XR (extended release dexmethylphenidate hydrochloride, used to treat attention deficit hyperactivity disorder).  View the complaint here.


    Celgene Corp. et al. v. Teva Pharmaceuticals Usa, Inc.

    2:11-cv-02356; filed April 25, 2010 in the District Court of New Jersey

    • Plaintiffs:  Celgene Corp.; Novartis Pharmaceuticals Corp.; Novartis Pharma AG
    • Defendant:  Teva Pharmaceuticals USA, Inc.

    Infringement of U.S. Patent Nos. 5,908,850 ("Method of Treating Attention Deficit Disorders with d-Threo Methylphenidate," issued June 1, 1999), 6,355,656 ("Phenidate Drug Formulations Having Diminished Abuse Potential," issued March 12, 2002, with a reexamination certificate issued March 27, 2007), 6,528,530 ("Phenidate Drug Formulations Having Diminished Abuse Potential," issued March 4, 2003), 5,837,284 ("Delivery of Multiple Doses of Medications," issued November 17, 1998), 6,635,284 (same title, issued October 21, 2003), and 7,431,944 ("Delivery of Multiple Doses of Medications," issued October 7, 2008) all licensed exclusively to Novartis in certain fields of use, following a Paragraph IV certification as part of Teva's filing of an ANDA to manufacture a generic version of Novartis' Focalin XR® (extended release dexmethylphenidate hydrochloride, used to treat attention deficit hyperactivity disorder).  View the complaint here.


    Genentech, Inc. et al. v. Sandoz Inc.

    1:11-cv-01076; filed April 22, 2011 in the District Court of Colorado

    • Plaintiffs:  Genentech, Inc.; Roche Palo Alto LLC
    • Defendant:  Sandoz Inc.

    Infringement of U.S. Patent No. 6,083,953 ("2-(2-amino-1,6-dihydro-6-oxo-purin-9-yl)methoxy-1,3-propanediol Derivative," issued July 4, 2000) following a Paragraph IV certification as part of Sandoz's filing of an ANDA to manufacture a generic version of Roche's Valcyte® (valganciclovir hydrochloride, used to treat cytomegalovirus retinitis and cytomegalovirus disease).  View the complaint here.


    Wyeth LLC v. Nostrum Pharmaceuticals, LLC et al.

    3:11-cv-02280; filed April 21, 2011 in the District Court of New Jersey

    • Plaintiff:  Wyeth LLC
    • Defendants:  Nostrum Pharmaceuticals, LLC; Nostrum Laboratories, Inc.; Enem Nostrum Remedies PVT., Ltd.

    Infringement of U.S. Patent Nos. 6,274,171 ("Extended release formulation of venlafaxine hydrochloride," issued August 14, 2001), 6,403,120 (same title, issued June 11, 2002), and 6,419,958 (same title, issued July 16, 2002) following a Paragraph IV certification as part of Nostrum's filing of an ANDA to manufacture a generic version of Wyeth's EFFEXOR XR® (venlafaxine hydrochloride, extended release, used to treat depression).  View the complaint here.


    AstraZeneca AB et al. v. Dr. Reddy's Laboratories Inc. et al.

    3:11-cv-02317; filed April 21, 2011 in the District Court of New Jersey

    • Plaintiffs:  AstraZeneca AB; AstraZeneca LP; Pozen Inc.
    • Defendants:  Dr. Reddy's Laboratories Inc.; Dr. Reddy's Laboratories Ltd.

    Infringement of U.S. Patent No. 6,926,907 ("Pharmaceutical Compositions for the Coordinated Delivery of NSAIDs," issued August 9, 2005) following a Paragraph IV certification as part of Dr. Reddy's filing of an ANDA to manufacture a generic version of AstraZeneca's Vimovo® (naproxen and esomeprazole magnesium, used to relieve the signs and symptoms of osteoarthritis, rheumatoid arthritis, and ankylosing spondylitis, and to decrease the risk of stomach (gastric) ulcers in patients at risk of developing stomach ulcers from treatment with non-steroidal anti-inflammatory drugs (NSAIDs).  View the complaint here.


    Novartis AG et al. v. Teva Pharmaceuticals USA, Inc. et al.

    2:11-cv-02289; filed April 21, 2011 in the District Court of New Jersey

    • Plaintiffs:  Novartis AG; Novartis Pharmaceuticals Corp.
    • Defendants:  Teva Pharmaceuticals USA, Inc.; Teva Pharmaceutical Industries, Ltd.

    Infringement of U.S. Patent Nos. 6,025,391 ("Enteric-Coated Pharmaceutical Compositions of Mycophenolate," issued February 15, 2000), 6,172,107 ("Entric-Coated Pharmaceutical Compositions," issued January 9, 2001), and 6,306,900 (same title, issued October 23, 2001) following a Paragraph IV certification as part of Teva's filing of an ANDA to manufacture a generic version of Novartis' Myfortic7 (mycophenolate sodium, used for the prophylaxis or prevention of organ rejection in patients receiving allogeneic renal transplants, administered in combination with cyclosporine and corticosteroids).  View the complaint here.

  • Calendar

    May 18-19, 2011 – Forum on Freedom to Operate (C5) – Munich, Germany

    May 23-25, 2011 – 6th International Judges Conference on Intellectual Property Law (Intellectual Property Owners Association) – Brussels, Belgium

    June 2, 2011 – EPO Opposition & Appeals — The Case Law (Patent Resources Group and Management Forum) – Chicago, IL

    June 3, 2011 – European Patents — The Case Law (Patent Resources Group and Management Forum) – Chicago, IL

    June 7-8, 2011 – Biosimilars*** (American Conference Institute) – New York, NY

    June 8, 2011 – Biotechnology/Chemical/Pharmaceutical (BCP) Customer Partnership Meeting (U.S. Patent and Trademark Office) – 10:00 am – 4:00 pm (EDT)

    June 21-22, 2011 – 10th Annual Forum on Pharma Patent Lifecycles (C5) – London, England

    July 18-19, 2011 – Hatch-Waxman Boot Camp*** (American Conference Institute) – San Diego, CA

    September 18-20, 2011 – Accelerating Intellectual Property and Innovation in South Africa (South African Department of Science and Technology) – Cape Town, South Africa

    ***Patent Docs is a media partner of this conference or CLE

  • San Diego American Conference Institute (ACI) will be holding the next session of its Hatch-Waxman Boot Camp conference on July 18-19, 2011 in San Diego, CA.  ACI faculty will help attendees:

    • Understand the interplay of the PTO and FDA in the patenting of drugs and biologics;
    • Learn about the essentials of the FDA approval process and its link to biopharmaceutical patents;
    • Develop an in-depth and practical knowledge of Hatch-Waxman protocols, including: Orange Book listings, bioequivalency, exclusivities, the 30-month stay, and the safe harbor;
    • Navigate the intricacies of patent term adjustment and patent term extension;
    • Comprehend how recent biosimilars legislation is changing industry dynamics; and
    • Recognize how pre-commercialization concerns relative to CMS approval and Medicare/Medicaid formulary selection are influencing the patenting and approval of drugs and biological products.

    Brochure In particular, ACI's faculty will offer presentations on the following topics:

    • Key agencies overview:  Understanding the jurisdiction and interplay of the FDA and PTO in the patenting of drugs and biologics;
    • Identifying and comprehending pre-commercialization concerns relative to small molecules and biologics;
    • Exploring the link between the FDA approval process and the patenting of drugs and biologics;
    • IP overview for drugs and biologics:  Hatch-Waxman, BPCIA, trade dress, and more;
    • Paragraph IV disputes and litigation;
    • How the dynamics of biosimilars are changing the Hatch-Waxman landscape;
    • Orange Book listings, de–listings, and related challenges;
    • Bioequivalence and the "same active ingredient" vis-a-vis patentability;
    • An in-depth look at 180-day exclusivity;
    • Comprehending the intricacies of non-patent/regulatory exclusivity;
    • Assessing patent protections afforded under the safe harbor; and
    • Examining pharmaceutical patent extensions:  Patent Term Adjustment and Patent Term Extension.

    A post-conference workshop, entitled "Biosimilars: The Law, Interpreting Regulations, and Anticipated Litigation," will be offered from 9:00 am to 12:00 pm on July 20, 2011.  The workshop will provide an overview of the law governing biosimilars, delve into the preparations that should be made in anticipation of litigation, and run through what the first biosimilars litigation could possibly look like.

    The agenda for the Hatch-Waxman Boot Camp conference can be found here.  A complete brochure for this conference, including an agenda, detailed descriptions of conference sessions, list of speakers, and registration form can be obtained here.

    ACI - American Conference Institute The registration fee for the conference is $2,295 (conference alone) or $2,895 (conference and workshop).  Those registering by May 20, 2011 will receive a $300 discount and those registering by June 17, 2011 will receive a $200 discount.  Those interested in registering for the conference can do so here, by calling 1-888-224-2480, or by faxing a registration form to 1-877-927-1563.
    Patent Docs is a media partner of ACI's Hatch-Waxman Boot Camp conference.

  • By Donald Zuhn —

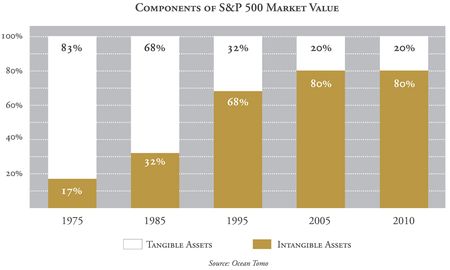
    Ocean Tomo A study released earlier this year by Ocean Tomo highlights the emergence of intellectual capital and how the U.S. has transitioned to an innovation-based economy over the past 35 years.  The company's Intangible Asset Market Value study indicated that in 2010, 80% of company value (for the S&P 500) was comprised of intangible assets, and that physical and financial accountable assets constituted only 20% of company value.  The study also showed that a significant portion of the intangible value was represented by patented technology.  The relative value of intangible and tangible assets has completely flipped since 1975, when 83% of corporate value was due to tangible assets and only 17% was derived from intangible assets.

    OceanTomoChart In a separate report — the Ocean Tomo Patent Quality Inventor study — the IP financial products company focused on the importance of inventors on corporate value by identifying the top inventors within four technology sectors:  biotech, industrials, semiconductors, and wireless.  Inventor ranking was based on the average patent IPQ® score for each inventor's 2010 issued and active U.S. patents within noted U.S. Patent Office Classifications.  In the biotech sector, the report awarded the top spot to Dr. Travis Mickle, who has received 16 patents falling within the 435 (Chemistry: Molecular Biology and Microbiology) and 436 (Chemistry: Analytical and Immunological Testing) classes that have an average IPQ score of 185.1 (where the median score is 100).  The report noted that this average placed Dr. Mickle's patents, which are directed to abuse-resistant amphetamine prodrugs, in the top 1% of all active U.S. utility patents.  The 16 patents are:  7,723,305; 7,718,619; 7,713,936; 7,700,561; 7,687,467; 7,687,466; 7,678,771; 7,678,770; 7,674,774; 7,671,031; 7,671,030; 7,662,788; 7,662,787; 7,659,254; 7,659,253; 7,655,630.

  •     By Kevin E. Noonan —

    European Court of Justice The European Court of Justice (ECJ) is poised to make a decision regarding the patent-eligibility of human embryonic stem cells (hESCs) in Europe, and that decision will be "no" if the court follows the recommendation of the court's advocate general.

    The case began in 2004, when Greenpeace sued in German federal court over a German patent to the University of Bonn involving methods for deriving neural cells from hESCs (DE 197568664 C1).  While German laws regarding stem cell research have been characterized as the "most restrictive in Europe," such research is permitted provided that it is performed with pluripotent (rather than totipotent) cells, using cell lines imported from abroad and only cell lines that were made prior to May, 2007.  Nevertheless, Greenpeace argued that claims to methods for using hESCs were "immoral and against public order," provisions of European law generally that define subject matter not eligible for patent (there is no corresponding provision under U.S. law).

    The German federal court ruled for Greenpeace in 2006, and the university appealed to the German supreme court.  That court decided that it needed to refer the question to the ECJ, since German law was closely patterned on European Union guidelines for biotechnology patenting.

    On March 17, 2011, the ECJ advocate-general, Judge Yves Bot, rendered an opinion that stem cell patents were "contrary to ethics and public policy" because they required "industrial use" of human embryos.  This is not a final decision of the ECJ, which will now consider the matter before the entire 13 judges of the court and render an opinion in the next several months.  However, it is expected that the court will agree with the advocate-general, since it is rare that such preliminary opinions are overruled, according to a court spokesman.

    Cover_nature This possibility prompted several stem cell scientists in Europe to send a letter to the court, published in Nature on April 28th, setting forth the case for stem cell patenting.  In the letter, the scientists expressed their "profound concern" in the capacities as "coordinators of multinational European stem-cell projects."  They contend that stem cells are cell lines, not embryos, and that they were derived from "surplus in vitro fertilized eggs donated after fertility treatments" that could not be maintained "indefinitely."  The existence of "more than 100" established stem cell likes make concerns about embryos "misplaced," they assert, and warn that it may be "premature to suggest that human embryonic stem cells can be replaced" (by iPS cells, for example) in developing stem cell therapies.  A stem cell ban in Europe will prevent scientists from "deliver[ing] clinical benefits without the involvement of biological industry," and such companies "must have patent protection as an incentive" to do their work in Europe.  This will result, these scientists predict, in "European discoveries [being] translated into applications elsewhere, at a potential cost to the European citizen."

    This action is the latest in a series of initiatives by individuals and groups with philosophical objections to patents involving biological materials, particularly material derived from human beings.  These include the Public Patent (PubPat) Foundation's attempts to invalidate the Thomson hESC patents in the U.S. (unsuccessful with regard to U.S. Patent Nos. 5,843,780 and 6,200,806 and currently pending against U.S. Patent No. 7,029,913) as well as PubPat and the ACLU's challenge to human gene patenting in the AMP v. USPTO (Myriad) case.  Opponents of such patents voice various arguments and rationales against patenting products of human biology, but ultimately the argument is the one raised in Europe, that there are moral objections to the practice.  Such objections are sincerely held by many and it is neither possible nor productive to make arguments on these grounds:  as it is in many public policy debates, these positions tend to be absolutes for some, not subject to merely reasoned argument.  Perhaps the best rejoinder of the argument is to posit the situation if patent opponents prevail (which also comprises a part of the scientists' argument in their letter).  Lack of patent protection will have two concrete and predictable consequences:  lack of financial investment to translate basic scientific discoveries to useful commercial products, and (where possible) reducing or eliminating public disclosure of inventions (and where not possible, investment in other technologies).  Neither outcome is conducive to reducing human morbidity or mortality or improving the human condition.  It is hard to understand how advocating such an outcome can be considered the more moral position.

    The Nature letter was signed by the following scientists:

    • Austin Smith EuroSyStem Project, Wellcome Trust Centre for Stem Cell Research, Cambridge, UK
    • Peter Andrews ESTOOLS Project, Centre for Stem Cell Biology, University of Sheffield, UK
    • Clare Blackburn EuroStemCell Project, MRC Centre for Regenerative Medicine, University of Edinburgh, UK
    • Anders Björklund NeuroStemcell Project, Wallenberg Neuroscience Center, Lund University, Sweden
    • Elena Cattaneo NeuroStemcell Project, University of Milan, Italy
    • Giulio Cossu OptiStem Project, Stem Cell Research Institute, San Rafaelle Institute of Milan, University of Milan, Italy
    • Juergen Hescheler ESNATS Project, Institute of Neurophysiology, University Hospital of Cologne, Germany
    • Olle Lindvall NeuroStemcell Project, Lund Stem Cell Center, Lund University, Sweden
    • Marc Peschanski STEM-HD, Institute for Stem Cell Therapy and Exploration of Monogenic Diseases, INSERM, Evry, France
    • Daniel Pipeleers BetaCellTherapy Project, Diabetes Research Centre, Free University of Brussels, Belgium
    • David Sassoon EndoStem Project, University of Paris VI/INSERM, Pitié Salpêtrière Medical School, Paris, France
    • Anna Veiga hESCreg Project, Centre of Regenerative Medicine, Barcelona, Spain
    • Catherine Verfaille HemiBio Project, Stem Cell Institute, K. U. Leuven, Belgium