• LESThe Greater Washington, DC Chapter of the Licensing
    Executives Society (LES) will be offering a program entitled "Patents and
    the Public Good" on November 14, 2012 from 5:30 – 8:00 pm (Eastern) at
    Sidley Austin LLP in Washington, DC. 
    Chief Judge Randall Rader of the U.S. Court of Appeals for the Federal
    Circuit, former Chief Judge Paul Michel of the U.S. Court of Appeals for the
    Federal Circuit, and Raymond Van Dyke of Van Dyke Law will review the positives
    of a strong patent system to our nation and the world.  A networking reception will be held from 5:30
    – 6:30 pm, and the presentation will be held from 6:30 – 8:00 pm.

    The
    registration fee for the program is $30 (LES members) or $40
    (non-members).  Those interested in
    registering for the program, can do so here.  The online registration deadline is November
    13, 2012.

  • Court Overrules PTO Interpretation of B-Delay Calculation

    By
    Kevin E. Noonan —

    District Court for the Eastern District of VirginiaJudge
    T.S. Ellis III of the U.S. District Court for the Eastern
    District of Virginia wrote the latest chapter in the story of judicial review of (and,
    generally, disagreement with) how the Office has interpreted the patent term
    adjustment provisions of the 1999 America Inventors Protection Act (AIPA).  This case involves so-called "B delay,"
    corresponding to a date that is three years after an application's filing date
    and the date on which the Office actually grants a patent pursuant to 35 U.S.C.
    § 154(b)(1)(B).  The issue
    was whether the Office had correctly implemented the statute by negating B delay
    in circumstances where an applicant had responded to a final rejection by
    filing a request for continued examination (RCE) (also a provision newly added
    to U.S. patent law by the AIPA).

    ExelixisThe
    case involved an application filed by Exelixis that resulted in grant of U.S.
    Patent No. 7,989,622 entitled "Phosphatidylinositol 3-Kinase Inhibitors
    and Methods of their Use."  The
    application was filed on January 15, 2008 and a first Office Action promulgated
    by the Office on February 22, 2010, resulting in an "A delay" (the
    time from the filing date to the first substantive action in a case pursuant to
    35 U.S.C. § 154(b)(1)(A)) of 283
    days.  The PTO issued Final Rejection on
    March 9, 2011 (38 months, i.e., 2 months from the three-year date calculated
    from the filing date), to which the applicants responded on April 11, 2011;
    this response was filed with an RCE.  "Less
    than three weeks later" (a time the Court characterized as "commendable,
    if, with respect to this application, uncharacteristic alacrity"), the
    Office issued a Notice of Allowance.  Despite payment of the Issue Fee almost immediately (on April 28, 2011),
    the patent did not grant until August 2, 2011.  At that time, the Office notified Exelixis that the PTA for this patent
    was a total of 368 days, corresponding to 283 days of A delay, 85 days of B
    delay, 0 days of C delay under 35 U.S.C. § 154(b)(1)(C) and
    subtracting 61 days for applicant delay in prosecution ("C reduction"
    under 35 U.S.C. § 154(b)(2)(C)).  The Office's calculation of the B delay, the
    issue in the case, credited Exelixis for 85 days from the time between the date
    that was three years after the application was filed and the date the Final
    Rejection was mailed; the Office did not give this patent the benefit of
    another 114 days corresponding from the date the RCE was filed until the grant date,
    implementing Office policy that filing an RCE stops the accumulation of B delay
    in the Office's PTA calculation.  These
    114 days and the Office's policies that deprived the '622 patent of this
    additional PTA prompted the lawsuit.  The Court set forth a helpful timeline:

    Timeline
    The Court recognized the context of the dispute being based in an assessment of
    Congressional intent in implementing the provisions of the General Agreement on
    Tariffs and Trade (GATT), which changed the term of U.S. patents from 17 years
    from issue to 20 years from the filing (or earliest priority) date, and the
    provisions of the AIPA apparently aimed at restoring term lost as the result of
    Patent Office delays in prosecution.  "Taken
    as a whole," the Court said:

    [T]he clear goal and purpose of these
    provisions is to provide a successful applicant with a patent that can be
    enforced against putative infringers for approximately 17 years — 20 years from
    the date of application less three years for prosecution and examination — and
    to reach this goal by providing applicants with day for day patent term
    extension for delays attributable to the PTO and day for day reductions for the
    patent term extension for delayed attributable to an applicant's failure to act
    with alacrity in certain circumstances.

    The Court characterizes the provisions of 35 U.S.C. § 154(b)
    as "guarantees" according to the titles of the three sections of § 154(b)
    relating to PTO delay, which is significant because the Court notes (in a
    footnote) that it is permissible for courts to use titles and headings in
    statutes in interpreting the meaning of the substantive provisions of law.

    On
    summary judgment, the Court ruled against the PTO's interpretation of the
    statute based on its construction of the statutory provisions and Congressional
    intent discerned inter alia from how Congress described and characterized those
    provisions in the statute.  The Court
    applied the deferential standard that agency action must be upheld unless it is
    "arbitrary, capricious, an abuse of discretion or otherwise not in
    accordance with law" under the Administrative
    Procedures Act (5 U.S.C. § 706(2)(A)), noting that an
    erroneous interpretation of the law satisfies this standard, citing Star Fruits S.N.C. v. U.S., 393 F.3d
    1277, 1281 (Fed. Cir. 2005), and Arnold
    Partnership v. Dudas
    , 362 F.3d 1338, 1340 (Fed. Cir. 2004).

    The Court's opinion begins with the "plain language of the statute," and
    the Court found that the meaning of the statute was "clear, unambiguous,
    and in accord with both the statute's structure and purpose."  Section 154(b)(2) clearly and "simply"
    is intended by its terms to provide a guarantee of no more than a three-year
    application pendency, according to the Court's opinion, and this goal is
    accomplished by "(i) starting a three year clock on the date the
    application is filed, (ii) tolling the running of this clock if, within the
    three year period, any of three events occur, including an RCE filing, and
    (iii) adding a day for day PTA to the patent term for any delay in the issuance
    of a patent after the three year clock, less any tolling, runs out."

    In
    the Court's view, the provisions of the statute involving RCEs have to do
    solely with tolling the clock with regard to the three-year date.  The opinion
    states that the portion of the statute concerning RCEs "clearly and
    unambiguously modifies and pertains to the three year period and does not apply
    to, or refer to, the day to day PTA remedy."  The Court characterizes as "[e]specially
    notable" that the statute "does not address the filing of an RCE after
    the expiration of the three year clock" (emphasis in original).  The statute "makes [it] clear that once
    the three year clock has run, PTA is to be awarded on a day for day basis
    regardless of subsequent events" in the Court's view.  The Court also notes that the filing of an
    RCE is not contained in the section setting forth the day to day reduction in
    PTA occasioned by applicant delay in prosecution, and concludes that the
    statute does not consider the filing of an RCE to be an action that warrants
    reduction in PTA under the statute.  The Court concludes that "RCE's gave no impact on the PTA after the three year
    deadline has passed."

    The Court rejected PTO arguments in favor of the Office's interpretation.  Specifically, the opinion swiftly dismissed
    the argument that the statute must be interpreted as if it included the word "then"
    in such a way that would support its interpretation by simply noting that the
    word "then" did not appear in the statute and on the grounds that the
    Office's interpretation would effectively treat the filing of an RCE as a failure
    to "engage in reasonable efforts to conclude prosecution of the
    application," i.e., the grounds for applicant delay where the Court had
    expressly found RCE filings were not included.  If this is the interpretation the Office wants to apply to RCEs, the Court tartly recommends that the Office should be sure to "issue any
    notice of rejection prior to the expiration of the three year period" as
    well as require prompt filing of an RCE.  The Court also rejects the Office's position that it should be accorded
    deference, saying that "deference is unwarranted when [] the statute is
    unambiguous."  In this regard the Court notes that the Federal Circuit had rejected this level of deference in
    the last case decided on PTA issues, Wyeth
    v. Kappos
    .  Finally, the Court noted
    that any disparate treatment that might result one application to the next
    (depending on whether an RCE was filed before or after the 3-year anniversary
    date) was something within the Office's power to control or minimize.

    It
    is likely that the government will appeal to the Federal Circuit, and thus that
    the finality of this interpretation of the statute will remain delayed until
    that review of complete.  This means that
    there is time for prudent patent practitioners to review the PTA status of their
    clients' patents to determine where there are instances where the PTA was "improperly"
    calculated under the interpretation set forth in this opinion.  But time may be of the essence for some because
    the statute also imposes a 180-day statute of limitations on filing challenges
    to Office PTA determinations (see "USPTO Posts Notice Regarding Wyeth Decision"), and while an
    intervening change in the law may permit challenges outside that period it
    makes better sense to identify important patents and file a lawsuit based on
    this interpretation of the B delay statutory provisions to best protect both
    clients and practitioners.

    Exelixis, Inc. v. Kappos (E.D. Va. 2012)
    Memorandum Opinion by Judge T.S. Ellis, III

  • By Donald Zuhn —

    California FlagOn Tuesday, while California
    voters were helping President Obama secure a second term in the White House, picking the President over Republican challenger Mitt Romney by a 59.1% to 38.6%
    margin, they were also responsible for defeating a ballot initiative, Proposition 37, that would have required raw or processed food to be
    labeled if such food was made from plants or animals having genetic material
    changed in specified ways.  The
    initiative measure, entitled "The California Right to Know Genetically
    Engineered Food Act," would have amended the California Health and Safety
    Code such that "any food offered for retail sale in California [would be considered to be] misbranded if it is or may
    have been entirely or partially produced with genetic engineering and that fact
    is not disclosed."  The margin of
    defeat for Proposition 37 was somewhat narrower than that in the Presidential
    race, with voters rejecting the ballot initiative 53.7% to 46.3%.

    According to Proposition
    37, "genetically engineered" food was defined as:

    [A]ny food that is produced from an organism or
    organisms in which the genetic material has been changed through the
    application of:
        (A)
    In vitro nucleic acid techniques, including recombinant deoxyribonucleic acid
    (DNA) techniques and the direct injection of nucleic acid into cells or
    organelles, or
        (B)
    Fusion of cells, including protoplast fusion, or hybridization techniques that
    overcome natural physiological, reproductive, or recombination barriers, where
    the donor cells/protoplasts do not fall within the same taxonomic family, in a
    way that does not occur by natural multiplication or natural recombination.

    Right to KnowUnder the Act, raw
    agricultural food that was genetically engineered would have to be labeled as
    "Genetically Engineered," and processed food ("any food produced from a raw agricultural
    commodity that has been subject to processing such as canning, smoking,
    pressing, cooking, freezing, dehydration, fermentation, or milling") would have to be
    labeled as "Partially Produced with Genetic Engineering."

    At
    the California Secretary of State website,
    arguments in favor of and against the ballot initiative were presented.  In addition, voters were directed by the
    website to the groups California Right to Know
    (in favor of the initiative) and NO on 37 (against
    the initiative) for additional information.

    No on 37Interestingly,
    the Ballotpedia webpage for Proposition 37
    listed the California Democratic Party and the Green Party of California as
    supporters of the failed ballot initiative and the California Republican Party as
    being opposed to the initiative.  Donors
    to the "No on 37" campaign included Monsanto, E.I. Dupont De Nemours
    & Co., DOW Agrisciences, Bayer Cropscience, BASF Plant Science, Syngenta
    Corporation, and the Biotechnology Industry Organization (BIO).

    According
    to a report
    in the San Jose Mercury News, proponents of the initiative, including
    California Right to Know, will now turn their efforts towards getting similar initiatives
    on the ballot in Washington and Oregon.

  •     By Andrew Williams —

    USPTO SealLast month, on October 18,
    2012, the U.S. Patent and Trademark Office published a notice of
    proposed rulemaking (77 Fed. Reg. 64190) to finally update its rules of professional conduct by conforming
    them to the ABA Model Rule of Professional Conduct.  The ABA Model Rules have been adopted in some
    form by every state (except California) and the District of Columbia.  These USPTO proposed rules are welcome news
    for patent practitioners that are also attorneys, because currently, such
    individuals are required to know and abide by the ethics rules of the state in
    which they practice, as well as the Patent and Trademark Office Code of
    Professional Conduct, which is based on the former ABA Model Code from
    1980.  Of course, there are differences
    between the ABA Model Rules and the proposed USPTO Rules, just are there are
    differences between the current USPTO Code and the proposed Rules of
    professional.  In highlighting any
    differences, this post will refer to the ABA Model Rules and not to the
    specific rules of any particular jurisdiction, because to do so would be too
    onerous (in other words, if you are an attorney licensed in a U.S.
    jurisdiction, do not rely on any statements found below regarding what ethical
    rules are required in your jurisdiction, but instead please consult the
    particular rules of the state in which you practice).  If you would like to provide any comments to
    the Patent Office regarding these proposed rules, they must be submitted to the Office on or before December 17, 2012.

    The Patent Office detailed its
    reasoning for proposing these rules at this time in the Federal Register
    notice, and these reasons were reiterated by David Kappos on his Public Blog.  The Office recognized that adopting
    harmonized rules of ethical and professional conduct is consistent with the
    mandate of the Leahy-Smith America Invents Act of 2011.  There are almost 42,000 registered patent
    practitioners, and at least 75% of them are attorneys.  Without the new rules, these patent attorneys
    are required to go back and forth between the old Model Code and the newer
    Model Rules.  More importantly, the
    amount of authority referring to or interpreting the Model Code is, at best,
    limited.  Instead, a practitioner has the
    benefit of comments, annotations, and case law interpreting the Model Rules
    which, even if not binding on the Patent Office, will be useful until a larger
    body of USTPO-specific precedent is established.  Finally, the Office pointed out that the new
    rules do not deviate significantly from the current rules of professional
    conduct, and largely codify obligations or professional and fiduciary duties
    that already exist.

    The differences between the
    current PTO Code of Professional Conduct, the ABA Model Rules of Professional
    Conduct, and the proposed USPTO Rules are highlighted in the Federal Register
    notice, as well as on the Patent Office website.  One of the more significant changes from the
    current Patent Office rules is the removal of the ability of the Patent Office
    to collect an annual practitioner maintenance fee, and the removal of the
    ability of the Patent Office to require continuing legal education reporting
    of all registered agents or attorneys.  Of course, these provisions were never enforced, but the new rules
    remove any threat of potential enforcement in the future.  There are also examples of differences
    between the ABA Model Rules of Professional Conduct and the proposed USPTO
    Rules that include the inclusion of ex
    parte
    proceedings in the section that requires candor towards the tribunal
    (proposed § 11.303) and the removal of the sections in the ABA Model Rules that
    deal exclusively with criminal proceedings.  However, probably the most important section to which patent attorneys need
    to pay attention is proposed § 11.106, dealing with the confidentiality of
    information.

    Confidentiality of Information

    One of the most significant
    changes found in the proposed USPTO Rules in terms of what is currently required
    by the Patent Office, and in terms of how it differs from the ABA Model Rules,
    is that relating to the handling of a client's confidential information.  This new rule differs from that currently in
    place by changing what is defined as confidential information.  The current USPTO Code defines "Confidence"
    as "information protected by the attorney-client or agent privilege under
    applicable law," and "Secret" as "other information gained
    in the professional relationship that the client had requested by held in
    inviolate of the disclosure of which would be embarrassing or would be likely
    to be detrimental to the client."  USPTO Code § 10.57(a).  As can be
    seen, and has been noted by commentators for years, this definition is somewhat
    limiting.  Most confidential information
    of a client does not fall under this definition of confidence, and information
    is only secret under this definition if the client has expressly stated that it
    is so (or if its disclosure would be embarrassing or detrimental).  Instead, the ABA Model Rules and the proposed
    USPTO Rules expand the scope of this term, defining confidential information as
    any "information relating to the representation of a client . . . ."  See, e.g., Proposed USPTO Rule §
    11.106(b).  Of course, this definition is
    provided in the context of what information a practitioner can or cannot
    (technically may or shall not) reveal, and it is important to note that this
    information can include information that is publically available.  In fact, the ABA rules have been interpreted
    to mean that it is possible for an attorney to be prohibited from revealing
    information related to his or her representation of a client even though such
    information could have otherwise been obtained from a publically available
    source.  As a result, it can be a
    difficult question figuring out whether something is truly "confidential"
    under this Rule.  Thus, this is a perfect
    example of a rule for which there is a lot of non-USPTO commentary, and
    hopefully the Patent Office will either provide its own guidance of the extent
    of "confidential" information, or will accepted this preexisting (and
    future) commentary as persuasive on the issue.

    The more troubling aspect
    of the proposed rule is where it deviates from the ABA Model Rules.  As with the Model rule, the proposed USPTO
    Rule explains what information a practitioner "shall not" reveal without
    informed consent, implied authorization, or permission under rules.  In addition, both sets of rules explain what
    information a practitioner "may" reveal, including such things as
    information that may prevent death, bodily harm, or fraud.  Of note, however, even in the cases where
    death, bodily harm, or fraud may be result, the practitioner is not required to
    reveal such information, but rather the practitioner has permission to do
    so.  The proposed USPTO Rules and the ABA
    Model Rules nevertheless diverge in one important respect:  the proposed USPTO
    Rules include a type of information that is mandatory to reveal:  "A
    practitioner shall disclose to the Office information necessary to comply with
    applicable duty of disclosure provisions." 
    In other words, under this proposed rule, a finding that relevant
    information was intentionally withheld by a practitioner involved in the
    prosecution of an application will not only cause a patent to become
    unenforceable, it will result in an ethical violation by the practitioner.

    This requirement to
    disclose information even though it may be confidential also has the
    possibility to trap a patent practitioner between two ethical obligations.  Importantly, this obligation to disclose
    information necessary to comply with the duty of disclosure is not limited to
    confidential information from that particular prosecution client.  Instead, it is possible under the rule that a
    patent attorney could be aware of confidential information belonging to another
    client that is nevertheless relevant, and he would therefore be ethically
    required to submit it with the prosecution client's application.  The Patent Office apparently considered this
    potential ethical quandary, and as a result, removed this type of disclosure
    from the types of information that are prohibited from being revealed.  Of course, try explaining to your other
    client why you revealed its confidential information and made it publically
    accessible in another client's patent file. 
    Moreover, any patent attorney in a jurisdiction that has adopted some
    form of the ABA Model Rules is subject to rules of professional conduct that do
    not have such a permissive cut-out. 
    Therefore, such a patent attorney would be required by the proposed
    USPTO Rules to reveal such third-party confidential information to satisfy the
    duty of disclosure, but would be prohibited from doing so by the professional
    rules of conduct of their state.

    To be fair, a similar
    quandary probably already exists, because if a patent attorney is aware of such
    third-party confidential information, that attorney is already required to disclose it
    to prevent the patent from becoming unenforceable (and therefore withholding it
    would violate the duty to act competently for your client).  Nevertheless, the proposed rules would
    explicitly make such action unethical, and subject to potential disciplinary
    proceedings.  What is unclear from the
    proposed USPTO rules is whether an attorney in such an ethical quandary could
    noisily withdraw from the case, thereby removing the problem.  However, if not, such an attorney would face
    certain disciplinary proceedings (either in their state or at the Patent Office), because it is impossible to "not reveal" information and, at
    the same time, submit it to the Patent Office in an information disclose
    statement.  Yet, this is what the
    proposed rule, in conjunction with the ABA Model Rules would require.

    Anyone wishing to provide comments regarding
    the new rules regarding confidential information, or provide comments to any
    other of the proposed rules, can do so by submitting them by e-mail to ethicsrules.comments@uspto.gov; by
    regular mail addressed to:  Mail Stop OED-Ethics Rules, United States
    Patent and Trademark Office, P.O. Box 1450, Alexandria, VA 22313–1450, marked
    to the attention of William R. Covey, Deputy General Counsel for Enrollment and
    Discipline and Director of the Office of Enrollment and Discipline; or via the
    Federal eRulemaking Portal.  Again, the deadline for submitting comments
    is December 17, 2012.  Additional information regarding the submission of
    comments can be found in the Office's Federal Register notice (77 Fed. Reg. 64190).

  • By Kevin E. Noonan —

    Mimicry
    is a classic (and classical) biological phenomenon, the appreciation of which
    dates to the time when biology was more accurately called "natural history"
    and was more diversion for English country gentlemen than proper science.  Perhaps
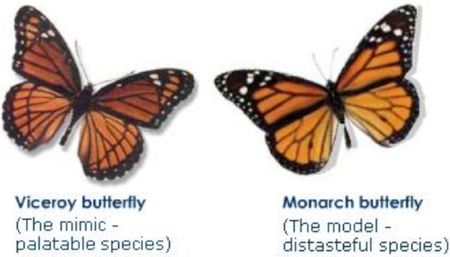
    the most well-known form of mimicry is Müllerian mimicry, and the
    best example of that is the viceroy butterfly (Limenitis
    archippus
    ),
    which although tasty to avian predators displays markings almost
    indistiguishable from the monarch butterfly (Danaus
    plexippus
    ),
    which is not only unpalatable but actually toxic to birds (see below).  Anyone with even a rudimentary appreciation
    of Linnean classification will recognize another interesting feature of
    mimicry:  despite their visual similarities, the monarch and viceroy butterflies
    are not only different species but members of different genera, suggesting that
    genetic inheritance cannot account for the convergence in wing pattern
    phenotype.

    Butterflies
    Heliconius melpomeneRecent
    work employing genomic sequencing and linkage analysis (from the Heliconius
    Genome Project) has begun to shed light on the genetic bases for Müllerian
    mimicry.  This work was performed on a
    South American butterfly species, Heliconius
    melpomene
    (at right).  As reported in Nature (doi:10.1038/nature11041), naturally occurring species of Heliconius show significant
    introgression in regions related to color pattern development.  Heliconius butterflies comprise 43 species and
    hundreds of races and make up rapidly radiating butterfly species in South
    America; certain of these species exhibit classic Müllerian mimicry, wherein palatable species adopt the
    color pattern of other species that are unpalatable to bird predators.  The
    authors sequenced almost 13,000 genes (12,669 open reading frames) in the 269
    megabase (Mb) butterfly genomic DNA and mapped the majority (~83%) of these
    genes to the 21 Heliconius chromosomes using a modern
    version of RFLP analysis termed restriction site DNA linkage mapping.  This work
    found that the chromosomal organization of Heliconius butterfly genomic DNA was "broadly
    conserved"
    since the evolutionary split from a common ancestor shared with Bombyx (silkmoth) species in the
    Cretaceous (~100 million years ago).  Comparison
    with the Bombyx mori (silkmoth) genome
    identified 6,010 orthologues and previously unidentified chromosomal fusions in
    the Heliconius chromosomal
    complement.  Biologically important expansion was found in families of
    genes related to chemosensation and body shape and form (homeobox, or Hox, genes).  The Heliconius genome showed an
    expected increase in ultraviolet opsins (due to the diurnal lifestyle adopted
    by butterflies) and an unexpected genetic complexity in olfactory genes (33 Heliconius
    olfaction genes, similar to what was found in monarch butterfly species having
    34 olfactory genes).  The researchers also
    found additional Hox genes related to body shape and form, similar to
    what is found in other butterflies and related dipteran species.

    The
    researchers compared genomic DNA from three co-mimicry species, Heliconius melpomene, Heliconius timareta, and Heliconius
    elevatus
    and produced a genetic tree between 84 individuals from H.
    melpomene
    and related species (see below).  In
    this comparison, the researchers found hybrid exchange especially at two genomic regions that control mimicry
    pattern based on ~4% (12Mb) of the
    genome.  The results obtained from Heliconius melpomene amaryllis
    and H. timareta ssp. were
    compared and the researchers report evidence of greater hybrid interbreeding
    between these species when compared with H.
    melpomene aglaope
    and H. timareta ssp, which do not
    have overlapping spatial ranges.  The
    researchers estimated 2-5% exchange of the genome; the introgression was not
    uniformly distributed over the 21 chromosomes but seems to be limited to 11
    chromosomes with two chromosomes showing the strongest frequency of
    introgression.  These regions, comprising loci known to be associated with red pattern (B/D) and yellow (N/Yb) pattern loci, showed blocks of sequence (~100 kb in size) that are
    shared between species and exchanged in hybrids.  The genetic makeup of these shared blocks of
    sequence were more consistent with hybrid introgression (and fixation) than in
    other alternatives, such as convergent evolution.

    Genetic Tree
    The researchers concluded that these species
    exchange protective color pattern genes promiscuously and that their evidence
    of hybrid exchange and fixation indicates that this mode of gene transfer
    between diverse species may be a more important mechanism for explaining
    phenomena like mimicry than had been previously appreciated.  The authors propose that hybridization
    between mimicking species is a plausible explanation for "parallel
    evolution of multiple mimetic patterns" in these species.

    Supplemental
    experimental details can be found here.

    Image of Heliconius melpomene by Greg Hume, from the Wikipedia Commons under the Creative Commons Attribution-ShareAlike 3.0 Unported license.

  • By Sherri Oslick —

    Gavel About Court Report:  Each week we will report briefly on recently filed biotech and pharma cases.

    Acura Pharmaceuticals Inc. v.
    Impax Laboratories Inc.

    1:12-cv-01371; filed October
    31, 2012 in the District Court of Delaware

    Acura Pharmaceuticals Inc. v.
    Par Pharmaceutical Inc.

    1:12-cv-01372; filed October
    31, 2012 in the District Court of Delaware

    Acura Pharmaceuticals Inc. v.
    Sandoz Inc.

    1:12-cv-01373; filed October
    31, 2012 in the District Court of Delaware

    Acura Pharmaceuticals Inc. v.
    Watson Laboratories Inc.-Florida et al.

    1:12-cv-01374; filed October
    31, 2012 in the District Court of Delaware

    • Plaintiff:  Acura
    Pharmaceuticals Inc.
    • Defendants:  Watson
    Laboratories Inc. – Florida; Watson Pharmaceuticals Inc.; Watson Pharma
    Inc.

    The complaints in these cases
    are substantially identical.  Infringement of U.S. Patent No. 7,510,726 ("Methods and
    Compositions for Deterring Abuse of Opioid Containing Dosage Forms,"  issued March 31, 2009) following a Paragraph
    IV certification as part of defendants' filing of an ANDA to manufacture a
    generic version of Oxecta® (oxycodone hydrochloride, used for the management of
    acute and chronic moderate to severe pain where the use of an opioid analgesic
    is appropriate, marketed by Pfizer). 
    View the Impax complaint here.


    Esoterix Genetic Laboratories,
    LLC et al. v. Life Technologies Corp., et al.

    1:12-cv-01173; filed October
    31, 2012 in the Middle District of North Carolina

    • Plaintiffs:  Esoterix Genetic
    Laboratories, LLC; The Johns Hopkins University
    • Defendants:   Life Technologies Corp.; Applied
    Biosystems, LLC; Ion Torrent Systems, Inc.

    Infringement of U.S. Patent
    Nos. 6,440,706 (" Digital Amplification,"  issued August 27, 2002), 7,824,889 (same
    title, issued November 2, 2010), and 7,915,015 (same title, issued March 29,
    2011) based on defendants' manufacture and sale of products, kits, and devices
    used in methods for the detection of ratios of genetic sequences in a
    biological sample and/or for the detection of allelic imbalances in a biological
    sample.  View the complaint here.


    Jazz Pharmaceuticals, Inc. v. Roxane
    Laboratories, Inc.

    2:12-cv-06761; filed October
    26, 2012 in the District Court of New Jersey

    Infringement of U.S. Patent
    No. 8,263,650 (" Microbiologically
    Sound and Stable Solutions of Gamma-Hydroxybutyrate Salt for the Treatment of
    Narcolepsy,"  issued September 11, 2012) following a
    Paragraph IV certification as part of Roxane's filing of an ANDA to manufacture
    a generic version of Jazz's Xyrem® (sodium oxybate, used to treat
    narcolepsy).  View the complaint here.

  • Calendar

    November 12-14, 2012 – Fall Intellectual Property
    Counsels Committee (IPCC) Conference
    (Biotechnology
    Industry Organization) – Charleston, SC

    November 15, 2012 – Successful Strategies for Overcoming Obviousness Rejections in the Patent
    Application Process
    (McDonnell
    Boehnen Hulbert & Berghoff LLP) – 10:00 – 11:15 am
    (CT)

    November 27, 2012 – Patents and the Written Description Requirement: Navigating
    Section 112 Disclosure Obligations and Withstanding Invalidity Challenges
    (Strafford) – 1:00 – 2:30 pm (EST)

    November 28-29, 2012 – Biotech Patents*** (American Conference
    Institute) – Boston, MA

    November 28-29, 2012 – Orphan Drugs and Rare Diseases*** (American Conference
    Institute) – Boston, MA

    December 3-5, 2012 – Drug and Medical Device
    Litigation
    *** (American Conference
    Institute) – New York, NY

    December 4-5, 2012 – Paragraph IV Disputes*** (American Conference
    Institute) – San Francisco, CA

    January 23-24, 2013 – The Comprehensive Guide to Patent Reform*** (American Conference
    Institute) – New York, NY

    ***Patent Docs is a media partner of this conference or CLE

  • New York #3American Conference
    Institute (ACI) will be offering "The Comprehensive Guide to Patent Reform"
    from January 23-24, 2013 in New York, NY. 
    The conference will allow attendees to:

    • Adapting to the
    first-to file regime and developing prosecution techniques to ensure timely
    filings that result in robust patents;
    • Dissecting the
    new rules covering the usage of declarations and assignments to avoid costly
    delays and confusion associated with recalcitrant or missing inventors;
    • Utilizing the inter partes review process to challenge
    troubling patents and understanding the discovery rules associated with this
    procedure;
    • Determining when
    it is appropriate to employ the post-grant review procedure to attack a rival
    patent and crafting defenses to it;
    • Grasping the
    advantages of supplemental examination, its potential to cure defects in
    prosecution, and recognizing its risks;
    • Maximizing the
    potential of third party prior art submission in attacking a rival patent
    pre-issuance and guarding against its effects with smart prosecution;
    • Understanding the
    implications of patent reform for Hatch-Waxman litigation; and
    • Demystifying the
    new standards for prior art encoded in the America Invents Act including
    discussion of the new "disclosures" definition, the impact on
    obviousness findings, and more.

    BrochureIn particular,
    ACI's faculty will offer presentations on the following topics:

    • USPTO keynote
    address: The USPTO's efforts to implement AIA provisions impacting patent
    prosecution — to be presented by Teresa Stanek Rea, Deputy Director, U.S.
    Patent and Trademark Office;
    • Preparing your
    company for the looming implementation of the first-to-file regime and
    preparing strategies to exploit the coming changes for maximum benefit;
    • Identifying what
    constitutes prior art under the AIA and planning a patenting strategy around
    the new standards;
    • Slipping the
    Gordian knot — How to avoid ethical conundrums presented by the AIA's
    inconsistent best mode requirements;
    • USPTO address:
    Detailing the new post grant opposition procedures, how they will impact
    patenting, and more — to be presented by Thomas Giannetti, Patent
    Administrative Judge, U.S. Patent and Trademark Office;
    • Learning the
    sweet science: Win the new boxing match known as inter partes review;
    • Preparing for the
    eventual employment of the post grant review procedure in opposition practice;
    • Deciding when to
    use supplemental examination to cure defects in a patent: Balancing the risks
    and rewards;
    • Gambit: Utilizing
    third party prior art submissions in the pre-issuance chess match;
    • Examining the
    ramifications of patent reform for Hatch-Waxman litigation and the
    brand/generic wars; and
    • Investigating the
    changes to declarations and assignments and understanding how these new provisions
    can be used to keep prosecution on track.

    In addition, a
    pre-conference workshop entitled "Patent Reform 101: A Primer on the
    Fundamental Provisions of the America Invents Act" will be offered from 9:00
    am to 12:00 pm on January 23, 2013, and a post-conference workshop entitled
    "Interactive Working Group Session: A Hypothetical Invention Being
    Patented Under the AIA" will be offered from 9:00 am to 12:00 pm on
    January 25, 2013.

    An agenda for the
    conference can be found here.  More information regarding the workshops can
    be found here.  A complete brochure for this conference,
    including an agenda, detailed descriptions of conference sessions, list of
    speakers, and registration form can be obtained here.

    ACI - American Conference InstituteThe registration
    fee for the conference is $2,295 (conference alone), $2,995 (conference and one
    workshop), or $3,495 (conference and two workshops).  Those registering by November 9, 2012 will
    receive a $300 discount and those registering by December 14, 2012 will receive
    a $200 discount.  Patent Docs readers who reference the discount code "PD 200" will receive $200 off the current price tier when registering.  Those interested in
    registering for the conference can do so here, by
    e-mailing CustomerService@AmericanConference.com, by calling 1-888-224-2480, or
    by faxing a registration form to 1-877-927-1563.

    Patent Docs is a media partner of the Patent Reform conference.

  • By Ann Palma —

    Gavel About
    Court Report Supplement:  Periodically, we will
    report on biotech and pharma cases that were inadvertently omitted from
    our Court Report column
    .

    W.H.P.M.,
    Inc. v. BiotechPharma, LLC et al.

    1:12-cv-137;
    filed February 10, 2012 in the Eastern District of Virginia

    • Plaintiff:  W.P.H.M., Inc.
    • Defendants:  BiotechPharma,
    LLC and Raouf A. Guirguis

    Infringement
    of U.S. Patent No. 7,927,562 ("Collection and Assay Device for Biological
    Fluid," issued April 19, 2011) based on BiotechPharma's manufacture, use,
    sale, and offers for sale of its products for drug testing and identity
    verification, including the "BTP tube device" or "DRG-SA06 Drug
    Testing & Identity Verification System."  View complaint here.


    Lonza
    Walkersville, Inc. v. Hyglos GmbH

    1:2012-cv-602;
    filed February 4, 2012 in the District of Maryland

    Infringement
    of U.S. Patent Nos. 5,716,834 ("Cloned Factor C cDNA of the Singapore
    Horseshoe Crab, Carcinoscorpius
    Rotundicauda
    and Purification of Factor C Proenzyme," issued February
    10, 1998), 5,712,144 (same title, issued on January 27, 1998), 5,858,706 ("Expression
    of Carcinoscorpius Rotundicauda Factor C in Eukaryotes," issued on January
    12, 1999), 5,985,590 (same title, issued on November 16, 1999), and 6,645,724 ("Assays
    for Endotoxin," issued on November 11, 2003) based on Hyglos GmbH's manufacture,
    use, offers for sale, sale, and importation of products that contain endotoxin
    detection assays, including EndoLISA®, or that contain an affinity matric for
    removal of bacterial endotoxins from solutions, including EndoTRAP®.  View the complaint here.


    ArcticDX,
    Inc. et al v. Sequenom, Inc. et al.

    2:12-cv-00081;
    filed February 29, 2012 in the Eastern District of Texas

    • Plaintiffs:  ArcticDx, Inc.;
    ArcticA, Inc.; ArticAx US Ltd.
    • Defendants:  Sequenom, Inc.;
    Sequenom Center for Molecular Medicine, LLC

    Infringement
    of U.S. Patent No. 8,114,592 ("Genetic Markers Associated with Age-Related
    Macular Degeneration, Methods of Detection and Uses Thereof," issued
    February 14, 2012) based on Sequenom's use, sale, and offer for sale of genetic
    diagnostic products, including RetnaGene®. 
    Declaratory Judgment of noninfringement and invalidity of U.S. Patent
    Nos. 8,053,190 ("Susceptibility Genes
    for Age-Related Maculopathy (ARM) on Chromosome 10q26," issued November 8,
    2011), 7,867,727 ("Methods and Reagents for Treatment and Diagnosis
    of Vascular Disorders and Age-Related Macular Degeneration," issued
    January 11, 2011), 7,695,909 ("Susceptibility
    Genes for Age-Related Maculopathy (ARM) on Chromosome 10q26," issued on
    April 13, 2010), 7,351,524 ("Diagnostics and Therapeutics for
    Macular Degeneration-Related Disorders," Issued on April 1, 2008) or
    8,088,579 ("Complement Factor H for Diagnosis of Age-Related Macular
    Degeneration," issues January 3, 2012) based on Arctic's manufacture, use,
    or sale of products for genetic diagnostics, including Macula Risk®.  View the complaint here.


    XY, LLC v.
    Trans Ova Genetics, LC

    5:12-cv-208;
    filed March 5, 2012 in the Southern District of Western District of Texas

    Infringement
    of U.S. Patent Nos. 7,820,425 ("Method of Cryopreserving Selected Sperm
    Cells," issued on October 26, 2010), 6,357,307 ("System and Method of
    Flow Cytometry and Sample Handling," issued on March 19, 2002), 6,604,435
    ("Low Acceleration Method of Flow Cytometry," issued on August 12,
    2003), 6,782,768 ("Flow Cytometer Nozzle," issued on August 31, 2004),
    6,263,745 ("Flow
    Cytometer Nozzle and Flow Cytometer Sample Handling Methods," issued on July 24,
    2001), 7,713,687 ("System to
    Separate Frozen-Thawed Spermatozoa Into X-Chromosome Bearing and Y-Chromosome
    Bearing Populations," issued on May 11, 2010), 7,771,921 ("Separation systems of Frozen-Thawed
    Spermatozoa into X-Chromosome Bearing and Y-Chromosome Bearing populations,"
    issued on August 10, 2010), 6,149,867 ("Sheath Fluids and Collection Systems for Sex-Specific
    Cytometer Sorting of Sperm," issued on November 21, 2000), 6,524,860 ("Methods for Improving Sheath Fluids
    and Collection Systems for Sex-Specific Cytometer Sorting of Sperm," issued on February
    25, 2003), and 7,195,920 ("Collection Systems for Cytometer Sorting of Sperm,"
    issued on March 27, 2007) based on Defendant's use of flow cytometers (or other
    similar apparatuses) for sex selection of sperm cells and the sale of semen
    produced using such apparatuses, in addition to Defendant's use of flow
    cytometry-based sex selection protocols and procedures.  View the complaint here.


    Proprietary
    Wellness, LLC v. Primordial Performance, LLC

    4:12-cv-11072;
    filed March 9, 2012 in the Eastern District of Michigan

    Infringement
    of U.S. Patent No. 8,084,446 ("Use of DHEA Derivatives for Enhancing
    Physical Performance," issued on December 27, 2011) based on Defendant's
    manufacture, use, offer for sale, sale, and importation of products or
    processes containing 4-DHEA or Andosterone and its esters, in addition to
    claims of breach of license agreements, unjust enrichment, quantum merit, and
    tortious interference. View the complaint here.


    Butamax™
    Advanced Biofuels LLC v. Gevo, Inc.

    12-cv-298;
    filed March 12, 2012 in the Eastern District of Delaware

    Infringement
    of U.S. Patent No. 8,129,162 ("Ketol-Acid Reductoisomerase Using NADH,"
    issued on March 6, 2012) based on Defendant's manufacture and use of recombinant
    microorganisms comprising mutant ketol-acid reductoisomerase enzymes.  View the complaint here.

  • By Kevin E. Noonan —

    Washington - Capitol #3Since
    1980, the Bayh-Dole Act has encouraged, facilitated, and required universities
    and other recipients of Federal grant money to license inventions resulting
    from funded research for commercialization.  While this has been one of the bedrocks upon which the biotechnology
    industry was built, and has enabled almost every research university to obtain
    patent protection on its technologies (instead of watching them be stolen by
    corporations foreign and domestic), it has never avoided opposition by "public
    interest" groups who believe that universities should not be engaged in
    profit-making ventures or that the U.S. citizenry should not have to pay
    businesses for inventions the public has already paid for.  Of course, this argument ignores the relative
    financial contributions of the parties:  while the Federally funded invention is
    the sine qua non of the commercial technology it is rarely enough, and
    (particularly in the pharmaceutical and biotechnology industries) the cost of
    research is vastly outweighed by the cost of development.

    However,
    these concerns, as well as the concern that a company could license university
    and other Federally funded inventions and then suppress them motivated (in
    part) a section of the law that gave the government "march-in" rights.  These rights, and the conditions triggering
    their exercise, are set forth in 35 U.S.C. § 203:

    35 USC § 203 – March-in rights

    (a) With respect to any subject invention in
    which a small business firm or nonprofit organization has acquired title under
    this chapter, the Federal agency under whose funding agreement the subject
    invention was made shall have the right, in accordance with such procedures as
    are provided in regulations promulgated hereunder to require the contractor, an
    assignee or exclusive licensee of a subject invention to grant a nonexclusive,
    partially exclusive, or exclusive license in any field of use to a responsible
    applicant or applicants, upon terms that are reasonable under the
    circumstances, and if the contractor, assignee, or exclusive licensee refuses
    such request, to grant such a license itself, if the Federal agency determines
    that such—

    (1) action is necessary
    because the contractor or assignee has not taken, or is not expected to take
    within a reasonable time, effective steps to achieve practical application of
    the subject invention in such field of use;
    (2) action is necessary to
    alleviate health or safety needs which are not reasonably satisfied by the
    contractor, assignee, or their licensees;
    (3) action is necessary to meet requirements
    for public use specified by Federal regulations and such requirements are not
    reasonably satisfied by the contractor, assignee, or licensees; or
    (4) action is necessary because the
    agreement required by section 204
    has not been obtained or waived or because a licensee of the exclusive right to
    use or sell any subject invention in the United States is in breach of its
    agreement obtained pursuant to section 204.

    (b) A determination
    pursuant to this section or section 202
    (b)(4) shall
    not be subject to chapter 71 of title 41.
    An administrative appeals procedure shall be established by regulations
    promulgated in accordance with section 206.
    Additionally, any contractor, inventor, assignee, or exclusive licensee
    adversely affected by a determination under this section may, at any time
    within sixty days after the determination is issued, file a petition in the
    United States Court of Federal Claims, which shall have jurisdiction to
    determine the appeal on the record and to affirm, reverse, remand or modify, as
    appropriate, the determination of the Federal agency. In cases described in
    paragraphs (1) and (3) of subsection (a), the agency's determination shall be
    held in abeyance pending the exhaustion of appeals or petitions filed under the
    preceding sentence.

    Regulations on how these
    rights can be petitioned for exercise by the relevant Federal agencies (such as
    the National Institutes of Health) have been promulgated:

    37 C.F.R. § 401.6 Exercise of march-in rights.

    (a) The following procedures shall govern
    the exercise of the march-in rights of the agencies set forth in 35 U.S.C. 203
    and paragraph (j) of the clause at § 401.14.

    (b) Whenever an agency receives information that it
    believes might warrant the exercise of march-in rights, before initiating any
    march-in proceeding, it shall notify the contractor in writing of the information
    and request informal written or oral comments from the contractor as well as
    information relevant to the matter. In the absence of any comments from the
    contractor within 30 days, the agency may, at its discretion, proceed with the
    procedures below. If a comment is received within 30 days, or later if the
    agency has not initiated the procedures below, then the agency shall, within 60
    days after it receives the comment, either initiate the procedures below or
    notify the contractor, in writing, that it will not pursue march-in rights on
    the basis of the available information.

    (c) A march-in proceeding shall be initiated by the
    issuance of a written notice by the agency to the contractor and its assignee
    or exclusive licensee, as applicable and if known to the agency, stating that
    the agency is considering the exercise of march-in rights. The notice shall
    state the reasons for the proposed march-in in terms sufficient to put the
    contractor on notice of the facts upon which the action would be based and shall
    specify the field or fields of use in which the agency is considering requiring
    licensing. The notice shall advise the contractor (assignee or exclusive
    licensee) of its rights, as set forth in this section and in any supplemental
    agency regulations. The determination to exercise march-in rights shall be made
    by the head of the agency or his or her designee.

    (d) Within 30 days after the receipt of the written
    notice of march-in, the contractor (assignee or exclusive licensee) may submit
    in person, in writing, or through a representative, information or argument in
    opposition to the proposed march-in, including any additional specific
    information which raises a genuine dispute over the material facts upon which
    the march-in is based. If the information presented raises a genuine dispute
    over the material facts, the head of the agency or designee shall undertake or
    refer the matter to another official for fact-finding.

    (e) Fact-finding shall be conducted in accordance
    with the procedures established by the agency. Such procedures shall be as
    informal as practicable and be consistent with principles of fundamental
    fairness. The procedures should afford the contractor the opportunity to appear
    with counsel, submit documentary evidence, present witnesses and confront such
    persons as the agency may present. A transcribed record shall be made and shall
    be available at cost to the contractor upon request. The requirement for a
    transcribed record may be waived by mutual agreement of the contractor and the
    agency. Any portion of the march-in proceeding, including a fact-finding hearing
    that involves testimony or evidence relating to the utilization or efforts at
    obtaining utilization that are being made by the contractor, its assignee, or
    licensees shall be closed to the public, including potential licensees. In
    accordance with 35 U.S.C. 202(c)(5), agencies shall not disclose any such
    information obtained during a march-in proceeding to persons outside the
    government except when such release is authorized by the contractor (assignee
    or licensee).

    (f) The official conducting the fact-finding shall
    prepare or adopt written findings of fact and transmit them to the head of the
    agency or designee promptly after the conclusion of the fact-finding proceeding
    along with a recommended determination. A copy of the findings of fact shall be
    sent to the contractor (assignee or exclusive licensee) by registered or
    certified mail. The contractor (assignee or exclusive licensee) and agency
    representatives will be given 30 days to submit written arguments to the head
    of the agency or designee; and, upon request by the con- tractor oral arguments
    will be held before the agency head or designee that will make the final
    determination.

    (g) In cases in which fact-finding has been
    conducted, the head of the agency or designee shall base his or her determination
    on the facts found, together with any other information and written or oral
    arguments submitted by the contractor (assignee or exclusive licensee) and
    agency representatives, and any other information in the administrative record.
    The consistency of the exercise of march-in rights with the policy and
    objectives of 35 U.S.C. 200 shall also be considered. In cases referred for
    fact-finding, the head of the agency or designee may reject only those facts
    that have been found to be clearly erroneous, but must explicitly state the
    rejection and indicate the basis for the contrary finding. Written notice of
    the determination whether march-in rights will be exercised shall be made by
    the head of the agency or designee and sent to the contractor (assignee of
    exclusive licensee) by certified or registered mail within 90 days after the
    completion of fact-finding or 90 days after oral arguments, whichever is later,
    or the proceedings will be deemed to have been terminated and thereafter no
    march-in based on the facts and reasons upon which the proceeding was initiated
    may be exercised.

    (h) An agency may, at any time, terminate a
    march-in proceeding if it is satisfied that it does not wish to exercise
    march-in rights.

    (i) The procedures of this part shall also apply to
    the exercise of march-in rights against inventors receiving title to subject
    inventions under 35 U.S.C. 202(d) and, for that purpose, the term ''contractor''
    as used in this section shall be deemed to include the inventor.

    (j) An agency determination unfavorable to the
    contractor (assignee or exclusive licensee) shall be held in abeyance pending
    the exhaustion of appeals or petitions filed under 35 U.S.C. 203(2).

    (k) For purposes of this section the term exclusive
    licensee
    includes a partially exclusive licensee.

    (l) Agencies are authorized to issue supplemental
    procedures not inconsistent with this part for the conduct of march-in
    proceedings.
    Abbott Laboratories #1Last
    Thursday, four groups (the American Medical Students
    Association (AMSA), Knowledge Ecology International (KEI), U.S. Public Interest
    Research Group (PIRG), and the Universities Allied for Essential Medicines
    (UAEM)) filed a petition with the NIH requesting the agency to exercise these
    march-in rights over the anti-AIDS drug ritonavir, exclusively sold by Abbott
    Laboratories.  This is but the fifth
    petition filed under the law, and none of them have been successful.  Before discussing the latest petition it will
    be helpful to review the circumstances and histories of the other petitions,
    all to the NIH, to discern the agency's reasoning for refusing to march-in in
    those instances.

    The first petition was by the
    start-up company CellPro, which produced a device (the Ceprate SC) for
    segregating hematopoietic stem cells from mixtures of cells containing not only
    stem cells but, most importantly, leukemia, lymphoma, and other blood cancer
    cells, thus enabling autologous reimplantation of hematopoietic stem cells in
    patients whose bone marrow had been irradiated to treat such cancers.  (The CellPro story, not surprisingly, told not
    dispassionately by the company founder, Rick Murdock, in Patient No. 1: A True Story of How One CEO Tool on Cancer and Big
    Business in the Fight of His Life
    , New York: Crown Publishers 2000) CellPro
    was sued by Baxter International and Johns Hopkins University, licensee and owner,
    respectively, of patents (U.S. Patent Nos. 4,965,680;
    5,130,144; 5,035,994; and 4,965,204) directed
    to monoclonal antibodies specific for antigens uniquely expressed on the stem
    cell surface.  After losing in the
    district court, CellPro faced an injunction that, while permitting the company
    to stay on the market until Baxter could get FDA approval on a competing device,
    was subject to a 100% (later reduced to 60%) royalty on its "incremental
    profits."  CellPro petitioned Donna
    Shalala, Secretary of Health and Human Services in the Clinton administration,
    alleging that Baxter and Hopkins had failed the provisions of §
    203(a)(1) and (2), for not having obtained FDA approval for its competing
    device in development and for obtaining an injunction that would remove the
    Ceprate SC device from the marketplace. 
    The petition was supported in public hearings by testimony (written and
    oral) by physicians attesting to the use and benefits of the device, and the
    negative medical consequences that would ensue if the NIH did not grant CellPro's march-in petition.

    National Institutes of HealthNonetheless, the NIH (headed at the time by Nobel
    Laureate Harold Varmus) refused to exercise its march-in rights.  In its refusal, the agency stated that it had
    considered whether Baxter had failed to commercialize and decided it
    had not failed, based on the totality of their development and regulatory approval
    activities but disregarding the fact that those activities did not result in a
    commercial, FDA-approved product.  With
    regard to whether there was an "unmet public health or safety need,"
    the agency again decided not to exercise the rights, based on the "considerable
    debate among scientists and clinicians" regarding whether the function of
    the Ceprate device (immunoselection of [hematopoietic] stem cells prior to
    [bone marrow] implantation) was beneficial.  Thus, the agency decided it was "premature" for either CellPro
    or Baxter to claim clinical benefit (although this was presumably at least one
    basis for FDA approval).  Baxter's
    agreement to permit CellPro to remain on the market (albeit under draconian
    royalty terms) was also a factor, and that CellPro did not introduce sufficient evidence that
    Baxter could not satisfy the medical need in the face of Baxter's averments
    that they could satisfy the market (including replacing any Ceprate devices
    removed from the market).  The agency
    also enunciated something close to a philosophy about attempts to have it
    exercise march-in rights under circumstances of a dispute between private
    parties:  the agency was wary of:

    Forced attempts to influence
    the marketplace for the benefit of a single company, particularly when such
    actions may have far-reaching repercussions on many companies' and investors'
    future willingness to invest in federally funded medical technologies.  The
    patent system, with its resultant predictability for investment and commercial
    development, is the means chosen by Congress for ensuring the development and
    dissemination of new and useful technologies.  It has proven to be an effective
    means for the development of health care technologies.  In exercising its
    authorities under the Bayh-Dole Act, NIH is mindful of the broader public
    health implications of a march-in proceeding, including the potential loss of
    new health care products yet to be developed from federally funded research.

    Finally,
    however, the agency reserved the right to "monitor the situation" to
    determine if the factual grounds for its decision proved true.

    In
    2004 a group called Essential Inventions filed a petition based
    not on private interests but on what it characterized as the "public
    interest" for lower prices on the HIV protease inhibitor ritonavir, sold
    by Abbott as Norvir® (the same drug that is the subject of the
    latest petition).  The factual basis for
    the petition was that Abbott had increased the price of the drug by ~400%.  In this case, there was no university party
    involved; Abbott had been funded by research monies from the Reagan
    administration in an effort to develop more effective anti-AIDS drugs.  The
    patents at issue (U.S. Patent Nos. 5,541,206;
    5,635,523; 5,648,497; 5,674,882; 5,846,987; 5,886,036) were the same
    patents that are the subject of the current petition, as was the basis in the
    statute:  that the requirement for licensing on "reasonable terms" ("upon
    terms that are reasonable under the circumstances")
    was violated by Abbott's pricing for Norvir®.  (It should be noted that this portion of the statute would appear to refer to
    the licensing terms between the "contractor" and the licensee and not
    to be related to the price the licensed drugs are sold for.)  This argument is supported by a 2001 law review
    article by Arno & Davis (Peter S. Arno &
    Michael H. Davis, Why Don't We Enforce Existing Drug Price Controls? The
    Unrecognized and Unenforced Reasonable Pricing Requirements Imposed upon
    Patents Derived in Whole or in Part from Federally Funded Research
    , 75
    Tulane L. Rev. 631, 660-661 (2001)), that argued that the legislative history
    and "purpose" of the Bayh-Dole Act indicated that price was part of
    the "reasonable terms" requirement.  The petitioners suggested a smorgasbord of remedies,
    including a 5% royalty, calculated on generic price; a requirement that "every
    manufacturer " contribute to R&D Funds for AIDS; that the NIH adopt
    the "Commonwealth" model of open licensing; and that these licenses
    include the right to distribute worldwide (disregarding that the NIH had no
    authority to countenance patent infringement in other countries).  The NIH held public hearings and received
    written and oral testimony from a "variety
    of groups and individuals representing universities, the AIDS community,
    pharmaceutical interests, drafters of the Bayh-Dole Act, and other interested
    parties."  Again, despite these
    arguments (including arguments that Abbott's pricing was preventing state
    government agencies from providing Norvir®
    to patients), the NIH again refused to exercise its march-in rights, saying:

    [T]he issue
    of the cost or pricing of drugs that include inventive technologies made using
    Federal funds is one which has attracted the attention of Congress in several
    contexts that are much broader than the one at hand.  In addition, because the
    market dynamics for all products developed pursuant to licensing rights under
    the Bayh-Dole Act could be altered if prices on such products were directed in
    any way by NIH, the NIH agrees with the public testimony that suggested that
    the extraordinary remedy of march-in is not an appropriate means of
    controlling prices.  The issue of drug pricing has global implications and,
    thus, is appropriately left for Congress to address legislatively.

    (Interestingly, as a result
    of this petition, Abbott agreed to exempt governmental purchasers, both Federal
    and state, from the price increase, a circumstance raised in the most recent
    petition.)

    Essential Inventions filed
    another petition in 2005, this time involving Pfizer's glaucoma drug Xalatan®.  The grounds for this petition were also drug
    price but here the actual costs ($64/4-6 week supply) were much less than the
    costs for Norvir® (up to ~$14,000/year).  In this case, the drug was developed by researcher at Columbia
    University (owner of U.S. Patent No. 4,359,353) and licensed exclusively to Pfizer; the NIH noted that there
    were other patents owned by Pfizer that relate to
    the drug but are not covered by march-in rights.  Petitioners compared the cost of the drug in
    the U.S. with costs in Canada and several European countries (and disregarded
    the fact that the comparison is with countries having government-subsidized
    medical systems).  The petitioners
    proposed substantially the same terms for licensing under the exercise of the
    NIH's march-in rights for this drug as they had for Norvir® and the NIH refused to exercise the rights for essentially
    the same reasons (repeating verbatim its opinion that controlling drug prices
    was something best left to Congress).

    The most recent petition where the NIH has made a determination arose
    under circumstances where the exclusive licensee, Genzyme, developed
    manufacturing difficulties that severely reduced the availability of its
    licensed drug for treating Fabray's disease, Fabrazyme®.  The petition was filed on behalf of three
    patients (Joseph
    M. Carik, Anita Hochendoner, and Anita Bova)
    and alleged as the basis for exercise of march-in rights that Genzyme could not
    adequately satisfy the public health need for the drug.  Here, the drug, agalsidase beta, was
    developed at Mt. Sinai School of Medicine (owner of U.S. Patent No. 5,356,804) and exclusively licensed
    to Genzyme.  The petitioner requested an "open"
    license (citing such licenses granted by IBM
    as part of computer standard setting and by Microsoft as part of the settlement
    of the government's antitrust action against that company), and also wanted any
    such license to extend to commercial "know-how" including recombinant
    cell lines expressing the drug.  The NIH
    once again refused to exercise its march-in rights, in this case on the grounds
    that permitting other licensees under march-in would
    not solve the problem, due to the time needed (for clinical studies and
    regulatory approval) to get the drug to market (see "HHS Denies Request to Exercise Bayh-Dole 'March-in' Rights for Fabrazyme").  The agency also cited the diligence
    exhibited by Genzyme in addressing the problem and planning to have full
    production sufficient to satisfy the patient population within about 24 months
    from when the manufacturing difficulties arose.  The agency also noted that there were "at least 5 other companies
    worldwide [] engaged in commercial development of alternative treatments"
    that would not need to license Mt. Sinai's patents to enter the marketplace.

    In the latest petition, the allegations again
    involve the cost of Abbott's Norvir® to
    U.S. patients.  In view of the agreement
    following the 2004 petition that has Abbott selling the drug to state and
    Federal government agencies at reduced cost (i.e., not subject to the ~400%
    price increase), the petition focuses on the impact of these costs on private
    parties.  In this regard the petitioners
    argue that, in view of the financial crisis, these high[er] drug costs were "undermining
    the international competitiveness of [U.S.] employers" and harming the
    economy (leaving unsaid the hope for a different outcome from a different
    administration).  The economic aspect of
    the argument is accented by a quotation from a speech by President Obama to the
    AMA on June 15, 2009, where he said, inter alia, that the cost of health care "is
    a threat to our economy."  Specifically, the petition argues that:

    While at one point in
    time the U.S. economy was so healthy and dominant it could ignore such
    concerns [regarding the effects of higher drug costs on the economy], this is no longer the case.  The U.S. economy as a whole is now
    smaller than the combined economy of the members of the European Union.  Between
    1970 and 2011, the U.S. share of the global GDP declined from 35 percent to 22
    percent, and the U.S. share of high-income country GDP has declined from 45
    percent to less than 34 percent.

    With the current
    financial crisis, U.S. unemployment is at high levels, our share of global GDP
    has shrunk sharply since 2000, and we no longer can afford the luxury of paying
    more for drugs invented on NIH grants than do our trading partners in other
    high income countries.

    The petition asks for two specific remedies to
    be imposed "without prejudice to" further march-in rights in response
    to "anticompetitive,
    abusive or unfair practices" by a licensee.  These remedies are:

    • A
    ceiling on prices for U.S. residents, to be imposed when US prices for a drug are higher than 7 of 10 comparison countries,
    among "high income" countries as determined by the World Bank, or prices for U.S. residents, when are 10% higher than the median price in those countries (these
    circumstances would be "presumptively not reasonable" under the
    statute); and

    • Licenses
    specific for use of a patented invention in the development of a "dependent"
    technology, such as a co-formulation of a patented drug with another drug.

    In
    addition, the petition requests imposition of "open" licensing for
    the drug under the agency's march-in rights provisions.  Petitioners suggest two additional "legal
    mechanisms" for achieving these ends:  royalty-free government licenses
    (involving participation by the government in drug distribution, etc.) and the
    grant of government licenses to third parties.

    In
    addition to the public health and unreasonable licensing grounds asserted in the
    petition, petitioners further argue that the Americans with Disabilities Act
    (as it has been interpreted by the Equal Employment Opportunities Commission)
    and the PPACA (the "healthcare law") impose requirements on employers
    that implicate the provisions of § 203(a)(3) that allow the
    agency to exercise march-in rights to permit compliance with Federal
    regulations.  The petitioners clearly are
    using this case as a stalking horse for a more extensive assault on drug
    prices, noting that there are several other drugs that they believe should be
    subject to the price ceiling and open licensing, including sitagliptin (Januvia®)
    and Tobrex® (ophthalmic
    tobramycin).  Finally, the petition
    asserts the policy rationale that "[t]he failure to grant a single
    march-in request in more than 30 years has sent a signal to the patent holder
    that the NIH will permit almost anything, no matter how abusive that action is
    to the public that paid for the research."

    Under
    the relevant regulations, the agency has 60 days to make a decision, which
    would occur after the election and may provide a Christmas present for the
    petitioners.  One fact is clear:  consumers of technology, not producers, are motivated to exercise the relevant
    provisions in the law to influence government to provide extra-agency
    regulation of the drug marketplace and to essentially chose the economic "winners"
    and "losers" in so doing.  It
    is well to remember how famously government control of markets failed during
    the major part of the last century, during a time when the West was
    experiencing technological and economic rejuvenation after a century of world
    warfare.  In a particular instance there
    may be a justification for the government to use its march-in rights to protect
    the public health, such as an epidemic or natural disaster, or when a company
    harms the public health by preventing commercialization of technology (something
    truly contrary to the "purpose" of the Bayh-Dole Act).  Prudence suggests, however, that government
    regulation of drug prices should occur only under circumstance where there is
    government regulation of everything else, and that that eventuality should
    arise sparingly and rare.