• By Andrew Williams —
    USPTO Seal - backgroundThe USPTO has been seeking feedback on the PTAB trial proceedings established by the Leahy-Smith America Invents Act.  A Federal Register Notice from June 27, 2014 contained the "Request for Comments on Trial Proceedings Under the America Invents Act Before the Patent Trial and Appeal Board" (79 Fed. Reg. 36474).  In it, the USPTO outlined 17 issues, or questions, for which the Office is most interested in receiving public comments.  The original deadline for submission was September 16, 2014, the two-year anniversary of the PTAB, but the period was extended this week until October 16, 2014.  Nevertheless, 11 comments were filed by the original deadline.  These can roughly be divided up into three categories: (1) companies, (2) individuals, and (3) intellectual property associations.  In this post, we review the first set of comments — those received from companies.

    Hospira

    Hospira's comments begin by explaining that it is a global specialty pharmaceutical and medication delivery company.  It then responded to each of the 17 questions posed by the Request for Comment found in the Federal Register notice.  Nevertheless, a common theme could be found — Hospira is generally satisfied with how the system has operated thus far and in most instances either advocated that no changes should be made or it deferred to the Board to use its discretion.  Even though it may sound incongruous to submit comments advocating no changes, it did so in anticipation of comments from others.  For example, in response to the question about the Board's practice regarding motions to amend, it is commonly believed that stakeholders are unhappy that the Board has only granted one such motion. Hospira noted that one particular proposal should be rejected –- the shifting of burdens to the petitioner to provide an expert to prove validity of any proposed amendment.  One problem that Hospira identified was that if the petitioner was required to provide an expert, the fixed timeline for PTAB trials would likely need to be adjusted (which is contrary to one of the perceived benefit of the IPR framework –- speed).  Moreover, Hospira noted that it is not unreasonable to require the patent owner to articulate its reasoning via an expert declaration.

    Hospira also commented on the factors enumerated in the Garmin v. Cuozzo case (IPR2012-00001) for requesting additional discovery.  Specifically, it noted that the Garmin factors are consistent with another of the perceived benefits of IPR — limited discovery.  With regard to the "multiple proceedings" questions (No. 7-13), Hospira deferred to the Board's exercise of discretion.  Nevertheless, it did argue that later petitioners should not be foreclosed from pursuing different arguments, even with regard to prior art already considered.  Hospira also addressed the topic of mandatory settlement discussions, and opposed any such requirement if it would impact the statutory timeline for IPR proceedings.

    Finally, in response to the "General" category of questions, Hospira had an interesting take on the interplay of post-issuance challenges before the PTAB and Hatch-Waxman-type litigations.  Hospira advocated that a PTAB written decision nullifying the claims of a challenged patent should serve to trigger the "failure-to-market" forfeiture provisions for first ANDA filers.  The argument went that, similar to a district court proceeding, the Patent Owner would have had the opportunity to present all of its evidence supporting validity.  Of course, this presumably would only become relevant if all remaining claims of the Orange Book listed patents were invalidity by the PTAB.  Nevertheless, this comment highlights the widely held belief that PTAB proceedings can be beneficial to secondary ANDA filers.

    SAS

    Two attorneys for SAS served as "commentators" for general question 17.  Specifically, they took issue with the PTAB's practice of instituting inter partes review on only some of the petitioned claims.  Instead, they argued, once the PTAB determines that the "reasonable likelihood" threshold has been met, all claims challenged in the petition should be included in the proceeding.  As it is, with partial institution, the petitioner is denied the right to appeal the non-instituted claims, which is allegedly contrary to the intent of Congress.

    SAS noted that in the past two years, IPR and CBM petitions have been used with increasing frequency.  In response, the Board has taken steps to ensure timely completion of the trials.  One of these is to treat institutions on a claim-by-claim basis, instead of the petition as a whole.  However, SAS points to several places in the statute which would suggest that this is inappropriate.  First, the standard for institution outlined in 35 U.S.C. § 314(a) is whether "there is a reasonable likelihood that the petitioner would prevail with respect to at least one of the claims challenged in the petition."  The use of the "at least one of the claims" language allegedly suggests that the trial would include more claims than the ones that exceed this threshold.  Moreover, the statute repeatedly uses the phrase "challenged patent claim," rather than "instituted patent claim."  For example, 35 U.S.C. § 318(a) provides that the final written decision must address "the patentability of any patent claim challenged by the petitioner."  Interestingly, SAS pointed out that in a CBM case, the Board noted that "[i]f Congress intended to limit the availability of the covered business method patent review on a claim-by-claim basis, . . . it could have used the term 'claim' rather than 'patent.'"  See Liberty Mutual Ins. Co. v. Progressive Casualty Ins. Co., CBM2012-00002, Paper 66 at 6 (Jan. 23, 2014).  The same logic applies to IPR institutions, SAS argued.

    So, what is the problem with the current claim-by-claim approach?  SAS alleged that partial institution complicates matters for district courts contemplating motions to stay litigation pending IPR trials.  If less than all of the claims are included in the PTAB trial, it might not serve to adequately simplify the issues in the district court.  Moreover, the claims or issues not deemed to have a reasonable likelihood of prevailing are non-appealable, and therefore not entitled to further review.  SAS pointed out that this issue had been raised by IBM when the rules were being developed.  In response, the PTO defended partial review because the statutory estoppel provision applies on a claim-by-claim basis.  Instead, SAS contended, this section demonstrated that Congress knew how to specify claim-by-claim treatment when it wanted to, which it clearly did not in the case of IPR institution.

    The solution — the PTAB should continue to address the patentability of all claims in the institution decision.  However, instead of foreclosing review of the other claims, they should be included in the institution.  The burden of providing sufficient attention to the various claims would fall to the parties because the page limits would force them to focus on the best arguments.  Nevertheless, if a party believed that a particular claim was important, it could provide the requisite evidence necessary for a complete record at the Federal Circuit.

    GEA Process Engineering, Inc.

    GEA Process responded to a single question — "#5 Should a patent owner be able to raise a challenge regarding a real party in interest at any time during trial."  The response — it depends.  The crucial factor, according to GEA Process, is which section of the statute is implicated.  On the one hand, if the challenge is to the Petitioner's standing under 35 U.S.C. § 315 as of the filing date of the petition, GEA Process believes that the patent owner should be allowed to raise this at any time.  However, if the challenge was whether all real parties in interest have been identified, this showing should be made reserved exclusively to the time before institution of the trial.  The distinction made by GEA Process was on jurisdictional grounds.  35 U.S.C. § 315 concerns whether a petitioner had standing, which in Federal court is a jurisdictional question.  As such, it should be allowed to be raised at any time during the trial.  However, 35 U.S.C. § 312, provides for determining whether the petition is complete, not whether someone can bring the suit in the first place.  Therefore, failure to satisfy this section should be waived if not raised before institution.

    Think Computer Corporation

    Think Computer prefaced its comments by pointing out that one of its two patents is being challenged in a CBM petition.  Also, it is a company that correlates a large amount of USPTO data for public use.  Therefore, it believes it has experienced the PTAB from two different perspectives.  In general, Think Computer is in favor of the PTAB and believes that it can serve a vital purpose.  Think Computer's criticism — the system should be set up such that it would not require an attorney.  Instead of the present system, Think Computer advocated for an error-checking web-based form which would allow even inexperienced individuals to construct arguments.  It also complained about the alleged "common practice" of experts submitting reports written entirely by law firm attorneys.  Think Computer did not address the fact that these experts are also subject to deposition, and that, regardless of who prepares what drafts, if the expert does provide final approval of the report, it will become evident during discovery.

    Think Computer's final complaint was with regard to the PTAB docketing system, which was described as "horrendous."  Of course, anyone who has attempted to use it will likely agree with this sentiment.  The PAIR system would have been an obvious alternative, but Think Computer believes that it "is a colossal disaster of its own."  Instead, it suggested that the Patent Office immediately issue an RFQ for both systems and to make a point to use a different vendor.

    Patent Docs will report on the additional comments in a future post, and will continue to monitor the PTAB website to see if any additional comments are submitted before the October 16, 2014 deadline.

  • By Donald Zuhn —

    USPTO SealDuring a session at today's biotechnology/chemical/pharmaceutical (BCP) customer partnership meeting, the U.S. Patent and Trademark Office provided an update on the status of the Myriad-Mayo Guidance.  The BCP session on the Guidance, coming at the end of the Office's first Bicoastal BCP (BCBCP) event, included presentations and comments from June Cohan, Legal Advisor with the USPTO's Office of Patent Legal Administration; and Suzannah K. Sundby of Canady & Lortz LLP at the USPTO's Alexandria, VA campus; and from Jeffery Tung, Patent Counsel, Isis Pharmaceuticals; Paul Naik, Vice President, Intellectual Property, Genentech, Inc.; and Dan Sullivan, Director, TC1600, USPTO at San Jose State University.  Although the BCP session encompassed both the Myriad-Mayo Guidance and the Alice Corp. Guidance, the bulk of the session focused on the former.

    Ms. Cohan spoke first, reminding attendees that at the last BCP meeting in April she had given a presentation on the Guidance following its March release (unlike her earlier presentation, this one was not accompanied by slides).  Since the April BCP meeting, she noted that the Office had conducted a forum on the Guidance in May, had collected comments on the Guidance over the summer, and had taken the opportunity to discuss the Guidance at other events (including during a BIO IPCC workshop in April and at the BIO International Convention in June).  Before discussing the impact of the eighty comments the Office had received in response to the Myriad-Mayo Guidance, Ms. Cohan posed the question that has been on many practitioners' and applicants' minds the past few months:  are there things in the Guidance that the Office can change?  To that question, her answer was a simple "yes".  As a result, she indicated that the Office intended to release "revised" Guidance in about a month.

    Following this welcome announcement, Ms. Cohan discussed what the Office had learned from the comments and public feedback as well as what aspects of the Guidance it intended to change.  She noted that the Office did not explain things as clearly as it could have in the Guidance, and that it was not the Office's intent to give the impression that the Supreme Court has set a high bar for patent eligibility.  Rather, it is the Office's belief that the Court has set a low bar for eligibility.  She also indicated that the Office did not intend to create a per se rule on purified products, and could have provided a better explanation on this aspect as well.

    With respect to questions about the eligibility of hybrid plants, hybrid animals, and antibodies expressed in mouse cells (which otherwise would not have been expressed in such cells), Ms. Cohan noted that all of these constituted patent eligible subject matter.  As for comments questioning the Office's interpretation of certain cases, she pointed out that the Office has gone back a reconsidered those decisions.

    In crafting the Guidance that was issued in March, Ms. Cohan suggested that the Office had "cast too broad a net," and that the Guidance's "significantly different" test, which focuses only on structural differences, might be too narrow.  She indicated that the revised Guidance would permit applicants to establish that a claimed product was significantly different from a natural product by pointing to functional differences (or differences in biological activity).

    As for the factor-based analysis of the Guidance, Ms. Cohan acknowledged that the scheme presented in the Guidance was "too complicated," and that the analytical framework in the revised Guidance would be "simplified".  Practitioners and applicants can also expect to see more examples in the revised Guidance.  Once the revised Guidance is released, Ms. Cohan noted that the Office wants "to keep the dialog going," explaining that several cases are working their way through the courts that could impact the manner in which the Office examines claims for subject matter eligibility.

    Following Ms. Cohan's presentation, Mr. Tung provided a few comments regarding the impact of the Guidance.  He argued, as have other commentators, that in contrast with the Guidance, the Myriad holding is narrow — even in the context of nucleic acids.  Ms. Cohan, however, responded that the Guidance applied to more than just nucleic acids because cases like Funk Bros., Chakrabarty, and Roslin concern more than just nucleic acids (i.e., bacteria and sheep), and therefore, "I don't see how we can limit this to DNA."

    Dr. Naik provided two observations regarding the Guidance.  First, he agreed that more examples were needed, pointing out that examples are extremely powerful tools, especially given something as nebulous and vague as subject matter eligibility.  He also suggested that more examples that were closer to the eligibility line were needed, rather than examples that encompassed subject matter that was too clearly eligible or too clearly ineligible.  Second, he argued that the Guidance's filter was too low, explaining that an analysis in which the recitation of a protein in a claim forced the examiner to quickly move to the third prong of the subject matter eligibility analysis was "unnecessarily broad," and not a good use of either the examiner's or applicant's time.

    Ms. Sundby suggested, as have other commentators, that it would be useful for the Office to create a subject matter eligibility webpage on which it would compile relevant decisions and new examples of patent eligible or ineligible subject matter.

    Note:  Ms. Cohan will be participating in a session that will highlight the Myriad-Mayo Guidance at next week's Biotechnology Industry Organization (BIO) IP & Diagnostics symposium in Washington, DC.  The session, entitled "Squaring the Circle: Obtaining Patents That Are Valid, Commercially Meaningful, and Enforceable," which is sponsored by McDonnell Boehnen Hulbert & Berghoff LLP, will also feature Sherry Knowles of Knowles IP Strategies (former Senior Vice President and Chief Patent Counsel at GlaxoSmithKline) and Patent Docs authors Kevin Noonan and Donald Zuhn.

  • By Paul Tully —

    FDAThe new phone book is here?  No, but close.  The Food and Drug Administration ("FDA") announced on Friday that it has published its first listing of approved biologic drugs.  The list will be supplemented with approved biosimilar alternatives to the biologic drugs, termed the "List of Licensed Biological Products with Reference Product Exclusivity and Biosimilarity or Interchangeability Evaluations."  Inevitably given a "color" designation (following the tradition of the "Orange Book" of approved small molecule drugs), the new listing of biologics is colloquially called the "Purple Book."  Both the Center for Drug Evaluation and Research ("CDER") and the Center for Biologic Evaluation and Research (CBER) have published their own versions of the Purple Book (CDER List of Licensed Biological Products; CBER List of Licensed Biological Products; FDA Purple Book webpage).

    Ucm412239The new lists disclose all approved biologic drugs and the date a product was licensed, as well as "whether the FDA evaluated the biological product for reference product exclusivity under section 351(k)(7) of the PHS Act."  The Purple Book is intended to permit users to determine whether the FDA has licensed a biosimilar and whether such a biosimilar is interchangeable with an already-licensed reference biological product (i.e., an already-licensed FDA biological product).  Both biosimilar and interchangeable biological products will be listed with regard to the reference product to which biosimilarity or interchangeability was demonstrated.  The agency further announced that these lists will be updated "periodically."

    This new resource is certain to become a valuable tool for life-sciences professionals, but it remains to be seen whether relevant patent information will ultimately make its way into the Purple Book, as is prevalent (and required) in its Orange Book cousin.  Stay tuned.

  • By Kevin E. Noonan —

    Ariosa DiagnosticsOn September 2nd, the Patent Trial and Appeals Board (PTAB) entered judgment in an inter partes review styled Ariosa Diagnostics v. Isis Innovation Ltd. (IPR 2012-00022).  The Board found that Ariosa demonstrated, by a preponderance of the evidence of record, that claims 1, 2, 4, 5, 8, 19, 20, 24, and 25 of U.S. Patent No. 6,258,540 are unpatentable under 35 U.S.C. § 102(b).  But the Board found that Ariosa did not carry its burden of showing by a preponderance of the evidence that claims 3, 12, 13, 15, 18, 21, and 22 of the '540 patent were unpatentable under 35 US.C. § 103(a).

    The claims under inter partes review are as follows:

    1.  A method for detecting a paternally inherited nucleic acid of fetal origin performed on a maternal serum or plasma sample from a pregnant female, which method comprises
        amplifying a paternally inherited nucleic acid from the serum or plasma sample and
        detecting the presence of a paternally inherited nucleic acid of fetal origin in the sample.

    2. The method according to claim 1, wherein the foetal nucleic acid is amplified by the polymerase chain reaction.

    3. The method according to claim 2, wherein at least one foetal sequence specific oligonucleotide primer is used in the amplification.

    4. The method according to claim 1, wherein the foetal nucleic acid is detected by means of a sequence specific probe.

    5. The method according to claim 1, wherein the presence of a foetal nucleic acid sequence from the Y chromosome is detected.

    8. The method according to claim 1, wherein the presence of a foetal nucleic acid from a paternally-inherited non-Y chromosome is detected.

    12. The method according to claim 5, for determining the sex of the foetus.

    13. The method according to claim 5, which comprises determining the concentration of the foetal nucleic acid sequence in the maternal serum or plasma.

    15. The method according to claim 13, for the detection of a maternal or foetal condition in which the level of foetal DNA in the maternal serum or plasma is higher or lower than normal.

    18. The method according to claim 13, for detection of a foetal chromosomal aneuploidy.

    19. The method according to claim 1, wherein the sample contains foetal DNA at a fractional concentration of total DNA of at least about 0.14%, without subjecting it to a foetal DNA enrichment step.

    20. The method according to claim 19, wherein the fractional concentration of foetal DNA is at least about 0.39%.

    21. A method of performing a prenatal diagnosis, which method comprises the steps of:
        (i) providing a maternal blood sample;
        (ii) separating the sample into a cellular and a non-cellular fraction;
        (iii) detecting the presence of a nucleic acid of foetal origin in the non-cellular fraction according to the method of claim 1;
        (iv) providing a diagnosis based on the presence and/or quantity and/or sequence of the foetal nucleic acid.

    22. The method according to claim 21, wherein the non-cellular fraction as used in step (iii) is a plasma fraction.

    24. A method for detecting a paternally inherited nucleic acid on a maternal blood sample, which method comprises:
        removing all or substantially all nucleated and anucleated cell populations from the blood sample,
        amplifying a paternally inherited nucleic acid from the remaining fluid and subjecting the amplified nucleic acid to a test for the Paternally inherited fetal nucleic acid.

    25. A method for performing a prenatal diagnosis on a maternal blood sample, which method comprises
        obtaining a non-cellular fraction of the blood sample
        amplifying a paternally inherited nucleic acid from the non-cellular fraction
        and performing nucleic acid analysis on the amplified nucleic acid to detect paternally inherited fetal nucleic acid.

    (Italics indicating claims found unpatentable by the PTAB.)

    The decision provided the following synopsis of the asserted grounds of unpatentability:

    Table
    wherein the cited references are:

    • Lo et al., Presence of Fetal DNA in Maternal Plasma and Serum, 350 LANCET 485–487 (1997).

    • Simpson et al., Isolating Fetal Cells in Maternal Circulation for Prenatal Diagnosis, 14 PRENATAL DIAGNOSIS 1229–1242 (1994).

    • Kazakov et al., Extracellular DNA in the Blood of Pregnant Women, 37(3) CYTOLOGY (TSITOLOGIA) 232–236 (1995).

    • Bianchi et al., Fetal Cells in Maternal Blood: Determination of Purity and Yield by Quantitative Polymerase Chain Reaction, 171 AM. J. OBST.GYNECOL. 922–26 (1994).

    • Schallhammer et al., Phenotypic Comparison of Natural Killer Cells from Peripheral Blood and from Early Pregnancy Decidua, 3 EARLYPREGNANCY: BIOLOGY AND MEDICINE 15–22 (1997).

    The decision noted that Ariosa abandoned at oral argument its first two grounds of unpatentability set forth above.

    Isis InnovationArriving at its bases for deciding the questions before it, the Board reiterated its decision (as set forth in its Decision on Petition) rejecting Isis's arguments that Ariosa lacked standing under § 315 to petition for inter partes review based on the litigation between the parties.  Specifically, the Board addressed Isis's contention that "it 'is of no moment'" that Ariosa raised its validity challenge as an affirmative defense (in the face of Isis's infringement allegations prompted by Ariosa's declaratory judgment action for judgment of non-infringement).  Isis also argued unsuccessfully that the exception under § 315(a)(3) does not apply because Isis (and its licensee, Sequenom) did not initiate the civil action.  Finally, Isis argued (again unsuccessfully) that granting Ariosa's petition for inter partes review would "thwart the intent of Congress," inter alia, by permitting "patent owner harassment."  For clarity, the Board set forth the precise question decided in granting Ariosa's petition:

    [W]hether filing a Declaratory Judgment of non-infringement in District Court bars Ariosa from later filing a petition for Inter Partes Review under 35 U.S.C. § 315(a), and whether the express mention of a counterclaim of invalidity in 35 U.S.C. § 315(a)(3) mandates interpreting the statute such that raising an affirmative defense of invalidity in response to a compulsory counterclaim of infringement deprives Ariosa of standing to file for inter partes review.

    The Board's decision to grant Ariosa's petition was based on its determination that the plain meaning of the statute, as interpreted by the definitions of "filing" and "a civil action" provided by the Federal Rules of Civil Procedure, is that the bar arises when a petitioner has filed a complaint prior to filing the inter partes review petition.  The Board also distinguished between raising an affirmative defense and filing a counterclaim, based on its understanding of the differences defined by the Supreme Court in Altvater v. Freeman, 319 U.S. 359 (1943).  In that case, the Court decided that a counterclaim is "justiciable" even in the face of a determination of non-infringement, while an affirmative defense of invalidity under similar circumstances would be to "decide a hypothetical case."  Citing Cardinal Chem. Co. v. Morton Int'l, Inc., 508 U.S. 83 (1993), the Board decided that "it is clear []that there is a fundamental difference between an affirmative defense of invalidity and a counterclaim of invalidity," and thus that the declaratory judgment of non-infringement by Ariosa in this case did not fall within the activities the statute provides would bar the grant of the inter partes review petition.

    The Board also rejected Isis's contention that § 315(b) barred grant of the petition because Isis had filed its infringement lawsuit more than a year before filing the inter partes review petition, because the infringement complaint had been dismissed without prejudice, citing the Board's prior decision to this effect in Macauto U.S.A. v. BOS GmbH & KG, Case IPR2012-00004 (PTAB January 24, 2013).

    In the last of the procedural issues discussed in the decision, the Board rejected Isis's contention that granting Ariosa's Motion for Joinder (of two separate inter partes actions against different combinations of claims) was improper (despite the statutory language being limited to joinder of parties; § 315(c)) based on its practice in prior proceedings permitting such joinder, including Microsoft Corp. v. Proxycann, Inc., Case IPR2013-00109 (PTAB February 25, 2013), and Samsung Elecs. Co., Ltd. v. Virginia Innovation Scis., Inc., Case IPR2014-00557 (PTAB June 13,2014).  The Board found no prohibition of issue joinder in either the statutory language or the legislative history, and relied on its rules (37 CFR § 42.1(b)) implementing § 316, specifying that "[t]he rules are to be construed so as to ensure the just, speedy, and inexpensive resolution of a proceeding."

    Turning to the merits of Ariosa's invalidity contentions, the Board construed the claim term "detecting."  In doing so, the Board made the following claim construction determination applying the "broadest reasonable interpretation" standard under 37 C.F.R. § 42.100(b):

    "detecting" means that the amplified DNA was detected, but not that such DNA be identified as being of paternal or fetal origin.

    This construction, according to the Board, is consistent with the construction used by Isis's expert at trial, and further that in the context of the claims Isis could have limited what was detected to paternal and fetal DNA by stating that "only" this DNA was detected (but, says the Board, "they chose not to do so," implying a decision point that there is no evidence Isis ever encountered).  While the Board relies on claim construction canons (such as not reading limitations from the specification into the claims), it is incongruous that the Board rendered this construction decision (which is dispositive; see below) in interpreting the scope and meaning of claims specifically directed towards detecting paternal or fetal DNA in pregnant female blood.

    With this construction established, the decision turned to the cited references.  Regarding the Kazkov reference, asserted against claims 1, 2, 4, 5, 8, 19, 20, 24 and 25, the Board found that the practice of the methods disclosed in the reference inherently anticipated these claims, because amplification of DNA from maternal blood would necessarily detect "paternally inherited nucleic acid of fetal origin."  Specifically, the Board found that the reference taught that:  1) extracellular DNA is present in human blood; 2) the amount of this DNA increases with pregnancy; 3) this DNA was detected using PCR; 4) the amplified DNA was detected (using gel electrophoresis); and 5) the primers Kazakov used would have detected paternal DNA (insofar as the primers amplified sequences present (albeit not exclusively) on the Y chromosome).

    Under the Board's construction, "Kazakov discloses the same method of claim 1."  The Board stated that "all that is required by the amplification step of claim 1 is a step of amplifying nucleic acid from a serum or plasma sample from a pregnant female, such as by PCR, as such amplified nucleic acid would necessarily include fetal nucleic acid, and the fetal nucleic acid necessarily includes paternally inherited nucleic acid," so long as "the detecting step does not require that the detected nucleic acid specifically be identified as being inherited from the father or even as being from the fetus, only that it be identified as containing some level of nucleic acid, which would include, necessarily, nucleic acid from the fetus that was inherited from the father."  The Board further reasoned that:

    [I]f the ordinary artisan were to follow the teachings of Kazakov, and perform PCR on the serum obtained from the blood obtained from a pregnant female, that blood would inherently contain paternally inherited fetal nucleic acid.  That nucleic acid would be amplified and detected by the experiments of Kazakov as such a result is necessarily inherent.  That is, the amplification and detection of paternally inherited fetal nucleic acid would be a new benefit of a known process.

    The Board further "credited" the testimony of Ariosa's expert, that the methods disclosed in Kazakov were all "conventional," although there is no discussion on the significance of these findings to the Board's decision.  And Isis's arguments that the possibility that fetal or paternal DNA would not be detected received no traction with the Board in refuting its inherent anticipation determination (which is not surprising, because the Board's error was in claim construction not the application of the facts to the incorrectly construed claims).

    Having made this determination regarding claim 1, the Board did not find (and in some cases Isis did not provide) any basis for the same reasoning not to be applied to claims 2, 4, 5, 8, 19, 20, 24, and 25, and thus found these claims to also be inherently anticipated by the Kazakov reference.

    Regarding obviousness, the Board held that Ariosa had not borne its burden of establishing invalidity by a preponderance of the evidence.  Limiting its considerations to the combination of the Kazakov and Simpson references, the Board found that the Simpson reference taught detection of fetal cells, not fetal cell-derived DNA in maternal whole blood, and that the art taught that detecting fetal cells in maternal blood was "a rare occurrence."  Under these circumstances, the Board found that:

    [T]he ordinary artisan would not have used the serum sample of Kazakov, which is a cell-free sample, for the whole blood sample of Simpson, which contains the cellular fraction, for the analysis of fetal DNA as taught by Simpson.  The ordinary artisan would not have had a reasonable expectation that the fetal DNA would have been present in maternal serum in sufficient quantities for detection using amplification methods such as PCR given the understanding in the art that fetal cells were a rare occurrence in maternal blood.  Moreover, Kazakov's lack of teaching that the increase in extracellular DNA in the serum of pregnant females is due to the presence of cell-free fetal DNA in the serum further supports that conclusion.

    Ironically, the Board's decision on obviousness relies on the failure of the art to recognize that fetal or paternal DNA could be detected in maternal blood, a distinction that the Board did not credit in its claim construction leading to its contrary decision regarding (inherent) anticipation by the Kazakov reference.

    The Board also rejected Ariosa's obviousness contentions based in the combination of the Kazakov, Simpson, Schallhammer (earlier determined by the Board to be cumulative over the teachings of the Simpson reference) and Bianchi references (wherein Bianchi taught Y-chromosome specific primers), based on the same lack of expectation that fetal cells (or their DNA) would be expected by the skilled artisan to be present in maternal blood.

    The Board also denied Isis's motion to amend its claims, once again rendering a decision that contradicts the rationale for construing claims according to the broadest reasonable interpretation of its terms (i.e., that when before the PTO the patentee can amend her claims).  The proposed amended claims are as follows, and were proposed to overcome the Board's broad construction of the term "detected" in the patented claims:

    28.  A method for detecting a paternally inherited nucleic acid of fetal origin performed on a maternal serum or plasma sample from a pregnant female, which method comprises
        amplifying a paternally inherited nucleic acid from the serum or plasma sample, and
        detecting the presence of a paternally inherited nucleic acid of fetal origin in the sample by determining that the paternally inherited fetal nucleic acid contains a sequence not possessed by the pregnant female.

    31.  A method for detecting a paternally inherited nucleic acid of fetal origin performed on a maternal serum or plasma sample from a pregnant female, which method comprises
        amplifying a paternally inherited nucleic acid from the serum or plasma sample, [and]
        detecting the presence of a paternally inherited nucleic acid of fetal origin in the sample by determining that the paternally inherited nucleic acid of fetal origin contains a sequence not possessed by the pregnant female,
    and
    determining fetal sex, fetal chromosomal aneuploidy, fetal mutation, fetal RhD status, or fetal paternally-inherited DNA polymorphism
    .

    In a Kafka-esque fashion, the Board denied the motion because the patentee is under a burden of showing she is "entitled" to the "relief" of amending claims, and the burden imposed is to establish that the claims are patentable under all sections of the Patent Act.  Even though inter partes review is limited to the issues of anticipation and obviousness based on prior art, the Board permits a challenge on the patentability of amended claims on all the statutory sections.  Here, citing the District Court's decision (and in the face of the pending Federal Circuit appeal), the Board determined that Isis had failed to carry its burden of showing that the proposed amended claims are patent-eligible under § 101.

    PTAB recognized that claims 1, 2, 4, 5, 8, 19–22, 24, and 25 of the '540 patent were declared invalid in Ariosa Diagnostics v. Sequenom.  With the exception of claim 21, this outcome suggests that Ariosa could have invalidated the claims in the district court under § 102(b) and done much less violence to the patent system in doing so.  It also raises the possibility that Sequenom may move the Federal Circuit to vacate the District Court's judgment based on § 101 in view of the Office's determination that the claims are invalid under § 102(b).

    The Board's decision provides Isis with another opportunity to appeal to the Federal Circuit, this time on the PTAB's decision invalidating these claims.  Assuming that the Court reverses the District Court on the patent eligibility issue, the Board has provided for review by the Court with issues of claim construction as well as procedural questions decided by the Board in instituting the petition in the first place.

  • By Andrew Williams —

    USPTO SealIf you were planning on letting the U.S. Patent and Trademark Office know how you feel about the use of the "Broadest Reasonable Interpretation" standard for claim construction during PTAB trials or the near impossibility of amending claims in IPRs, you have an additional month.  As we have previously reported, the USPTO is seeking feedback on the PTAB trial proceedings established by the Leahy-Smith America Invents Act.  This request can be found in a Federal Register Notice from June 27, 2014, entitled "Request for Comments on Trial Proceedings Under the America Invents Act Before the Patent Trial and Appeal Board."  Specifically, the USPTO outlined 17 issues, or questions, for which the Office is most interested in receiving public comments.  The original deadline for submission was tomorrow, September 16, 2014, the two-year anniversary of the PTAB.  However, the period for public comment was extended until October 16, 2014 according to a document found on the Patent Office website entitled "USPTO Extending Comment Period on Trial Proceedings Under the America Invents Act Before the Patent Trial and Appeal Board.  The Patent Office stated that the extension was made "[i]n view of stakeholder requests for additional time to submit comments on the new administrative trial proceedings."  Comments on all aspects of PTAB trial proceedings are to be e-mailed to TrialsRFC2014@uspto.gov.

  •         By Sherri Oslick —

    Gavel About Court Report:  Each week we will report briefly on recently filed biotech and pharma cases.

    Millennium Pharmaceuticals Inc. v. Glenmark Pharmaceuticals Ltd. et al.
    1:14-cv-01156; filed September 10, 2014 in the District Court of Delaware

    • Plaintiff:  Millennium Pharmaceuticals Inc.
    • Defendants:  Glenmark Pharmaceuticals Ltd.; Glenmark Generics Ltd.; Glenmark Generics Inc. USA

    Infringement of U.S. Patent Nos. 6,713,446 ("Formulation of Boronic Acid Compounds," issued March 30, 2004) and 6,958,319 (same title, issued October 25, 2005), licensed exclusively to Millennium, following a Paragraph IV certification as part of Glenmarks' filing of an ANDA to manufacture a generic version of Millenium's Velcade® (bortezomib, used to treat multiple myeloma).  View the complaint here.

    Teva Pharmaceuticals USA Inc. et al. v. Sandoz Inc. et al.
    1:14-cv-01171; filed September 10, 2014 in the District Court of Delaware

    • Plaintiffs:  Teva Pharmaceuticals USA Inc.; Teva Pharmaceutical Industries Ltd; Teva Neuroscience Inc; Yeda Research and Development Co Ltd.
    • Defendants:  Sandoz Inc.; Momenta Pharmaceuticals Inc.

    Teva Pharmaceuticals USA Inc. et al. v. Dr Reddy's Laboratories Ltd et al.
    1:14-cv-01172; filed September 10, 2014 in the District Court of Delaware

    • Plaintiffs:  Teva Pharmaceuticals USA Inc.; Teva Pharmaceutical Industries Ltd; Teva Neuroscience Inc; Yeda Research and Development Co Ltd.
    • Defendants:  Dr Reddy's Laboratories Ltd; Dr Reddy's Laboratories Inc.

    The complaints in these cases are substantially identical.  Infringement of U.S. Patent Nos. 8,232,250 ("Low Frequency Glatiramer AcetateTherapy," issued July 31, 2012) and 8,399,413 (same title, issued March 19, 2013) following a Paragraph IV certification as part of defendants' filing of an ANDA to manufacture a generic version of Teva's Copaxone® (glatiramer acetate injection, used for the reduction or frequency of relapses in patients with relapsing-remitting multiple sclerosis).  View the Sandoz complaint here.

    Fresenius Kabi USA, LLC v. Emcure Pharmaceuticals USA Inc. et al.
    1:14-cv-01141; filed September 8, 2014 in the District Court of Delaware

    • Plaintiff:  Fresenius Kabi USA, LLC
    • Defendants:  Emcure Pharmaceuticals USA Inc.; Emcure Pharmaceuticals Ltd.

    Fresenius Kabi USA, LLC v. Emcure Pharmaceuticals USA, Inc. et al.
    1:14-cv-05584 filed September 8, 2014 in the District Court of New Jersey

    • Plaintiff:  Fresenius Kabi USA, LLC
    • Defendants:  Emcure Pharmaceuticals USA Inc.; Emcure Pharmaceuticals Ltd.

    The complaints in these cases are substantially identical.  Infringement of U.S. Patent No. 8,476,010 ("Propofol Formulations with Non-Reactive Container Closures," issued July 2, 2013) following a Paragraph IV certification as part of Emcure's filing of an ANDA to manufacture a generic version of Fresenius' Diprivan® (propofol injectable emulsion, used for the induction and maintenance of general anesthesia and sedation in certain patient populations).  View the Delaware complaint here.

    Salix Pharmaceuticals, LTD et al. v. Mylan Pharmaceuticals Inc.
    1:14-cv-00152; filed September 8, 2014 in the Northern District of West Virginia

    • Plaintiffs:  Salix Pharmaceuticals, LTD; Salix Pharmaceuticals, Inc; Glycyx Pharmaceuticals, LTD
    • Defendant:  Mylan Pharmaceuticals Inc.

    Infringement of U.S. Patent Nos. 6,197,341 ("Formulations of Balsalazide and Its Derivatives," issued March 6, 2001) and 8,497,256 ("Formulations and Uses of 2-Hydroxy-5-Phenylazobenzoic Acid Derivatives for the Treatment of Males," issued July 30, 2013) following a Paragraph IV certification as part of Mylan's filing of an ANDA to manufacture a generic version of Salix's Giazo® (balsalazide disodium, used for the treatment of mildly to moderately active ulcerative colitis in male patients 18 years of age and older).  View the complaint here.

    AbbVie Inc. v. Hetero Labs Ltd. et al.
    1:14-cv-01137; filed September 5, 2014 in the District Court of Delaware

    • Plaintiff:  AbbVie Inc.
    • Defendants:  Hetero Labs Ltd.; Hetero USA Inc.; Hetero Labs Ltd.; Hetero USA Inc.

    Infringement of U.S. Patent No. 8,691,878 ("Solid Pharmaceutical Dosage Form," issued April 8, 2014) following a Paragraph IV certification as part of Hetero's filing of an ANDA to manufacture a generic version of AbbVie's Norvir® (ritonavir, used to treat human immunodeficiency virus (HIV) infection).  View the complaint here.

    Bristol-Myers Squibb Co. et al. v. Merck & Co Inc.
    1:14-cv-01131; filed September 4, 2014 in the District Court of Delaware

    • Plaintiffs:  Bristol-Myers Squibb Co.; Ono Pharmaceutical Co., Ltd.
    • Defendant:  Merck & Co. Inc.

    Infringement of U.S. Patent No. 8,728,474 ("Immunopotentiative Composition," issued May 20, 2014), licensed to BMS, based on Merck's anticipated manufacture and sale of pembrolizumab for the treatment of cancer.  View the complaint here.

    Otsuka Pharmaceutical Co., Ltd. V. Zhejiang Huahai Pharmaceutical Co., Ltd., et al.
    1:14-cv-05537; filed September 4, 2014 in the District Court of New Jersey

    • Plaintiff:  Otsuka Pharmaceutical Co., Ltd.
    • Defendants:  Zhejiang Huahai Pharmaceutical Co., Ltd.; Huahai US Inc.; Prinston Pharmaceutical Inc.; Solco Healthcare U.S., LLC

    Infringement of U.S. Patent Nos. 8,017,615 ("Low Hygroscopic Aripiprazole Drug Substance and Process for the Preparation Thereof," issued September 13, 2011), 8,580,796 (same title, issued November 12, 2013), and 8,642,760 (same title, issued February 4, 2014) following a Paragraph IV certification as part of Zhejiang Huahai's filing of an ANDA to manufacture a generic version of Otsuka's Abilify® (aripiprazole, used to treat bipolar disorder and schizophrenia).  View the complaint here.

    Astrazeneca Pharmaceuticals LP et al. v. Sagent Pharmaceuticals, Inc.
    1:14-cv-05539; filed September 2, 2014 in the District Court of New Jersey

    • Plaintiffs:  Astrazeneca Pharmaceuticals LP; Astrazeneca UK Ltd.; Astrazeneca AB
    • Defendant:  Sagent Pharmaceuticals, Inc.

    Infringement of U.S. Patent Nos. 6,774,122 ("Formulation," issued August 10, 2004), 7,456,160 (same title, issued November 25, 2008), 8,329,680 (same title, issued December 11, 2012), and 8,466,139 (same title, issued June 18, 2013) following a Paragraph IV certification as part of Sagent's filing of an ANDA to manufacture a generic version of AstraZeneca's Faslodex® (fulvestrant injection, used to treat hormone receptor positive metastatic breast cancer in postmenopausal women with disease progression following antiestrogen therapy).  View the complaint here.

  • CalendarSeptember 16, 2014 – "Section 103 and Obviousness: Capitalizing on CCPA and Early Federal Circuit Precedent — Strategies for Withstanding Obviousness Rejections and Attacks on Patent Validity and Patentability" (Strafford) – 1:00 to 2:30 pm (EDT)

    September 16, 2014 – First Inventor to File America Invents Act Roadshow (U.S. Patent and Trademark Office) – Concord, NH

    September 17, 2014 – Biotechnology/Chemical/Pharmaceutical (BCP) Customer Partnership Meeting (U.S. Patent and Trademark Office) – 12:00 to 5:00 pm (EDT)

    September 18, 2014 – First Inventor to File America Invents Act Roadshow (U.S. Patent and Trademark Office) – Madison, WI

    September 18-19, 2014 – FDA Boot Camp (American Conference Institute) – Boston, MA

    September 23, 2014 – First Inventor to File America Invents Act Roadshow (U.S. Patent and Trademark Office) – Alexandria, VA (webcast)

    September 25, 2014 – "Charting New Territory: Prosecution and Diligence Strategies in the Wake of the AIA" (American Bar Association Center for Professional Development and Section of Intellectual Property Law) – 1:00 to 2:30 pm (ET)

    September 26, 2014 – IP & Diagnostics Symposium (Biotechnology Industry Organization) – Alexandria, VA

    September 30, 2014 – 2014 Intellectual Property Continuing Legal Education Seminar (DuPont and Widener University School of Law) – Wilmington, DE

    September 30, 2014 – First Inventor to File America Invents Act Roadshow (U.S. Patent and Trademark Office) – Dallas, TX

    September 30 – October 1, 2014 – Paragraph IV Disputes Master Symposium (American Conference Institute) – Chicago, IL

    October 2, 2014 – "USPTO Guidance for Determining Subject Matter Eligibility In View of U.S. Supreme Court's Mayo and Myriad Decisions" (McDonnell Boehnen Hulbert & Berghoff LLP) – 10:00 to 11:15 am (CT)

    October 2, 2014 – "Conflicts in Patent Prosecution: Minimizing Risks of Malpractice Liability and Ethics Sanctions" (Strafford) – 1:00 to 2:30 pm (EDT)

    October 2, 2014 – First Inventor to File America Invents Act Roadshow (U.S. Patent and Trademark Office) – Denver, CO (webcast)

    October 9, 2014 – First Inventor to File America Invents Act Roadshow (U.S. Patent and Trademark Office) – Atlanta, GA

    October 20-22, 2014 – Business of Biosimilars (Institute for International Research) – Boston, MA

    October 29-30, 2014 - Congress on PIV Litigation (Momentum) – Philadelphia, PA

    ***Patent Docs is a media partner of this conference or CLE

  • USPTO Building FacadeThe U.S. Patent and Trademark Office will be holding its next biotechnology/chemical/pharmaceutical (BCP) customer partnership meeting on Wednesday, September 17, 2014.  The latest BCP meeting will be the Office's first Bicoastal BCP (BCBCP) event, with the meeting being held simultaneously in the Madison Auditorium on the USPTO Alexandria Campus (East Coast) and Engineering Building Conference Room of San Jose State University (West Coast).  During the inaugural BCBCP, participants will be able to interact with Office personnel in person on both coasts, or alternatively, via webcast.

    The agenda for the meeting is as follows:

    • Welcoming Remarks (12:00 – 12:10 pm EDT) — Margaret A. Focarino, Commissioner for Patents, USPTO (Alexandria)

    • Opening Remarks (12:10 – 12:30 pm EDT) — John Cabeca, Director, Silicon Valley Office, USPTO (San Jose); Jerry Lorengo Director, TC1600, USPTO (Alexandria); Dan Sullivan Director, TC1600, USPTO (San Jose); and Wanda Walker Director, TC1600, USPTO (Alexandria)

    • Improper Markush (12:30 – 1:20 pm EDT) — Joe Mallon of Knobbe, Martens, Olson & Bear, LLP (San Jose)

    • Break (1:20 – 1:30 pm EDT)

    • OPAP Procedure and Tips (1:30 – 2:10 pm EDT) — Kevin Little Office of Patent Application Processing (OPAP), USPTO (Alexandria)

    • RCE Statistics and Reduction Initiatives. Brief Overview of AFCP 2.0 & QPIDS, and other Patent Application Initiatives (2:10 – 2:40 pm EDT) — Directors, TC1600 and Silicon Valley Office, USPTO

    • Discussion/Lunch (2:40 – 3:40 pm EDT) — Directors, TC1600 and Silicon Valley Office, USPTO

    • Break (3:40 – 3:50 pm EDT)

    • Myriad Guidance and CLS Decision (3:50 – 4:50 pm EDT) — Paul Naik Vice President, Intellectual Property, Genentech, Inc. (San Jose); Suzannah K. Sundby, Partner, Canady & Lortz LLP (Alexandria); Jeffery Tung Patent Counsel, Isis Pharmaceuticals (San Jose); June Cohan Legal Advisory, OPLA, USPTO (Alexandria)Jerry Lorengo Director, TC 1600, USPTO (Alexandria); and Dan Sullivan Director, TC1600, USPTO (San Jose)

    • Closing Remarks/Discussion (4:50 – 5:00 pm EDT) — John Cabeca, Director, Silicon Valley Office, USPTO (San Jose); Jerry Lorengo Director, TC1600, USPTO (Alexandria); Dan Sullivan Director, TC1600, USPTO (San Jose); and Wanda Walker Director, TC1600, USPTO (Alexandria)

    The meeting can be viewed online here.  The event number is 642 123 660 and the event password is 12345.  Additional information regarding the BCP customer partnership meeting can be found here.

  • USPTO SealThe U.S. Patent and Trademark Office will be hosting seven roadshow events between September 16 and October 9 to increase understanding of the First Inventor to File (FITF) provisions of the Leahy-Smith America Invents Act (AIA).  At each roadshow, panelists will discuss FITF statistics to date, the applicability of the FITF provisions on patent applications filed today, the FITF statutory framework and its exceptions, and AIA evidentiary declaration practice useful to invoke these exceptions.

    Roadshow events will be held on the following dates:

    Tuesday, September 16 — Franklin Pierce IP Center at UNH School of Law (Concord, NH) — 1:00 to 5:00 pm

    Thursday, September 18 — University of Wisconsin-Madison, Engineering Hall (Madison, WI) — 1:00 to 5:00 pm

    Tuesday, September 23 — USPTO Headquarters (Alexandria, VA) — 1:00 to 5:00 pm

    Tuesday, September 30 — Dallas Bar Association Belo Mansion (Dallas, TX) — 1:00 to 5:00 pm

    Thursday, October 2 — USPTO Rocky Mountain Regional Office, Byron G. Rogers Federal Building (Denver, CO) — 9:00 am to 1:00 pm

    Tuesday, October 7 — Cupertino Community Hall (Silicon Valley, CA) — 9:00 am to 1:00 pm

    Thursday, October 9 — Georgia Institute of Technology, Scheller College of Business (Atlanta, GA) — 1:00 to 5:00 pm

    The agenda for the roadshows will be as follows:

    Introductory Remarks (10 min)
    FITF – A Year Plus In Review (20 min)
    • Review of examiner training
    • Statistics to date
    Will My Application Be Examined Under AIA (FITF) Or Not? (75 min)
    • 1.55/1.78 statements
    • The power of the ADS
    • Have you checked your filing receipt?
    • Scenarios
    Break (15 min)
    FITF Overview and Tips on Responding to Prior Art Rejections (60 min)
    • 35 USC 102(a)(1) and 102(a)(2) – bases for rejections
    • 35 USC 102(b)(1) and 102(b)(2) – exception provisions
    • Scenarios
    Effective Use of AIA (FITF) Evidentiary Declarations (45 min)
    • 37 CFR 130(a) and 130(b)
    • Scenarios
    Tour of the AIA (FITF) Website and Q&A (15 min)

    The roadshows in Alexandria on September 23 and Denver on October 2 will be webcast live through the USPTO website.  The September 23 roadshow can be accessed here, using the event number 643 625 972, and event password 12345.  Those wishing to call in can use the toll number 1-571-270-7000 and the access code 643 625 972.  The October 2 roadshow can be accessed here, using the event number 649 219 415, and event password 12345.  Those wishing to call in can use the toll number 1-571-270-7000 and the access code 649 219 415.

    The roadshow events are free and open to the public, but seating is available on a first-come, first-served basis.  Additional information regarding the roadshows can be found here.

    Fitf_roadshow_map

  • By Andrew Williams —

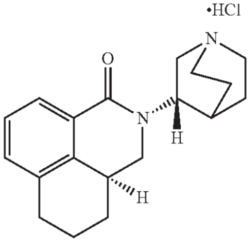
    Accord HealthcareLast week, on September 2, 2014, Accord Healthcare, Inc. ("Accord") filed what appears to be the second-ever Post-Grant Review ("PGR") (see Petition).  This PGR was for U.S. Patent No. 8,598,219 ("the '219 Patent"), which is jointly assigned to Helsinn Healthcare S.A. and Roche Palo Alto, LLC (collectively "Helsinn").  As a reminder, PGRs are the third type of post-issuance review procedures established by the America Invents Act ("AIA") — the other two being Inter Partes Review and Covered Business Method Patent Review (IPR and CBM, for short).  However, the reason that only one other PGR has been filed to date is because this type of proceeding only applies to patents that were examined pursuant to the new First Inventor to File scheme established by the AIA.  And because such applications could only be filed on or after March 16, 2013, there are only a limited number of such patents that are presently eligible for PGR.  One of the other significant differences between IPRs and PGRs is that the latter is not limited to certain types of prior art validity attacks (such as 102 or 103), but instead any type of validity challenge available in District Court is essentially available in front of the Patent Trial and Appeals Board ("PTAB").  This includes attacks under 35 U.S.C. § 112, such as allegations of a lack of enablement, a lack of written description, and a failure to distinctly claim the invention.  Accord took full advantage of this in its petition for PGR2014-00010, in which Accord alleged that Helsinn's patent related to liquid pharmaceutical formulations of palonosetron should not have been issued by the Patent Office.

    The '219 patent had been asserted in several Hatch-Waxman litigations involving ALOXI®, which is a palonosetron formulation indicated to help prevent nausea and vomiting following chemotherapy.  Palonosetron hydrochloride, the active pharmaceutical ingredient, has the following structural formula:

    Structure
    The '219 patent is a member of a family of patents directed to formulations of palonosetron hydrochloride.  Importantly, this patent was filed as a continuation-in-part application on May 23, 2013, with a letter that asserted that claim 9 only had support because of a newly added example, and therefore was subject to the AIA.  Only claims 1-5 and 8 of the '219 patent are the subject of this petition, with claim 1 reading:

    1.    A pharmaceutical single-use, unit-dose formulation for intravenous administration to a human to reduce the likelihood of cancer chemotherapy-induced nausea and vomiting, comprising a 5 mL sterile aqueous isotonic solution, said solution comprising:
        palonosetron hydrochloride in an amount of 0.25 mg based on the weight of its free base;
        from 0.005 mg/mL to 1.0 mg/mL EDTA;
        and from 10 mg/mL to 80 mg/mL mannitol,
        wherein said formulation is stable at 24 months when stored at room temperature.

    Claim 8 is the only other challenged independent claim, and it reads identically, except for a stability limitation of 18 months when stored at room temperature.  This patent issued on December 3, 2013, and the PGR petition was filed within the requisite nine months.

    The petition pointed out that during the prosecution of the '219 patent and its family, the Patent Office had rejected the claimed formulations as obvious.  In response, the applicants submitted a declaration from inventor Daniele Bonadeo ("the Bonadeo declaration") and argued that one of skill in the art would not have combined the features of the invention as a matter of routine optimization.  Instead of routine, the applicants continued, the claimed formulations were obtained after a sequence of experiments, each of which built upon the others like building blocks.  If the experimental sequence had varied, the applicants alleged that they would have obtained a different formulation.  The Bonadeo declaration explained that the first two parameters studied were palonosetron concentration and pH.  None of the studies described in this declaration, however, occurred at a pH other than 5.0, which makes sense because palonosetron was described as extremely stable at this pH.

    Considering that the '219 patent ultimately issued, the applicants were apparently successful in overcoming these obviousness rejections.  In other words, the applicants convinced the examiner that a person of ordinary skill in the art would not have found it obvious to combine the teachings in the prior art to derived the claimed inventions.  The positions taken by the applicant, however, were utilized by the petitioner, Accord, to allege that a person of ordinary skill in the art would not have, for example, found the specification enabling.  This highlights the problem that PGRs pose for patent applicants.  Before such procedure, arguments could be made without much fear that they would be coopted by the Office for making alternative rejections.  And, if the Office did, there would still an opportunity to provide a response or amend the claims.  Even if such arguments were made in district court litigation, the patent would at least enjoy a presumption of validity.  Now, all applicants must take extreme caution in making any arguments, because anything said can (and probably will) be used against them at the PTAB.

    What follows is an identification of the 35 U.S.C. § 112 arguments made by Accord.  Considering that the patent owner has not yet filed any response, and the PTAB has not weighed in, no position is taken here as to the merits of these arguments.

    Written Description – Stability

    Accord first alleged that the '219 was unpatentable for failing to provide an adequate description of the claimed subject matter being stable at 18 or 24 months when stored at room temperature, as required by 35 U.S.C. § 112(a).  Specifically, the petition asserted that the specification does not show that the inventors were in possession of any formulation that would have achieved the stability limitations of the claims.  Instead, the argument went, the patent contained general statements that it is possible to increase the stability of the formulations, but did not provide any examples with stability beyond a couple of weeks.  Accord included a declaration from Dr. Arnold J. Repta, which explained how a person of ordinary skill in the art would have understood the teaching of the specification.  However, Dr. Repta did not include any additional testing of the formulations taught in the application in his declaration.

    Enablement

    The second assertion made in the petition was that the '219 patent does not enable a pH range for the claimed formation outside of about 4.0 to 6.0, and therefore it is not enabled as required by 35 U.S.C. § 112(a).  This is because, according the petition, the only relevant formulation in the specification was disclosed as having a pH of 5.0±0.5.  Moreover, the specification was alleged to claim that palonosetron is most stable at pH 5.0.  Accord also cited to the Bonadeo Declaration, which was submitted during prosecution by the applicants, which alleged claimed that palonosetron formulations containing mannitol or EDTA required a pH of 4-6.  Therefore, according to the petition, because the challenged claims do not recite any pH limitations, they were broader than the teaching of the specification.

    "Regards as the Invention"

    35 U.S.C § 112(b) requires that a patent "conclude with one or more claims particularly pointing out and distinctly claiming the subject matter which the inventor or a joint inventor regards as the invention."  Accord challenged the claims of the '219 patent as not including the invention as regarded by the inventors.  Specifically, the petition alleges that the Bonadeo Declaration made clear that palonosetron was extremely stable at a pH of 5.0, and that there was no hint that a pH outside of the range of about 4.0 or 6.0 would be suitable.  A similar argument was made about the language found in the specification.  The petitioner concluded by pointing out that, even though the inventors believed that the inventive formation should be in a range of 4.0 to 6.0, such a limitation was not included in the claims.

    Written Description – pH Range

    Finally, Accord made a similar argument when alleging that the specification did not support claims that did not include a pH range of 4.0 to 6.0.  Citing to the Gentry Gallery, Inc. v. Berkline Corp., 134 F.3d 1473 (Fed. Cir. 1998), line of cases, the petitioner alleged that the pH range was an essential or critical feature which was omitted from that claims.  In other words, Accord alleged that the broad claims without a pH limitation were invalid because the "entirety of the specification" demonstrates that the invention was of much narrower scope.

    Of course, similar to an IPR, the patent owner now has a chance to submit a preliminary response to the petition.  The standard used for instituting a PGR differs from that required for an IPR.  Instead of the "reasonable likelihood" standard, a PGR will only be instituted when it is more likely than not that at least one of the claims challenged is unpatentable.  In essence, this should be a slightly more stringent standard, because with both positions being equally likely, an IPR petition would have a reasonable likelihood of demonstrating claims as unpatentable, but a PGR petition would not be more likely than not to demonstrate unpatentable claims.  However, it remains to be seen if less PGRs are instituted than IPRs.  We will continue to monitor PGR2014-00010, and provide updates as warranted.