• By Kevin E. Noonan —

    One of the hallmarks of the current coronavirus pandemic, caused by SARS-Cov-2, is its zoonotic origins.  The virus is thought to have risen in bats and "jumped" to humans, and despite conspiracy theories of intentional human engineering (reminiscent of Soviet-backed propaganda in Africa in the 1980's that HIV/AIDS was produced in a laboratory in the U.S. as a way to reduce the African population) such transfer of disease-causing microorganisms from animals to humans is not so rare.  Influenza, for example, is a virus that infects humans, fowl (chickens and ducks) and pigs, and due to the segmented nature of the influenza virus RNA genome passage through these species (with concomitant "shuffling" of genes for variants of the neuraminidase and hemagglutinin) results each year or so in a virus to which the human immune system is naïve, even for previously infected individuals.

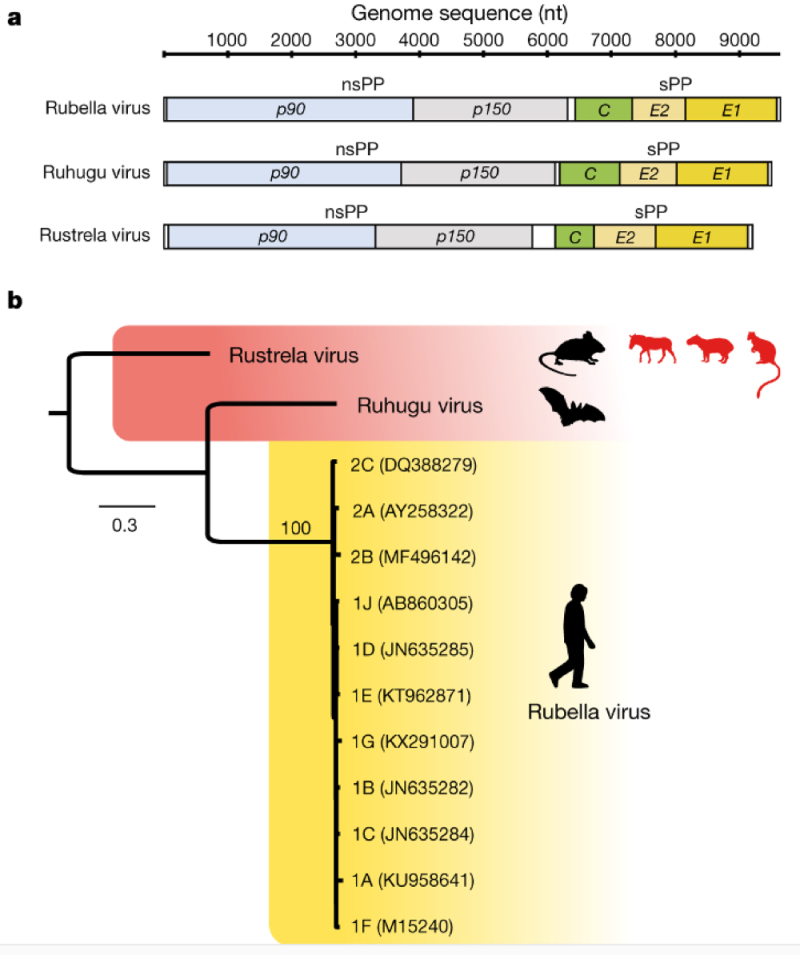
    Recently, an international* team of researchers reported in Nature their finding of viruses related to the Rubella virus (RuV) in a paper entitled "Relatives of rubella virus in diverse mammals," Nature 586: 424-28 (October 7, 2020).  One, named the ruhugu virus (RuhV), was found in "apparently healthy cyclops leaf-nosed bats (Hipposideros cyclops).  The other, a less closely related virus named the rustrela virus (RusV), is endemic in wild yellow-necked field mice (Apodemus flavicollis) in Germany.  This latter virus was also found by these researchers to cause acute encephalitis in placental and marsupial mammals (a donkey, Equus asinus; a capybara, Hydrochoeris hydrochaeris; and a kangaroo, Dendrolagus bennettianus) at a German zoo.

    Genetic analyses showed clear homology of the genetic structure of these viruses, and the evolutionary relationships between them and the rubella virus that causes disease in humans.  The E1 protein of the RuV virus is described as "a receptor-binding, class-II fusion protein, contain[ing] an immune-reactive region (amino acid residue positions 202–283) with immunodominant T cell epitopes and four linear, neutralizing B cell epitopes (NT1–NT4)."  These researchers report that the E1 protein of the RusV and RuhV viruses share this structure and epitope mapping confirms the activity of neutralizing antibodies that block trimerization of this protein, which is necessary for virus fusion with host cell membrane.  Consistent with evolutionary divergence between RuV/RusV and RuhV are two non-conservative amino acid differences in one of the E1 protein fusion loops, disclosed as P134A and T135A.  Nevertheless, these researchers report "regions of marked conservation and stabilizing selection" in certain regions of the viral genomes, consistent with their putative evolutionary relationships.

    While the authors are appropriately cautious in stating that "[t]he implication that RuhV or RusV are zoonotic agents is currently speculative," they also note that "bats and rodents possess biological attributes that predispose them to hosting many zoonotic viruses" and that RusV can infect both placental and marsupial mammals and cause neurological disease that resemble the severe encephalitis cause in infected human infants.

    These observations are also relevant, the authors note, to efforts to eliminate rubella in infection (particularly congenital infections) in humans in the near future.  The possibility of future zoonotic transfer of these related viral species is a prospect that could influence the likelihood of realistically removing this class of viruses from human pathogens, and may in the interim provide a pathway for developing animal models for infection with these viruses.

    Image

    * University of Wisconsin-Madison, WI; Uniformed Services University of the Health Sciences, Bethesda, MD; Leidos, Reston, VA; Fort Detrick, Frederick, MD; Friedrich-Loeffler-Institut, Greifswald-Insel Riems, DE; Food Safety and Fisheries, Rostock, DE

  • By James Lovsin —

    Supreme Court SealToday, the Supreme Court granted petitions for a writ of certiorari to review the Federal Circuit's decision in Arthrex, Inc. v. Smith & Nephew, Inc., in which the court of appeals held how administrative patent judges were appointed to the Patent Trial and Appeal Board ("PTAB") violated the Appointments Clause of the Constitution (Art. II, sec. 2, cl. 2).  To prospectively remedy that violation, the court severed a portion of the America Invents Act restricting the way APJs can be removed, and vacated and remanded the PTAB's judgment for a new hearing.  The Federal Circuit denied rehearing, with four judges dissenting.

    The Supreme Court has granted certiorari on two questions that the government argued were common questions across the petitions filed by the petitioner (Smith & Nephew), patent owner (Arthrex), and government:

    1.  Whether, for purposes of the Appointments Clause, U.S. Const. Art. II, § 2, Cl. 2, administrative patent judges of the U.S. Patent and Trademark Office are principal officers who must be appointed by the President with the Senate's advice and consent, or "inferior Officers" whose appointment Congress has permissibly vested in a department head.

    2.  Whether, if administrative patent judges are principal officers, the court of appeals properly cured any Appointments Clause defect in the current statutory scheme prospectively by severing the application of 5 U.S.C. 7513(a) to those judges.

    Notably, the Supreme Court did not grant cert. on the government's third common question, which involved waiver/forfeiture of an appointments clause challenge.  Of course, the Court would not have granted cert. if the Justices thought the appointments issue was waived.

    Merits briefing from Smith & Nephew, Arthrex, and the government will begin on the two granted questions.  There will likely be many briefs from amici.  The Supreme Court has scheduled one hour of oral argument divided between Smith & Nephew, Arthrex, and the government.  A decision in the case is likely expected no later than June 2021.

    Arthrex presents yet another constitutional challenge to the PTAB.  Two years ago, the Supreme Court upheld the constitutionality of IPR proceedings in Oil States v. Greene, holding that IPRs did not violate Article III or the Seventh Amendment.  Justice Gorsuch and Chief Justice Roberts dissented in Oil States and it will be interesting to see if they are persuaded by the appointment clause challenge in this case.  The Justices' views in Supreme Court cases involving judicial review of PTAB rulings (Thryv, SAS, and Cuozzo) may also be worth considering here.  For example, Justice Alito, Justice Sotomayor, and Justice Gorsuch have supported strong judicial review in these cases.

    The Supreme Court's decision in Arthrex could affect pending and future PTAB proceedings.  There are, for example, around 40 PTAB cases in which the Federal Circuit remanded to the PTAB for a new hearing in light of Arthrex.  The government filed a consolidated cert. petition on these cases.  And for its part, the PTO has stayed Arthrex remands until the Supreme Court resolves Arthrex.  These cases will likely sit until after the Supreme Court's decision in Arthrex.

    If the Supreme Court answers the first granted question, yes, APJs are principles officers, the Court will likely address the second granted question, the remedy for the violation.  Some Federal Circuit judges and commentators have argued that the Federal Circuit's remands were unnecessary because judicial severance of the America Invents Act is both prospective and retroactive.  Bedgear v. Fredman Bros (Judges Dyk and Newman concurring opinion); A. Michaels, Arthrex Remands: Treat or Trick? Patently-O (July 19, 2020).  The retroactivity issue was not significantly addressed, if at all, at the cert. stage.  It will be interesting to see if retroactivity is briefed and considered by the Court in the context of the second question.

    If the Supreme Court concludes that Congress needs to remedy how PTAB judges are appointed with legislation, the Court could stay the effect of its ruling.  This would give Congress time to enact such legislation.  The House Subcommittee on Courts, Intellectual Property, and the Internet held a hearing on the appointment of PTAB judges in November 2019.  However, the appointment of PTAB judges may be a low priority for Congress in 2021.

    MBHB will monitor developments in Arthrex.  In the meantime, litigants should carefully consider the potential impact of the Supreme Court's decision in Arthrex on their PTAB proceedings.

  • By Kevin E. Noonan —

    Court of Appeals - 3d Cir. SealThe Federal Trade Commission carried out an (in)famous crusade against reverse payment (more provocatively, "pay for delay") settlements in ANDA litigation for almost a decade before eventually having the Supreme Court see things their way (to some extent) in FTC v Actavis.  The Commission has not lost its enthusiasm for such interventions in drug patenting matters but without (so far) overwhelming success.  This pattern is illustrated in the Third Circuit's recent decision in Federal Trade Commission v. AbbVie Inc.

    The Commission's interest arose over Androgel, which the opinion characterizes as a "blockbuster testosterone replacement therapy that generated billions of dollars in sales," the latter characteristic no doubt playing a large part in attracting the FTC's investigation.  The Commission brought suit against AbbVie and related companies under 15 U.S.C. § 53(b) (Section 13(b) of the Federal Trade Commission Act), which provides FTC jurisdiction over false advertisements with provisions for injunctions and restraining orders:

    Power of Commission; jurisdiction of courts

    Whenever the Commission has reason to believe—

    (1) that any person, partnership, or corporation is engaged in, or is about to engage in, the dissemination or the causing of the dissemination of any advertisement in violation of section 52 of this title, and

    (2) that the enjoining thereof pending the issuance of a complaint by the Commission under section 45 of this title, and until such complaint is dismissed by the Commission or set aside by the court on review, or the order of the Commission to cease and desist made thereon has become final within the meaning of section 45 of this title, would be to the interest of the public,

    the Commission by any of its attorneys designated by it for such purpose may bring suit in a district court of the United States or in the United States court of any Territory, to enjoin the dissemination or the causing of the dissemination of such advertisement. Upon proper showing a temporary injunction or restraining order shall be granted without bond. Any suit may be brought where such person, partnership, or corporation resides or transacts business, or wherever venue is proper under section 1391 of title 28. In addition, the court may, if the court determines that the interests of justice require that any other person, partnership, or corporation should be a party in such suit, cause such other person, partnership, or corporation to be added as a party without regard to whether venue is otherwise proper in the district in which the suit is brought. In any suit under this section, process may be served on any person, partnership, or corporation wherever it may be found.

    The basis of the allegations were that defendants filed "sham" patent infringement suits (which seems to be the allegation the FTC makes against any patent infringement or ANDA lawsuit brought by a drug company) against generic drug companies, followed by entering into an "anticompetitive reverse-payment agreement" with one such company (Teva Pharmaceuticals).  (Indeed, Perrigo Co., one of the generic competitors, made that allegation before the District Court.)

    The various parties settled, concomitant with payments from Abbott/AbbVie for "avoided litigation expenses" (a category sanctioned by the Supreme Court's Actavis opinion) and an agreement of a date certain, prior to patent expiration, when the generic competitor could market its Androgel generic drug.  In Teva's case, the Androgel settlement was paired with settlement over another generic drug (brand name TriCor) which permitted Teva to take advantage of a 180-day exclusivity period as first filer, with Abbott being the supplier and being entitled to "the costs of production, an additional percentage of that cost, and a royalty."  (This agreement formed the basis for FTC's allegation of an illegal reverse payment.)

    FTC's complaint recited two counts:

    • Count 1 (Monopolization): that AbbVie and Besins willfully maintained a monopoly through a course of anticompetitive conduct, including sham patent litigation against Teva and Perrigo.

    • Count 2 (Restraint of Trade): that AbbVie restrained trade by entering into an anticompetitive reverse payment agreement with Teva.

    As explained by the Third Circuit, the District Court dismissed FTC's complaint as to the reverse payment liability theory but permitted the case to move forward on the sham patent allegation.  Thereafter, the District Court held for FTC on both prongs of the sham litigation theory (i.e., that AbbVie had monopoly power in the relevant market and the sham litigation was willfully acquired or maintained by that litigation) and entered judgment requiring AbbVie (and co-defendant Besins) to disgorge $448 million in "ill-gotten profits" but declined to enter an injunction.  This appeal (by FTC, AbbVie and Besins) followed.

    The Third Circuit held that the District Court erred in rejecting the reverse-payment theory and concluding that litigation against Teva was a sham.  The District Court did not err, according to the Third Circuit, in concluding that litigation against another generic company, Perrigo, was a sham, that defendants had monopoly power in the relevant market, or in denying FTC's request for an injunction.  The Court also held that the FTC had overstepped its authority in seeking disgorgement, because Section 13(b) of the FTC Act contained no provision authorizing this remedy, thus forestalling any remedy against defendants unless the district court found antitrust liability on the reverse payment theory on remand.

    With regard to the reverse payment allegation, the Court set forth its understanding of the fact pattern supporting such an allegation:

    "Company A sues Company B for patent infringement.  The two companies settle under terms that require (1) Company B, the claimed infringer, not to produce the patented product until the patent's term expires, and (2) Company A, the patentee, to pay B many millions of dollars" citing Actavis.

    The Court's basis of finding anticompetitive behavior under these circumstances was that "Company B might have prevailed by proving Company A's patent invalid.  Even if the patent were valid, Company B might prevail by showing it did not infringe.  In either case, generic drugs would have entered the market before Company A's patent was set to expire, and consumers would have benefited from lower drug prices."

    The lack of a bright line set forth in Actavis by the Supreme Court (which also involved Androgel) was the basis for the Court's reversing and remanding the District Court's grant of defendants' motion to dismiss.  And the opinion asserted that the Third Circuit's decision here was consistent with other cases decide by the Court, including King Drug Co. of Florence, Inc. v. SmithKline Beecham Corp., 791 F.3d 388 (3d Cir. 2015) (no authorized generic), In re Wellbutrin XL Antitrust Litig. Indirect Purchaser Class, 868 F.3d 132 (3d Cir. 2017), and In re Lipitor Antitrust Litig., 868 F.3d 231 (3d Cir. 2017) (settling a related claim for much less than it was objectively worth on one drug and a "no authorized generic" agreement on another).  From these cases the Third Circuit recognized two principles set forth in this opinion:

    • First, a reverse payment's legality depends mainly on its economic substance, not its form.

    • The second principle . . . is that the law of pleading applies to reverse-payment theories. [Specifically, i]f a plaintiff plausibly alleges that an agreement's anticompetitive effects outweigh its procompetitive virtues, the district court must accept that allegation and allow the plaintiff to take discovery.

    The Third Circuit's application of these principles led it to reverse the District Court's dismissal of FTC's reverse payment theory in their complaint, because "FTC plausibly alleged an anticompetitive reverse payment."  Although filing the putatively sham lawsuits engaged the 30-month delay in FDA approval provided by the Hatch-Waxman Act, FTC alleged the AbbVie and Besins were fearful that Teva would prevail and thus "turned to other ways to preserve their monopoly."  Under the terms of this agreement, the Third Circuit held that the "payment" to Teva (specifically, being permitted to bring its TriCor generic on the market) was "plausibly 'large'" due to inter alia its 180-day exclusivity as first filer.  (As it turns out, according to the opinion, Teva's actual sales were much higher than predicted and "far exceed[ed] the litigation costs that [any of the parties] saved by settling"; this analysis adds not a little hindsight to a determination that is usually grounded in the parties' expectations in determining whether there was antitrust intention in the agreement).  These agreements were also "plausibly 'unjustified," in the Third Circuit's view, on the basis that FTC alleged the TriCor arrangement "'cannot be explained as an independent business deal from Abbott's perspective'" (presuming that FTC has any basis for determining Abbott's perspective).  Thereafter the opinion cites a litany of such putatively "highly unusual" aspects of the agreements.  Finally, the opinion asserts that "it is plausible that the anticompetitive effects of AbbVie's settlement with Teva outweighed any procompetitive virtues of the TriCor deal" (which is a calculation FTC is entitled to assert as part of an antitrust allegation), citing similar considerations in its King Drug opinion.  The Third Circuit critiqued the District Court's grounds for finding that FTC had not plead an antitrust violation, finding it wanting at least as being inconsistent with King Drug in limiting its determination to the absence of a large cash payment from AbbVie/Besins to Teva.  And the Third Circuit found the District Court's lack of consideration of the supply arrangement between Teva and AbbVie/Besins for antitrust purposes to be inconsistent with its Lipitor decision, if only because a proper determination of such an agreement's compliance with antitrust principles requires "'factual assessments, economic calculations, and expert analysis that are inappropriate at the pleading stage.'"  The opinion was also critical of the District Court finding that the agreement was procompetitive as a matter of law, but the Third Circuit was careful not to come to any decision on whether the District Court had "correctly concluded the TriCor deal was procompetitive," leaving the question of where the balance will be struck on pro- and anticompetitive effects of these agreements to be decided on remand.

    The basis of the Third Circuit's decisions that AbbVie's assertion of its patents in ANDA litigation was (Perrigo) or was not (Teva) a sham were grounded in the Noerr-Pennington doctrine (i.e., that antitrust liability was not raised when a party petitioned the government for redress), recognizing the standard for exceptions to this immunity.  Specifically, the opinion noted that the Supreme Court has recognized two prongs for determining an exception:

    First, the lawsuit must be objectively baseless in the sense that no reasonable litigant could realistically expect success on the merits.  If an objective litigant could conclude that the suit is reasonably calculated to elicit a favorable outcome, the suit is immunized under Noerr, and an antitrust claim premised on the sham exception must fail.  Only if challenged litigation is objectively meritless may a court examine the litigant's subjective motivation.  Under this second part of our definition of sham, the court should focus on whether the baseless lawsuit conceals an attempt to interfere directly with the business relationships of a competitor through the use of the governmental process—as opposed to the outcome of that process—as an anticompetitive weapon.  This two-tiered process requires the plaintiff to disprove the challenged lawsuit's legal viability before the court will entertain evidence of the suit's economic viability (emphasis in opinion.)

    The Court noted that the statutory nature of an ANDA litigation mitigates against a determination that a litigation is a sham (although the 30-month stay mitigates in a generic company's favor as being "a collateral injury" as an "anticompetitive weapon" an NDA holder can use regardless of whether there is a good faith basis that the generic drug infringes any Orange Book-listed patent).  Regarding the District Court's basis for deciding the Teva litigation was not objectively baseless, the opinion found that it was not objectively baseless for AbbVie to assert infringement under the doctrine of equivalents based on the amendment's "tangential relationship" to prosecution history estoppel with regard to the penetration enhancers specified by AbbVie/Besins' patents and Teva's accused infringing formulation.

    On the other hand, this tangential relationship rationale did not convince the Third Circuit in the Perrigo litigation, because there "[n]o reasonable litigant in AbbVie and Besins's position would believe it had a chance of winning on these arguments."  The opinion rejected AbbVie/Besins's argument that the voluntary nature of the amendment avoids prosecution history estoppel, correctly noting that the Federal Circuit has rejected the argument that voluntary amendments have any less effect in supporting the estoppel than involuntary ones (i.e., ones made for reasons of patentability), citing Festo Corp. v. Shoketsu Kinzoku Kogyo Kabushiki Co., 344 F.3d 1359 (Fed. Cir. 2003).  Nor was it effective for AbbVie/Besins to argue that such amendments were made to expedite prosecution, citing Biogen, Inc. v. Berlex Labs., Inc., 318 F.3d 1132 (Fed. Cir. 2003) (and providing a cautionary tale for prosecutors who make such statements routinely, at least with regard to the estoppel-producing effects of such amendments).

    Having satisfied the objectively baseless prong of the sham litigation test, the opinion then reviewed whether AbbVie/Besins's suit met the subjective motivation prong.  In this regard the Third Circuit reviewed with approval the logical syllogism used by the District Court:

    (1) A lawsuit is objectively baseless if "no reasonable litigant could realistically expect success on the merits," . . . (2) and a litigant who files an objectively baseless lawsuit must have had some subjective motivation for suing; (3) but because the lawsuit was objectively baseless, the litigant's subjective motivation could not have been success on the merits, unless the litigant was unreasonable; (4) thus, a reasonable litigant's subjective motivation for filing an objectively baseless lawsuit must be something besides success on the merits.

    As applied to this case, the District Court:

    [F]irst held that AbbVie and Besins's lawsuits were objectively baseless.  It then reasoned that because AbbVie and Besins's decisionmakers were all very experienced patent attorneys who had reviewed Perrigo's paragraph IV notices and consulted outside counsel, they knew the lawsuits were baseless.  Finally, it reasoned that because the decisionmakers knew the lawsuits were baseless, they must have been motivated by something other than success on the merits.

    (although there appears to be a circularity in this reasoning that approaches the tautological).  The opinion also credited the Hatch-Waxman-mandated 30-month stay as being sufficient motivation to satisfy (under the right circumstances) the subjectively baseless prong of the test.

    The discussion in the opinion of monopoly power was unremarkable, in view of the interactions between the regulatory regime and the Orange Book patents listed in support of AbbVie/Besins's NDA ("a generic drug has significant capital, technical, regulatory, and legal barriers to overcome").  Also relevant was AbbVie's commercial behavior regarding price in the marketplace, specifically the ability to raise prices consistently despite the existence of cheaper (albeit somewhat less desirable) alternatives, i.e., injectable formulations.

    It was the Third Circuit's decision on remedies that was (for now) most significant, absolving (for now) AbbVie from disgorging almost $500 million as a penalty.  The error, according to the Third Circuit, was that the section of the FTC Act relied upon by the Commission in bringing suit does not give FTC the authority to order such disgorgement.  The Court based its decision on a review of the legal authority underlying FTC's enforcement capabilities and finding no indication that Congress gave FTC the authority under Section 13(b) it claims in this case.  And the wording of Section 13(b) was unambiguous to the Court in limiting FTC to seeking injunctive relief only in cases where an antitrust violation is imminent or ongoing, neither circumstance arising here.  While the opinion conceded that courts in sister circuits have permitted FTC to pursue and obtain disgorgement under Section 13(b) this court founds such outcomes not to be supported by the plain meaning of the statute nor an understanding of the operation of the Section in the context of the statute as a whole.

    Finally, the Third Circuit affirmed the District Court's denial of the injunctions sought by FTC as not being an abuse of discretion.  The standard for granting FTC an injunction relied upon by the Court is that "there is 'cognizable danger of recurrent violation, something more than the mere possibility which serves to keep the case alive,'" citing United States v. W.T. Grant Co., 345 U.S. 629, 633 (1953).  Those circumstances did not exist here given the nature of the underlying litigation, according to the Third Circuit.

    While the case is remanded, giving FTC another opportunity to obtain an injunction, and proceedings on remand will include trial on whether Counts 1 and 2, one judgment that survived appeal was the District Court's finding that AbbVie/Besins were liable for monopolization under Count 1 for bringing a sham litigation against Perrigo.  What penalty FTC can successfully extract from these defendants will be determined on remand and could form the basis for a settlement, especially should the possibility of a successful disgorgement request on grounds other than Section 13(b) be made by FTC.

    Federal Trade Commission v. AbbVie Inc. (3d Cir. 2020)
    Panel: Circuit Judges Hardiman, Porter, and Phipps
    Opinion by Circuit Judge Hardiman

  • By Michael Borella —

    USPTO SealLast year, the U.S. Patent and Trademark Office (USPTO) issued a request for comments (RFC) on patenting artificial intelligence (AI) based inventions.  Topics of the RFC included AI's impact on inventorship and ownership, patent eligibility, disclosure, prior art, and the level of ordinary skill in the art.  The USPTO received 99 comments from a variety of stakeholders.  In parallel to that effort, the USPTO also released a second RFC related to the impact of AI on copyright, trademark, database protections, and trade secret law.  Stakeholders provided a similar number of comments.

    With this feedback in hand, the USPTO collated it into a 56-page report.  This article focuses on issues related to the drafting and prosecution of AI-related patent applications.  Readers are encouraged to read the rest of the report to obtain the USPTO's summary of the remaining issues.

    The report begins with a high level overview that can be expressed as three main points:

    1.  Most commenters agreed that AI-based inventions are a subset of computer-implemented software inventions, and thus the current framework for examining such inventions is suitable for examining AI-based inventions.  Still, some commenters exhibited concern that it may be difficult to meet the requirements of 35 U.S.C. § 112(a) for certain AI-based inventions.

    2.  Most commenters agreed that AI may lead to changes in how the standard of a person of ordinary skill in the art (POSITA) is interpreted.

    3.  There were some concerns that AI would lead to a proliferation of prior art making it difficult for examiners to conduct searches for relevant prior art.

    A brief list of issues of note follows.

    Definitions

    Interestingly, there was no clear consensus on what constitutes an AI-based invention.  Of the definitions provided, the one that seems to be the most helpful (in my subjective opinion) states that:

    AI inventions can be categorized (in no particular order) as follows:

    (a) inventions that embody an advance in the field of AI (e.g., a new neural network structure of an improved machine learning (ML) model or algorithm)

    (b) inventions that apply AI (to a field other than AI)

    (c) inventions that may be produced by AI itself.

    Patent Eligibility

    As noted, the consensus is that, as a special form of computer-implemented invention, AI-based inventions can be fairly evaluated under the current patent-eligibility of 35 U.S.C. § 101 as interpreted by the courts.  Some commenters indicated that AI-based inventions could be characterized as falling into the abstract idea subject matter exception, as a method of organizing human activity, a mental process, or a mathematical concept.  But to the extent that these inventions provide technological improvements and "amount to significantly more than the abstract idea," they would be found to pass through the § 101 filter.

    Indeed, this approach is consistent with the Supreme Court's refusal to draw a line between what is and is not eligible.  Each invention is to be considered on its own merits, and there are no "magic words" or drafting techniques that can be used to guarantee a claim is eligible.

    Written Description and Enablement

    While many commenters did not believe that AI-based inventions had any special written description requirements, one commenter stated that "AI inventions can be difficult to fully disclose because even though the input and output may be known by the inventor, the logic in between is in some respects unknown."  This is true, in the sense that a learning model can be viewed as a "black box" that receives input (e.g., words or images) and produces output (e.g., a classification).  One can view the constituent elements that make up the model (e.g., nodes, connections, and weights), but these elements can number in the millions.  As a consequence, they cannot be used to concisely characterize the invention, nor should they as the specific values of weights can be different each time the model is trained.

    Regarding enablement, the USPTO notes that "the amount of guidance or direction needed in the specification to enable the invention is inversely related to the amount of knowledge in the state of the art, as well as the predictability in the art."  But there is no clear understanding of the predictability of AI systems.  Some contain an inherent amount of intentional randomness that allow these systems to produce superior results when compared to more deterministic systems.  Others may be predictable if their training data and training technique is fixed.

    One possible way of addressing both of these concerns is to allow AI models that involve machine learning to meet the written description and enablement requirements with proper specification of how the models can be trained.  If, from the specification, a POSITA can glean enough knowledge to reasonably reproduce the training technique with their own data, the invention is likely to be well-described and enabled.

    Level of Ordinary Skill in the Art

    While the ubiquity of AI may impact the level of ordinary skill in the art, commenters seems to agree that the present legal framework is "adequate to determine the impact of AI-based tools in a given field."  As the level of skill in the art generally increases over time, use of AI-based tools would be just another way for the level of skill to rise, albeit at a potentially faster rate than what we have seen in the past.

    Prior Art Considerations

    While most commenters believed that there are no specific concerns regarding prior art and AI-based inventions, some indicated that AI itself may be used to generate massive amounts of prior art that would be challenging to thoroughly search.  Others pointed out that prior art unique to AI may eventually exist as AI systems evolve toward more general intelligence.  A key to these considerations would be proper examiner training.

    Data Protection

    A critical component of AI goes beyond the algorithms and models, and is embodied in the data sets that the algorithms use to train the models.  Currently, such data sets have relatively weak protection (mostly in the realm of trade secret law).  Some commenters would like to have provisions that allow incumbent companies to maintain proprietary rights over data sets that they have collected, while still allowing newcomers to use these data sets to train their own models.

    Examination

    There were suggestions that the USPTO coordinate with the world's other patent offices when considering its approach to AI-based inventions.  Of note was that the Japan Patent Office and Korean Intellectual Property Office have established specific and dedicated AI examination units.  Further, the Intellectual Property Office of Singapore has "reportedly created an expedited examination path for AI technologies."

  • CalendarOctober 13, 2020 – "An In-House Counsel’s Guide to Product Clearance: Best Practices in Freedoms-to-Operate, Landscaping and Opinions of Counsel" (McDonnell Boehnen Hulbert & Berghoff LLP) – 10:00 am to 11:15 am (CT)

    October 13, 2020 – "Preparing & Prosecuting Patents That Stand Up To Challenge" (LexisNexis IP and IPWatchdog) – 12:00 pm (ET)

    October 15, 2020 – "Myths of Litigated Patents" (LexisNexis IP and IPWatchdog) – 12:00 pm (ET)

    November 9-10, 2020 – Summit on Life Sciences IP Due Diligence (American Conference Institute) – virtual conference

  • ACIAmerican Conference Institute (ACI) will be holding its 4th Annual Summit on Life Sciences IP Due Diligence on November 9-10 as a VIRTUAL conference.

    The conference will begin with a Pre-Conference Primer at 8:00 am Monday, November 9th, entitled "Life Science Due Diligence 101 – Preparing Your Due Diligence Checklist and Managing the Process from Soup to Nuts."

    The conference will offer presentations on:

    • Evaluating COVID-19's Impact on Biopharma Deal Making and Valuation of IP Assets
    •
    Conducting Due Diligence in the Age of "Six Feet Apart": Strategies for Performing a Robust Virtual/Remote IP Due Diligence Review
    •
    Unique IP Due Diligence Considerations for Medical Devices, MedTech, and Artificial Intelligence
    •
    Mitigating International Legal Risk: IP Due Diligence Considerations for Global Deals
     
    Assessing the Intricacies of Deal-Related IP Valuation
    •
    An Interactive Interview with a Private Equity Expert and Venture Capitalist on the Ins and Outs of Securing Funding for Life Sciences Companies
    •
    Understanding the Impact of New Developments at FDA on the IP Due Diligence Review Process
    •
    "War Stories" on Due Diligence Gone Wrong: When Defective IP Due Diligence Wrecks the Deal
    •
    Interactive Ethics Lab on Overcoming Common Challenges and Ethical Dilemmas in Life Sciences IP Due Diligence

    The conference will also offer an interactive case study focusing on "Avoiding Missteps When Conducting IP Due Diligence in Connection with Distressed Companies and Distressed Assets" and "Uncovering Red Flags in IP Ownership and Solving Conundrums Caused by Government Funding Initiatives on IP Rights and Evaluation."

    There will be a Post-Conference Workshop entitled "Freedom to Operate BootCamp: A Comprehensive Tactical Guide to Mastering the FTO Search" starting at 2:15 pm on November 10th.

    A complete brochure for this conference, including an agenda, detailed descriptions of conference sessions, list of speakers, and registration form can be obtained here.

    The registration fee is $1,795 (registered and paid by November 8th).  The Pre-Conference Primer is priced at $195 and the Post-Conference Workshop is $295; the registration fee for the combination of Virtual conference and workshop is $2,085.  Patent Docs readers are entitled to a 10% discount off of registration using discount code D10-657-657JX01.  Those interested in registering for the conference can do so here, by e-mailing CustomerService@AmericanConference.com, or by calling 1-888-224-2480.

    Patent Docs is a media partner of ACI's 4th Annual Summit on Life Sciences IP Due Diligence.

  • LexisNexisLexisNexis IP and IPWatchdog and will be offering a webinar entitled "Myths of Litigated Patents" on October 15, 2020 at 12:00 pm (ET).  Dr. Sean Tu, Professor, WVU College of Law; Megan McLoughlin of LexisNexis® IP; and Gene Quinn of IPWatchdog will discuss whether patent examiners who issue litigated patents have common characteristics?  While intuition would argue that those examiners who issue the most patents (approximately one patent every three business days) would exhibit a higher litigation rate, surprisingly, two studies by Professor Sean Tu suggest that this is wrong.  The panel will address the following issues:

    • Which types of examiners tend to issue patents that later undergo litigation
    • Whether examiners who are "rubber stamping" patents issue litigated patents at a disproportionately higher rate
    • Whether examiners with less experience are issuing more litigated patents
     The issues at stake during prosecution for different types of examiners
    • How this knowledge can impact your prosecution practice

    There is no registration fee for this webinar.  However, those interested in registering for the webinar, should do so here.

  • MBHB Logo 2McDonnell Boehnen Hulbert & Berghoff LLP will be offering a live webinar entitled "An In-House Counsel’s Guide to Product Clearance: Best Practices in Freedoms-to-Operate, Landscaping and Opinions of Counsel" on October 13, 2020 from 10:00 am to 11:15 am (CT).  In this presentation, MBHB attorneys Grantland Drutchas and Jason Kray will address the steps in-house counsel need to take to protect their companies from assertions by competitors and NPE's and the goals and expectations they should you have for their outside counsel in conducting clearances and drafting opinions of counsel,  In addition, the panel will explore the following topics:

    • What steps you should take to explore the IP landscape when developing or on-boarding new products.
    • How to get the most from your freedom-to-operate opinions.
    • How to ensure that a search addresses the full scope of potential IP in all relevant countries.
    • What you need to do to make sure that your outside counsel's opinion will pass muster in litigation.
    • What you can do to evaluate NPE challenges, both in the U.S. and in the growing international NPE market.
    • How to evaluate risks of competitors re-crafting IP to cover your products.
    • What you should do to assess clearance risks for products from suppliers.

    While there is no fee to participate, attendees must register in advance.  Those wishing to register can do so here.  CLE credit is pending for the states of California, Illinois, New Jersey, New York, North Carolina, and Virginia.

  • LexisNexisLexisNexis IP and IPWatchdog and will be offering a webinar entitled "Preparing & Prosecuting Patents That Stand Up To Challenge" on October 13, 2020 at 12:00 pm (ET).  Todd Van Thomme of Nyemaster Goode P.C., Kristin Murphy of Honigman LLP, and Bernie Tomsa of Brooks Kushman P.C. will discuss: (1) Pros and cons of having a Background (101 vs. 103); (2) Carefully considering claim terms and infringers; (3) Fixing problems before and after allowance; (4) When should you be willing to take a case to appeal? (5) Portfolio building techniques: Identifying valuable claims to add.

    Those interested in registering for the webinar can do so here.

  • What Quantum of Culpable Conduct Is Required for an ANDA Applicant to Induce Infringement?

    By Kevin E. Noonan —

    Federal Circuit SealThe back-and-forth, (almost) cat-and-mouse-like competition between branded innovator and generic drug makers sanctioned under the Hatch-Waxman Act has been on-going for over thirty years.  As part of this regime, Congress has provided a pathway for generic drug companies to obtain FDA approval for less than all the indications a branded drug has obtained, using a "carve out" strategy resulting in a so-called "skinny label."  This has raised the possibility of "off-label" use, where physicians prescribe the generic drug for an indication not approved for the generic drug but known to be clinically appropriate from the innovator's approval for the product.  The extent to which a generic drug company can use this strategy to avoid liability for inducing infringement was tested in the Federal Circuit's recent decision in GlaxoSmithKline LLC v. Teva Pharmaceuticals USA, Inc.

    The matter arose in litigation over GSK's Coreg® product (carvedilol) for treatment of hypertension (the initial approved indication; U.S. Patent No. 4,503,067), congestive heart failure (CHF) (the subject of U.S. Patent No. 5,760,069) and left ventricular dysfunction following myocardial infarction (LVD-MI).  The '069 patent recites a method of treating CHF with a combination of carvedilol and "one or more of an angiotensin-converting enzyme ("ACE") inhibitor, a diuretic, and digoxin."

    Teva's ANDA was filed with a Paragraph III certification over the '067 patent and a Paragraph IV certification over the '069 patent.  The FDA tentatively approved Teva's generic product for "treatment of hypertension and heart failure" which Teva launched on expiration of the '067 patent.  Teva's label indicated that the product was approved treatment of LVD-MI and hypertension and announced that FDA had given its product an "AB rating" (which the opinion explained "allow[s] users to determine quickly whether the Agency has evaluated a particular approved product as therapeutically equivalent to other pharmaceutically equivalent products").  Thereafter, FDA required Teva to amend its label to be identical to the GSK label for Coreg®, which introduced treatment of heart failure into the approved treatments recited in Teva's label.

    GSK filed for reissue of the '069 patent which was duly granted by the U.S. Patent and Trademark Office as Reissue Patent No. RE40,000; claim 1 is representative of the invention as claimed in the '000 reissue patent:

    1.  A method of decreasing mortality caused by congestive heart failure in a patient in need thereof which comprises administering a therapeutically acceptable amount of carvedilol in conjunction with one or more other therapeutic agents, said agents being selected from the group consisting of an angiotensin converting enzyme inhibitor (ACE), a diuretic, and digoxin,
        wherein the administering comprises administering to said patient daily maintenance dosages for a maintenance period to decrease a risk of mortality caused by congestive heart failure, and said maintenance period is greater than six months.

    (Where the italicized portion of the claim represents the modifications introduced in prosecution of the reissue application.)

    GSK filed suit against Teva for inducement of infringement based on the Teva label, based on direct infringement by physicians prescribing the drug for the label indications.  Teva argued that it had "carved out" the indication for CHF pursuant to 21 U.S.C. § 355(j)(2)(A)(viii), resulting in a "skinny label" with regard to this indication.  Thereafter, FDA compelled Teva to amend its label to include that indication.  In addition, Teva argued that it could be liable for inducement only if GSK could show that Teva had "directly communicated with the direct infringers and 'caused' them to directly infringe the method in the '000 patent."  In a jury instruction the court informed the jury that circumstantial evidence could be used to satisfy this burden.

    The jury found that Teva induced infringement of the '000 reissue patent both before and after the label amendment (albeit infringing several claims after but not before that change).  The District Court granted Teva's motion for judgment as a matter of law (JMOL) on the basis that GSK had not "caused" physicians to prescribe their product for the infringing uses.  Because proof of such causation was required, according to the District Court, its absence precluded the jury from basing its decision on substantial evidence.  The Court relied on the "many sources of information available to prescribing physicians" other than Teva's label (including paradoxically GSK's label and promotion of its Coreg® product) in finding this evidentiary deficiency.  Also, the Court based its decision on physician testimony that their prescribing behavior relied on "guidelines and research, as well as their own experience" and not Teva's label.  "In sum," the Court said, "substantial evidence [did] not support the jury's finding on causation, and therefore [did] not support its verdict that Teva is liable for induced infringement, during both the skinny and full label periods."  This appeal followed.

    The Federal Circuit reversed, in an opinion by Judge Newman joined by Judge Moore; Chief Judge Prost provided a lengthy, comprehensive dissent.  The panel majority relied on the Supreme Court's decision in Global-Tech Appliances, Inc. v. SEB S.A., 563 U.S. 754 (2011), that copying is evidence of inducement, as applied in to generic drugs (which are required by regulatory statutes to "copy") in Eli Lilly & Co. v. Teva Parenteral Meds., Inc., 845 F.3d 1357, 1369 (Fed. Cir. 2017), and Sanofi v. Watson Labs. Inc., 875 F.3d 636, 645 (Fed. Cir. 2017) (as well as the earlier precedent of AstraZeneca LP v. Apotex, Inc., 633 F.3d 1042, 1060 (Fed. Cir. 2010), and Mentor H/S, Inc. v. Med. Device All., Inc., 244 F.3d 1365, 1379 (Fed. Cir. 2001)).  The majority also found compelling evidence from Teva's website regarding its product's AB rating with GSK's Coreg® product and other promotional content, as well as testimony from GSK's witnesses regarding physician reliance on information from generic drug makers.

    Less persuasive to the Federal Circuit majority was Teva's evidence regarding "deliberately omit[ting]" reference to CHF in its skinny label, particularly in light of the continued promotion of the product by Teva in a manner not consistent with complete exclusion of the CHF indication (albeit without express inclusion of that indication).

    The panel majority opined that the District Court erred in applying the correct legal standard, stating that "precedent makes clear that when the provider of an identical product knows of and markets the same product for intended direct infringing activity, the criteria of induced infringement are met."  Considering this precedent, the majority held that "[t]here was ample record evidence of promotional materials, press releases, product catalogs, the FDA labels, and testimony of witnesses from both sides, to support the jury verdict of inducement to infringe the designated claims for the period of the '000 reissue patent."  The opinion cites Vanda Pharm. v. West-Ward Pharm. Int'l Ltd., 887 F.3d 1117, 1129 (Fed. Cir. 2018), and Sanofi for the principle that the label's contents can be evidence of inducement to infringe.

    The majority also takes its opportunity to address (briefly) the Chief Judge's dissent, stating that it is not proper for the appellate court to "find facts afresh" nor to engage in policy arguments regarding FDA approval standards.  In the majority's view, "[t]he implications of the dissent's position are vast, and if enforcement of patents on new discoveries varies with the extent to which the patentee has profited from past discoveries, this is a policy matter for Congress, not a factor in judicial review of jury verdicts."

    The majority's opinion concludes by considering Teva's objections to the bases for the District Court's damages instructions and calculations, affirming because they concluded that "the jury instructions are in conformity to law."

    Chief Judge Prost dissented based, as the majority noted, on her objections to the quanta of evidence adduced and policy consequences should the majority's position be sustained.  The Chief begins by referencing the need, in her view, for balance between the incentives patents provide for pharmaceutical innovation and the public's need for access to that innovation once the patent term has expired, noting that this was one motivation for Congress to establish the generic drug approval system.  In the Chief's view the majority's decision undermines these policy goals by finding Teva induced infringement by marketing its generic drug produce for unpatented uses (emphasis in dissent) using its skinny label.  The dissent sets out the opinion that Teva acted just as Congress intended when it enacted the skinny label provisions, by waiting (using its Paragraph III certification) until GSK's patent had expired before launching its product for unpatented indications.  And Teva's inclusion on its label of the CHF indication (recited in GSK's '000 reissue patent) was compelled by FDA; even then according to the dissent "there was still no inducement via the full label."  The Chief Judge believes that:

    The district court got it right: no evidence established that Teva actually caused the doctors' infringement for either label.  No communication from Teva encouraged doctors to use generic carvedilol to practice the patented method.  And no evidence showed that doctors relied on Teva's label.  Indeed, GSK's own expert admitted that he had not read Teva's label before prescribing generic carvedilol.  Rather than suggest inducement, the record established that doctors relied on other sources of information, not Teva, in making their decision to prescribe carvedilol.  And in any case, the record showed that the switch from Coreg® to generic carvedilol occurred "automatically," often without doctors' knowledge at all.

    The dissent not only disagrees with the majority's decision, but recognizes that "it nullifies Congress's statutory provision for skinny labels—creating liability for inducement where there should be none," contrary to Congressional intent and "slowing, rather than speeding, the introduction of low-cost generics."

    After stating her opinion, the Chief Judge spends the rest of her lengthy dissent (at 33 pages much longer than the majority opinion) setting forth the following subjects:  1. The Statutory Background; 2. The Factual n Procedural Background; 3. [That] The Majority Nullifies Congress's Provision for Skinny Labels; and 4. [That] The Majority Misapplies the Law and Misconstrues the Facts.  The bulk of the Chief's arguments are in the third and fourth sections of her dissent (as commented upon by the majority).  Regarding the Chief's opinion that the majority engaged in judicial nullification of the skinny label option enacted by Congress, the Chief stated that:

    To hold [that Teva infringes based on its label], as the Majority does, undermines Congress's provision for skinny labels by substantially nullifying section viii.  According to the Majority, a generic company that carves out from its label a patented method of use can nonetheless be found to infringe that patented method based on the content of the FDA-approved label.  See Maj. 16.  By finding inducement based on Teva's skinny label, which was not indicated for—and did not otherwise describe—the patented method, the Majority invites a claim of inducement for almost any generic that legally enters the market with a skinny label.  That is directly contrary to Congress's intent," citing Caraco Pharm. Labs., Ltd. v. Novo Nordisk, 566 U.S. 399 (2012) (emphasis in dissenting opinion).

    The Chief also found Federal Circuit precedent contrary to the majority's opinion in Takeda Pharmaceuticals U.S.A. v. West-Ward Pharmaceutical Corp., 785 F.3d 625 (Fed. Cir. 2015), and Warner-Lambert v. Apotex Corp., 316 F.3d 1348 (Fed. Cir. 2003).  And rather than finding anything nefarious in Teva's carveout, the Chief opined that Teva "was acting in accordance with Congress's goals for it" as sanctioned by the Supreme Court in Caraco Pharm.

    Chief Judge Prost's disapproval of the majority's application of the law to the facts revisited the jury's determination and the District Court's consideration that the jury lacked substantial evidence to support its verdict of induced infringement, unavoidably at least considering if not finding facts afresh as the majority noted in their opinion.

    While not expressly discussed by either the majority or the dissent, there is a hint that the majority was concerned with generic drug companies improperly relying on the skinny label strategy to profit from "off-label use" by physicians for the very indication excluded by the skinny label carveout.  The majority extended the scrutiny regarding this stratagem perhaps farther than it can reliably be stretched, not requiring the level of promotion of the carved out indication that would be expected to attract inducement liability.  And the dissent argued strongly regarding the impropriety of the policymaking consequences of the majority's decision usurping Congress's role in deciding the extent to which a generic drug company can use the skinny label strategy to avoid infringement inducement liability.  It is likely that (at a minimum) the Court will hear this case en banc; perhaps less likely but certainly within the realm of possibility is Supreme Court review in light of the important policy considerations raised by the majority and even more strongly by Chief Judge Prost's dissent.

    GlaxoSmithKline LLC v. Teva Pharmaceuticals USA, Inc. (Fed. Cir. 2020)
    Panel: Chief Judge Prost and Circuit Judges Newman and Moore
    Opinion by Circuit Judge Newman; dissenting opinion by Chief Judge Prost