• Plus ça change, plus c'est la même chose

    By Kevin E. Noonan —
    Federal Circuit SealJudge Moore, in Athena Diagnostics, Inc. v. Mayo Collaborative Services, LLC stated the obvious when she said in her dissent:

    My colleagues' refusal deflates the Amici's hopeful suggestion that our precedent leaves the eligibility of a diagnostic claim in front of the Federal Circuit "uncertain."  It is no longer uncertain.  Since Mayo, every diagnostic claim to come before this court has been held ineligible.  While we believe that such claims should be eligible for patent protection, the majority of this court has definitively concluded that the Supreme Court prevents us from so holding.  No need to waste resources with additional en banc requests.

    In the interim it has become clear that even asking the Court to provide any answer other than an affirmance of a district court decision below invalidating claims to diagnostic methods is too much to ask, a reality made evident once again by the Court's recent decision in CareDx, Inc. v. Natera, Inc.

    To recap proceedings below, the case arose over the claims in U.S. Patent Nos. 8,703,652, 9,845,497, and 10,329,607 directed to "methods to help predict the status or outcomes of transplant recipients through sequencing of cell-free nucleic acids ("cfDNA") found in the bodily fluids of a recipient."  The rationale behind the invention is rejection of a transplanted organ in a recipient is accompanied by cell death, which releases donor-specific DNA into the recipient's bodily fluids.  Claim 1 of the '652 patent, claim 1 of the '497 patent, and claim 1 of the '607 patent were illustrative:

    Claim 1 of the '652 patent recites:

    1.  A method for detecting transplant rejection, graft dysfunction, or organ failure, the method comprising:
        (a) providing a sample comprising cell-free nucleic acids from a subject who has received a transplant from a donor;
        (b) obtaining a genotype of donor-specific polymorphisms or a genotype of subject-specific polymorphisms, or obtaining both a genotype of donor-specific polymorphisms and subject-specific polymorphisms, to establish a polymorphism profile for detecting donor cell-free nucleic acids, wherein at least one single nucleotide polymorphism (SNP) is homozygous for the subject if the genotype comprises subject-specific polymorphisms comprising SNPs;
        (c) multiplex sequencing of the cell-free nucleic acids in the sample followed by analysis of the sequencing results using the polymorphism profile to detect donor cell-free nucleic acids and subject cell-free nucleic acids; and
        (d) diagnosing, predicting, or monitoring a transplant status or outcome of the subject who has received the transplant by determining a quantity of the donor cell-free nucleic acids based on the detection of the donor cell-free nucleic acids and subject cell-free nucleic acids by the multiplexed sequencing,
        wherein an increase in the quantity of the donor cell-free nucleic acids over time is indicative of transplant rejection, graft dysfunction or organ failure, and wherein sensitivity of the method is greater than 56% compared to sensitivity of current surveillance methods for cardiac allograft vasculopathy (CAV).

    Claim 1 of the '497 patent recites:

    1.  A method of detecting donor-specific circulating cell-free nucleic acids in a solid organ transplant recipient, the method comprising:
        (a) genotyping a solid organ transplant donor to obtain a single nucleotide polymorphism (SNP) profile of the solid organ transplant donor;
        (b) genotyping a solid organ transplant recipient to obtain a SNP profile of the solid organ transplant recipient, wherein the solid organ transplant recipient is selected from the group consisting of: a kidney transplant, a heart transplant, a liver transplant, a pancreas transplant, a lung transplant, a skin transplant, and any combination thereof;
        (c) obtaining a biological sample from the solid organ transplant recipient after the solid organ transplant recipient has received the solid organ transplant from the solid organ transplant donor, wherein the biological sample is selected from the group consisting of blood, serum and plasma, and wherein the biological sample comprises circulating cell-free nucleic acids from the solid organ transplant; and
        (d) determining an amount of donor-specific circulating cell-free nucleic acids from the solid organ transplant in the biological sample by detecting a homozygous or a heterozygous SNP within the donor-specific circulating cell-free nucleic acids from the solid organ transplant in at least one assay,
        wherein the at least one assay comprises high-throughput sequencing or digital polymerase chain reaction (dPCR), and
        wherein the at least one assay detects the donor-specific circulating cell-free nucleic acids from the solid organ transplant when the donor-specific circulating cell-free nucleic acids make up at least 0.03% of the total circulating cell-free nucleic acids in the biological sample.

    Claim 1 of the '607 patent recites:

    1.  A method of quantifying kidney transplant-derived circulating [cfDNA] in a human kidney transplant recipient, said method comprising:
        (a) providing a plasma sample from said human kidney transplant recipient, wherein said human kidney transplant recipient has received a kidney transplant from a kidney transplant donor, wherein said plasma sample from said human kidney transplant recipient comprises kidney transplant-derived circulating [cfDNA] and human kidney transplant recipient-derived circulating [cfDNA];
        (b) extracting circulating [cfDNA] from said plasma sample from said human kidney transplant recipient in order to obtain extracted circulating [cfDNA], wherein said extracted circulating [cfDNA] comprises said kidney transplant-derived circulating [cfDNA] and human kidney transplant recipient-derived circulating [cfDNA];
        (c) performing a selective amplification of target [DNA] sequences, wherein said selective amplification of said target [DNA] sequences is of said extracted circulating [cfDNA], wherein said selective amplification of said target [DNA] sequences amplifies a plurality of genomic regions comprising at least 1,000 single nucleotide polymorphisms, wherein said at least 1,000 single nucleotide polymorphisms comprise homozygous single nucleotide polymorphisms, heterozygous single nucleotide polymorphisms, or both homozygous single nucleotide polymorphisms and heterozygous single nucleotide polymorphisms, and wherein said selective amplification of said target deoxyribonucleic acid sequences is by polymerase chain reaction (PCR);
        (d) performing a high throughput sequencing reaction, wherein said high throughput sequencing reaction comprises performing a sequencing-by-synthesis reaction on said selectively-amplified target [DNA] sequences from said extracted circulating [cfDNA], wherein said sequencing-by-synthesis reaction has a sequencing error rate of less than 1.5%;
        (e) providing sequences from said high throughput sequencing reaction, wherein said provided sequences from said high throughput sequencing reaction comprise said at least 1,000 single nucleotide polymorphisms; and
        (f) quantifying an amount of said kidney transplant-derived circulating [cfDNA] in said plasma sample from said human kidney transplant recipient to obtain a quantified amount, wherein said quantifying said amount of said kidney transplant-derived circulating [cfDNA] in said plasma sample from said human kidney transplant recipient comprises using markers distinguishable between said human kidney transplant recipient and said kidney transplant donor, wherein said markers distinguishable between said human kidney transplant recipient and said kidney transplant donor comprises single nucleotide polymorphisms selected from said at least 1,000 single nucleotide polymorphisms identified in said provided sequences from said high throughput sequencing reaction, and wherein said quantified amount of said kidney transplant-derived circulating [cfDNA] in said plasma sample from said human kidney transplant recipient comprises at least 0.03% of the total circulating [cfDNA] from said plasma sample from said human kidney transplant recipient.

    The Magistrate Judge resolved the issue of whether these claims were ineligible for patenting under 35 U.S.C. § 101 under the first step of the Supreme Court's test enunciated in Mayo and Alice Corp. v. CLS Bank Int'l.  Defendants argued (as they must) that the claims in the patents-in-suit were directed to one of the judicial exceptions (a natural phenomenon, specifically "the correlation between transplant rejection and the presence of naturally occurring [cfDNA] in the bodily fluids of transplant recipients").  The Magistrate, relying on Federal Circuit precedent permitting a court to consider the patent specification in determining "what a patent claim is really directed to at step one [of the Mayo/Alice test]" (Enfish LLC v. Microsoft Corp.) found that:

    [T]he patents' [related] specification repeatedly and consistently states that this basic "correlation" between the presence of increased levels of donor-specific cfDNA and transplant rejection . . . — i.e., the thing that, according to Defendants, the asserted claims were purportedly "directed to" — had already been well-known in the art for quite a long time.

    The District Court, while granting parties the opportunity for discovery and expert testimony, ultimately granted Natera's motion for summary judgment that the claims were invalid under Section 101 for lack of subject matter eligibility, and this appeal followed.

    The Federal Circuit affirmed, in an opinion by Judge Lourie joined by Judges Bryson and Hughes.  The reasoning is depressingly predictable:  that the claims fail the first prong of the Alice eligibility test for being directed to a natural phenomenon and fail the second prong of the test by reciting only conventional, well-understood, and routine methods that did not rise to the ineluctable "something more" required for eligibility.  In this, patentees fell into the trap that was sprung on unwary applicants ever since Ariosa v Sequenom.  As in that case, the particular petard upon which patentees' eligibility hopes were hoisted was disclosure in the specification regarding this conventionality, the opinion setting out in a footnote in detail the extent of what the Court found was an admission:

    See, e.g., '652 patent at col. 9 ll. 8–14 (stating that "[d]etection, identification and/or quantitation of the donor-specific markers (e.g.[,] polymorphic markers such as SNPs) can be performed using real-time PCR, chips (e.g., SNP chips), high throughput shotgun sequencing of circulating nucleic acids (e.g.[,] [cfDNA]), as well as other methods known in the art"); id. at col. 10 ll. 11–12 (stating that, to obtain cfDNA samples, "any technique known in the art may be used, e.g. a syringe or other vacuum suction device"); id. at col. 13 ll. 51–53 (stating that step 2 of claimed methods can be performed "using existing genotyping platforms know[n] in the art"); id. at col. 15 ll. 6–8 (stating that techniques recited in step 2 of claimed methods "can be accomplished through classic Sanger sequencing methods which are well known in the art"); id. at col. 13 ll. 58–61 (stating that "[c]ompanies (such as Applied Biosystems, Inc.) currently offer both standard and custom designed TaqMan probe sets for SNP genotyping that can in principle target any desired SNP position for a PCR based assay"); id. at col. 20 ll. 31–34 (stating that genotyping recited in claimed methods "may be performed by any suitable method known in the art including those described herein such as sequencing, nucleic acid array or PCR"); id. at col. 15 ll. 22–65 (discussing commercial high throughput sequencing products); id. at col. 14 ll. 58–67 (citing articles from 2006 and 2007 as supporting the statement that "digital PCR is a much more accurate and reliable method to quantitate nucleic acid species"); id. at col. 18 l. 55–col. 19 l. 2 (stating that "[m]ethods for quantifying nucleic acids," including high throughput genotyping, "are known in the art"); id. at col. 21 ll. 5–9 (stating that "[t]he presence or absence of one or more nucleic acids from the transplant donor in the transplant recipient may be determined by any suitable method known in the art including those described herein such as sequencing, nucleic acid arrays or PCR").

    The opinion states summarily that "[t]he claimed methods are indistinguishable from other diagnostic method claims the Supreme Court found ineligible in Mayo and that we found ineligible on multiple occasions."  Natera recites and the panel agrees with the familiar litany of cases coming to the same conclusion, i.e., Athena Diagnostics, Inc. v. Mayo Collaborative Servs., LLC, 915 F.3d 743 (Fed. Cir. 2019); Genetic Veterinary Scis., Inc. v. LABOKLIN GmbH & Co. KG, 933 F.3d 1302 (Fed. Cir. 2018); Roche Molecular Sys., Inc. v. Cepheid, 905 F.3d 1363 (Fed. Cir. 2018); Cleveland Clinic Found. v. True Health Diagnostics LLC, 859 F.3d 1352 (Fed. Cir. 2017); Ariosa Diagnostics, Inc. v. Sequenom, Inc., 788 F.3d 1371 (Fed. Cir. 2015).  The similarity to the Ariosa decision (which in some ways propelled the Court down this path of per se ineligibility) is express:

    Here, as in Ariosa, the claims boil down to collecting a bodily sample, analyzing the cfDNA using conventional techniques, including PCR, identifying naturally occurring DNA from the donor organ, and then using the natural correlation between heightened cfDNA levels and transplant health to identify a potential rejection, none of which was inventive.  The claims here are equally as ineligible as those in Ariosa.

    To the extent there is anything remotely new in this opinion it is the acknowledgement that conventionality is an element of step one of the Alice eligibility test, citing Athena and Cleveland Clinic decisions for the principle.

    With regard to that conventionality, the opinion illuminates the logical error of its treatment of this part of the equation.  The opinion asserts that the methods are conventional because "CareDx does not actually claim any improvements in laboratory techniques—rather, as previously discussed, the actual claims of the patent merely recite the conventional use of existing techniques to detect naturally occurring cfDNA.  Furthermore, the specification admits that the laboratory techniques disclosed in the claims require only conventional techniques and off-the-shelf technology" and "the asserted claims add nothing inventive because they merely recite standard, well-known techniques in a logical combination to detect natural phenomena."  The case also contains a convenient mantra for this rationale:  "We have repeatedly held that applying standard techniques in a standard way to observe natural phenomena does not provide an inventive concept," citing Ariosa, Athena, and Roche.  According to the Court, a conclusion of ineligibility is justified because the claimed combination of steps adds nothing inventive, analogous to the factual circumstances in Mayo v. Prometheus.

    But what the Court has consistently ignored is the difference between the claims in Mayo and the ones in Ariosa and the Court's other diagnostic method cases.  And that difference is that the detection methods reciting in the Prometheus claims were conventional because they were actually being performed in the art on the subject matter and for the purpose (assessing the amount of drug in a patient's blood after administration) recited in the claims.  The only distinction from these conventional methods in those claims was the recognition that there were boundary levels of detected drug concentrations that indicated whether the dosage should be increased or decreased.  In contrast, in all the diagnostic method cases that have fallen under the Court's ineligibility ax since Mayo there had not been any recognition, much less practice, in the prior art of these methods on this subject matter to detect the natural phenomenon that has been used to satisfy step one of the Alice test.  The inventiveness resides there, and refusal to recognize that distinction is the principal reason for the Court's continuing invalidity jurisprudence.

    In her dissent in Athena, Judge O'Malley noted that:

    Since Mayo, every diagnostic claim to come before this court has been held ineligible.  While we believe that such claims should be eligible for patent protection, the majority of this court has definitively concluded that the Supreme Court prevents us from so holding.  No need to waste resources with additional en banc requests.  Your only hope lies with the Supreme Court or Congress.  I hope that they recognize the importance of these technologies, the benefits to society, and the market incentives for American business.  And, oh yes, that the statute clearly permits the eligibility of such inventions and that no judicially-created exception should have such a vast embrace.  It is neither a good idea, nor warranted by the statute.

    In view of the Supreme Court's denial of certiorari in American Axle v. Neapco it appears Congress (other than reliance on trade secret protection) remains the only source of any respite from the scourge of ineligibility for diagnostic methods claims.

    CareDx, Inc. v. Natera, Inc. (Fed. Cir. 2022)
    Panel: Circuit Judges Lourie, Bryson, and Hughes
    Opinion by Circuit Judge Lourie
  • By Michael Borella —

    Federal Circuit SealCan a prior art reference with an error be considered to be a disclosure of the erroneous teaching?  A Federal Circuit panel split over this issue, with their disagreement largely based on how apparent the error would be to one skilled in the art.

    LG filed two Inter Partes Reviews (IPRs) against U.S. Patent No. 6,844,990, challenging claims 5 and 21, respectively.  Both Patent Trial and Appeal Board (PTAB) panels sided with Immervision, finding that a critical section of the prior art used in LG's obviousness contentions contained an error that "would have been disregarded or corrected by a person of ordinary skill in the art."

    The invention is described by the Court as follows:

    The '990 patent relates to capturing and displaying digital panoramic images.  Panoramic (e.g., super-wide angle) objective lenses typically have linear image point distribution functions.  This means there is a linear relationship between the distance of an image point from the image's center and the corresponding relative angle of the object point to the image's center.  While this linearity allows digital panoramic images to be easily rotated, shifted, and enlarged or shrunk, it also limits image quality to the resolution of the image sensor used when taking the initial image.

    * * *

    The '990 patent purports to improve the resolution of particular sectors of a digital panoramic image [by] capturing an initial digital panoramic image using an objective lens having a non-linear image point distribution function that expands certain zones of the image and compresses other zones of the image.

    Along these lines, representative claim 5 recites:

    5.  The method according to claim 1, wherein the objective lens compresses the center of the image and the edges of the image and expands an intermediate zone of the image located between the center and the edges of the image.

    LG's arguments that claims 5 and 21 are obvious relied on disclosure from U.S. Patent No. 5,861,999 ("Tada"), which described four embodiments "relating to the aspheric characteristics of various lens elements."  Embodiment 3 of Tada described a set of optical parameters in Table 5.

    LG contended that Embodiment 3 disclosed the features of claim 5.  As noted by the Court, "Tada, however, does not explicitly discuss the image point distribution functions of its lenses" and "[i]nstead LG relied on its expert Dr. Russell Chipman's declaration for the proposition that Tada's third embodiment has a distribution function" as claimed.  Indeed, Dr. Chipman used Table 5 of Tada to model the lens of Embodiment 3 and calculated that this embodiment produces the compression and expansion of claim 5.  On this basis, the PTAB instituted both IPRs.

    Immervision hired its own expert, Mr. David Aikens, who independently modeled the lens of Embodiment 3 using the parameters from Table 5.  Mr. Aikens, however, found that the lens did not match the corresponding example in one of Tada's figures.  After some investigation, Mr. Aikens concluded that the lens would provide an image that was "distorted with precisely the kind of uncorrected field curvature that Tada was explicitly trying to prevent."  Thus, Mr. Aiken's wrote that "a person of ordinary skill in the art would be convinced that there was an error in the model and that the error was significant."

    Looking into the matter further, Mr. Aikens found that "the aspheric coefficients from Table 3, which corresponds to Tada's Embodiment 2, were exactly the same as in Table 5, which corresponds to Embodiment 3.  Further, Tada claimed priority to a Japanese patent application.  Mr. Aikens also reviewed this application and found that the parameters in its version of Table 5 were different from those of the '999 patent.  Using the different parameters from the Japanese application, Mr. Aikens confirmed that they produced a lens surface that perfectly matched the other disclosure in Tada.  In other words, Table 5 of Tada, which was relied on by LG, was the product of a copy and paste error and thus contained incorrect values.

    In both IPRs, the PTAB concluded that the "disclosure of aspheric coefficients in Table 5 of Tada is an obvious error that a person of ordinary skill in the art would have recognized and corrected."  Thus, the PTAB ruled that LG had failed to prove claims 5 and 21 obvious.  LG appealed.

    Judge Stoll, writing in majority for herself and Judge Cunningham, rapidly determined that "[i]t is undisputed that the aspheric coefficients in Tada's Table 5 were erroneous" and that "there is no dispute that if a lens were constructed using the (correct) aspherical data from Tada's Japanese priority application, the lens would not satisfy the compression and expansion zone limitation of claims 5 and 21."  The question to the Court, then, was "whether substantial evidence supports the Board's fact finding that the error would have been apparent to a person of ordinary skill in the art such that the person would have disregarded the disclosure or corrected the error."

    In making this determination, the majority relied on In re Yale, a Court of Customs and Patent Appeals case from 1970.  Therein, a similar fact pattern was present — a claim was rejected over a reference that erroneously disclosed a chemical compound that was not discovered until several years after the reference was published.  Notably, the reference's description of this compound was inconsistent, and later an author of the reference admitted that disclosure of the compound was erroneous.

    The holding from Yale, and described by the majority, was:

    [W]here a prior art reference includes an obvious error of a typographical or similar nature that would be apparent to one of ordinary skill in the art who would mentally disregard the errant information as a misprint or mentally substitute it for the correct information, the errant information cannot be said to disclose subject matter.  The remainder of the reference would remain pertinent prior art disclosure.  This standard for reviewing errors in disclosures has been undisturbed for half a century and we are bound to apply it.

    Applying this law to the facts of this case, the majority found that "[t]he Board correctly identified several aspects of the disclosure in Table 5 that would alert the ordinarily skilled artisan that the disclosure was an obvious error of a typographical or similar nature."  The first of these aspects was that Table 5 in the Japanese priority application had different parameters than the equivalent Table 5 in Tada.  Second, Tada's Table 5 was inconsistent with other disclosure in Tada.  Third, the fact that the parameters from Tables 3 and 5 were identical "is incongruous with the differences in the values of other data for the lens systems."

    LG made two arguments against the PTAB's conclusion.  The first was that "Mr. Aikens' convoluted process that took ten to twelve hours to complete clearly weighed against the obviousness of the error."  The majority disagreed, noting that Yale does not impose a temporal requirement regarding how long it would take one to determine that a reference contains an error.  The second was that Yale should be limited to just typographical errors.  Again, the majority disagreed, finding that the distinction between the typographical error of Yale and the copy-and-paste error of Tada was not significant enough to overrule the PTAB.

    Thus, the majority affirmed the PTAB's final written decisions that LG had not established the obviousness of claims 5 and 21.

    Judge Newman wrote in dissent.  The crux of her disagreement with the majority was that the error in Tada was not found "until an expert witness conducted a dozen hours of experimentation and calculation."  Thus, she did not believe that the error to be merely typographical.

    Judge Newman observed that the error in Table 5 was not noticed by prosecuting patent attorneys, the patent examiner, or in a certificate of correction that was obtained to address other errors in Tada.  The error was also not noticed by two PTAB panels that instituted the IPRs.[1]  Judge Newman also noted that Mr. Aikens only noticed the error after creating a model for the lens and hours of subsequent investigation.  In Judge Newman's view, this distinguished the situation surrounding Tada from that of Yale, mainly because "Yale did not require calculations or experimentation" and "the correct information is not readily evident."[2]

    Thus, Judge Newman would reduce the scope of Yale to typographical errors that are readily recognized as such by a quick review of the prior art reference.

    [1] In modest disagreement with Judge Newman, once an error like this is in a patent application, it would be unlikely to be noticed during prosecution and subsequent proceedings that are largely focused on the language of the claims.  In contrast, the hypothetical person of ordinary skill in the art is presumed to review the patent in its totality.

    [2] Judge Newman admitted that by looking to the Japanese priority application, one could readily identify the error.  She wrote that "[i]t should not be necessary to search for a foreign document in a foreign language to determine whether there is an inconsistency in a United States patent."  But such an activity is certainly within the ambit of one of ordinary skill in the art, especially as this individual is presumed to be aware of all relevant prior art.

    LG Electronics v. Immervision, Inc. (Fed. Cir. 2022)
    Panel: Circuit Judges Newman, Stoll, and Cunningham
    Opinion by Circuit Judge Stoll; opinion dissenting in part by Circuit Judge Newman

  • CalendarJuly 18, 2022 – Southeast Asia intellectual property roadshow (U.S. Patent and Trademark Office, U.S.-ASEAN Business Council, the U.S. Department of Commerce's International Trade Administration, and the U.S. Commercial Service) – 7:00 pm to 9:00 pm (ET)

    July 21 2022 – "AI Is Coming to an Invention Near You" (Fitch Even) – 12:00 pm (ET)

    July 21, 2022 – "Boosting the Bottom Line Strategies for Managing Patents During a Recession" (IPWatchdog and UnitedLex) – 12:00 pm (ET)

  • Fitch EvenFitch Even will be offering a webinar entitled "AI Is Coming to an Invention Near You" on July 21 2022 at 12:00 pm (ET).  Steven G. Parmelee of Fitch Even will discuss the following topics:

    • A general explanation of AI and machine learning (ML) in particular
    • A statistical overview of ML in US patenting
    • A patent eligibility overview of ML claims
    • Various approaches to claim drafting
    • Practice tips

    While there is no cost to participate in the program, advance registration is required.  Those interested in attending the webinar can register here.

  • IPWatchdogIPWatchdog and UnitedLex will be offering a webinar entitled "Boosting the Bottom Line Strategies for Managing Patents During a Recession" on July 21, 2022 at 12:00 pm (ET).  Gene Quinn of IPWatchdog, Inc. will moderate a panel consisting of Erica Helgerson, pSemi Corporation, a Murata Company; Nidhi Nahar, Block Inc.; Patrick Woolley of Polsinelli; and Joe Dearing, UnitedLex.  The panel will cover 4 strategies that can impact attendees' bottom lines in 2022, and help attendees prune with confidence and precision, including:

    • Effective methods for pruning low-value patents from your portfolio
    • Synching your patent filing strategy to the competitive landscape
    • Identifying and divesting patents no longer core to your business
    • Monetization strategies on high-value and infringed-upon patents

    There is no registration fee for this webinar.  However, those interested in registering for the webinar, should do so here.

  • USPTO SealThe U.S. Patent and Trademark Office, in cooperation with the U.S.-ASEAN Business Council, the U.S. Department of Commerce's International Trade Administration, and the U.S. Commercial Service, will be offering a Southeast Asia intellectual property roadshow on July 18, 2022 from 7:00 pm to 9:00 pm (ET).  Policy makers, thought leaders, and experienced practitioners in the field of intellectual property (IP)—including speakers from the U.S. Patent and Trademark Office (USPTO), the U.S. Association of Southeast Asian Nations (US-ASEAN) Business Council, the U.S. Department of Commerce, and the private sector—will be presenting a free webinar on IP issues in Southeast Asia and the ASEAN Economic Community.  Topics to be covered during the webinar will include:

    • Protecting and enforcing IP in ASEAN Economic Community markets
    • Practical market experiences in Southeast Asia
    • Cross-border e-commerce digital asset protection considerations
    • U.S. government resources

    Those interested in registering for the webinar can do so here. 

  • By Kevin E. Noonan —

    Federal Circuit SealAlmost four years ago, in a relatively rare occurrence based on there being an insufficient factual record to permit proper appellate review, the Federal Circuit vacated a District Court decision rendering invalid the claims in five patents asserted by Tris Pharma, Inc. against Teva Laboratories FL, Inc. and remanded.  The Court has had the opportunity to review the District Court's decision on remand, delayed by the COVID pandemic, and today affirmed the determination that Teva had not shown the claims to be invalid by clear and convincing evidence.

    To recap, the case arose in ANDA litigation regarding Quillivant XR®, an extended release, liquid methylphenidate (MPH) formulation for the treatment of Attention Deficit Hyperactive Disorder (ADHD).  The District Court found the asserted claims of U.S. Patent Nos. 8,465,765 ('765 patent), 8,563,033 ('033 patent), 8,778,390 ('390 patent), 8,956,649 ('649 patent), and 9,040,083 ('083 patent) were invalid as being obvious under 35 U.S.C. § 103.  As explained in the earlier opinion, MPH is a widely used psychostimulant that has been used for treating ADHD since the 1950's.  Both immediate-release (IR) and extended-release (ER) formulations of the drug were also known in the art, and sustained-release (SR) formulations were later developed to overcome drawbacks of both IR and ER formulations.  Unfortunately, prior art SR formulations were disadvantageous for having slow onset action properties.  The claimed invention was a combination formulation comprising an IR component and an SR component that showed "a 45-minute therapeutic onset and 12 hours of therapeutic effect."

    Asserted claim 6 (dependent on claim 1) was at issue in the litigation and was deemed to be representative:

    1.  A methylphenidate aqueous extended release oral suspension comprising (1) an immediate release methylphenidate component, (2) a sustained release methylphenidate component comprising a water-insoluble, water-permeable, pH-independent, barrier coated methylphenidate-ion exchange resin complex, and (3) water, wherein said suspension has a pH of about 3.5 to about 5 and said suspension provides a single mean average plasma concentration peak for methylphenidate and a therapeutically effective plasma profile for methylphenidate for about 12 hours.

    6.  The suspension according to claim 1, wherein the suspension has a pharmacokinetic profile in which the single mean plasma concentration peak for d-methylphenidate has an area under the curve (AUC)0-∞of about 114 ng-hr/mL to about 180 ng-hr/mL, Cmax of about 11 ng/mL to about 17 ng/mL, Tmax of about 4 hours to about 5.25 hours and T1/2 of about 5 hours to about 7 hours following a single oral administration of an aqueous suspension at a dose equivalent to 60 mg racemic methylphenidate HCl in adults.

    The prior art considered by the District Court in the earlier case included several commercially available, extended-release MPH formulations (Concerta®, Daytrana®, Focalin XR®, Metadate CD®, and Ritalin LA®), U.S. Patent Application Publication No. 2010/0260844, and certain scientific publications.  The commercial products all exhibited various pharmacokinetic and pharmacodynamics properties regarding a single or multiple PK peak profile, initial onset time and total duration.  The '844 patent publication disclosed "a formulation of MPH that provides a rapid onset of action within 1 to 1.5 hours, a single Tmax of 5.5 to 7.5 hours, and a therapeutic duration of about 12 to 14 hours."  Actavis relied on the '844 application in combination with Concerta®, Daytrana®, and Metadate CD® as teaching "a single mean peak PK profile, exhibiting an early onset of action, and exhibiting an extended duration of effect" that would have suggested to the skilled worker the early onset and extended duration formulation in the asserted claims.  Tris Pharma argued to the contrary that the prior art taught combinations of IR and ER MPH formulations that resulted in a bimodal PK profile.  This feature of combined IR and ER MPH formulations was designed to "mimic the peaks and valleys of multiple immediate release dosing regimens" which was not a feature of the claimed invention.  In addition, the commercially available formulations showed a later Tmax that was important to achieve the longer duration.

    The District Court found the claims to be obvious and the Federal Circuit disagreed, vacating and remanded, in an opinion by Judge Chen, joined by Judges Newman and O'Malley.  The opinion was based on Fed. R. Civ. P. 52(a)(1), which states that "[i]n an action tried on the facts without a jury or with an advisory jury, the court must find the facts specially and state its conclusions of law separately."  Citing Gechter v. Davidson, 116 F.3d 1454, 1457 (Fed. Cir. 1997), the Federal Circuit stated that "[w]hen the opinion explaining the decision lacks adequate fact-findings, meaningful review is not possible, frustrating the very purpose of appellate review as well as this court's compliance with its statutory mandate."  Such considerations were important in this case because the paucity of the District Court's fact-finding made it impossible for the panel to properly assess the correctness of its legal conclusion of obviousness.

    On remand, the District Court considered dependent claim 10 of the '033 patent to be exemplary:

    1.  A methylphenidate aqueous extended release oral suspension comprising (1) an immediate release methylphenidate component, (2) a sustained release methylphenidate component, and (3) water, said suspension having a pH of about 3.5 to about 5, wherein said suspension provides a single mean average plasma concentration peak and a therapeutically effective plasma profile for about 12 hours for methylphenidate, and wherein the suspension has a pharmacokinetic profile in which the single mean plasma concentration peak for methylphenidate has an area under the curve (AUC)0 ® ∞ of about 114 to about 180 ng-hr/mL, Cmax of about 11 to about 17 ng/mL, Tmax of about 4 to about 5.25 hours, and T1/2 of about 5 to about 7 hours following a single oral administration of said suspension at a dose equivalent to 60 mg racemic methylphenidate HCl in adults.

    9.  A method for treating a patient having a condition susceptible to treatment with methylphenidate, the method comprising administering to the patient the suspension according to claim 1, wherein said suspension provides a therapeutically effective amount of methylphenidate within 45 minutes after administering of said suspension and a single average plasma concentration peak.

    10.  The method according to claim 9, wherein the suspension which has a pH from about 4 to about 4.5.

    (In a footnote the opinion notes that claims 6 and 20 of the '765 patent, claims 4 and 10 of the '033 patent, and claims 15, 16, and 20 of the '390 patent were under consideration on appeal.)  The opinion set forth the claim limitations relevant to the issues on appeal to be:

    • "aqueous,"
    • "therapeutically effective plasma profile . . . for about 12 hours" (12-hour duration, relevant to extended-release formulations),
    • "therapeutically effective amount of methylphenidate within 45 minutes" (45-minute onset),
    • "Tmax of about 4 to about 5.25 hours" (early Tmax range), and
    • "single mean plasma concentration peak" (single mean peak).

    All of the claims before the Court recited "a liquid MPH formulation with a 12-hour duration and single mean peak," and claim 6 of the '765 patent, claim 4 of the '033 patent, and claims 15,16, and 20 of the '390 patent recited the early Tmax range.  Claim 10 was considered representative because it was the only claim that recited all these features.

    On remand, Actavis relied upon the same factual evidence it adduced in the earlier trial, relating to five commercially available prior art MPH formulations, prior art patent publication 2010/0260844 and scientific articles.  According to the panel opinion "no single reference disclose[d] all of the limitations in any given claim" (although some references disclosed "various subsets of the claim limitations").  Having considered "the trial record, post-trial record, and the parties' post-remand briefing" the District Court found Actavis failed to show the asserted claims were invalid for obviousness by clear and convincing evidence, and this appeal followed.

    The Federal Circuit affirmed this time around, in an opinion by Judge Chen joined by Chief Judge Moore and Judge Hughes.  Considering the asserted claims seriatim, the Court affirmed the District Court's determination that Actavis failed to prove that "an artisan of ordinary skill would have been motivated to combine with a reasonable expectation of success a liquid MPH formulation with a single mean peak, 12-hour duration, and 45-minute onset" with regard to claim 20 of the '765 patent.  In particular, Actavis did not adduce evidence that "a skilled artisan would have pursued a single mean peak concentration profile," instead merely showing that single or bimodal peaks were potential options and the art taught away from a single mean peak.  These determinations were supported, according to the Court, by objective indicia of non-obviousness, specifically unexpected results, and the existence of a long-felt need for the claimed formulations.

    For claim 6 of the '765 patent, claim 4 of the '033 patent, and claims 15, 16, and 20 of the '390 patent, the Federal Circuit affirmed the District Court's decision that Actavis had not established that "an artisan of ordinary skill would have been motivated to combine with a reasonable expectation of success a liquid MPH formulation with a single mean peak, 12-hour duration, and the claimed Tmax range."  Specifically, the District Court held (and the Federal Circuit agreed) that while one of the references disclosed these features the prior art formulations were not liquid ones, which was required by the asserted claims.  And the District Court found that Actavis failed to establish that claim 10 of the '033 patent was invalid for obviousness because of a failure in showing a motivation to combine these references.

    On appeal, the panel rejected Actavis's arguments that the District Court erred in finding no motivation to combine the asserted references and no reasonable expectation of success.  In the Court's opinion, the lower court's findings were supported by the evidence of record as supplemented by additional evidence required by the Federal Circuit in its remand decision.  The District Court held that Actavis had not met this burden on remand.

    Specifically with regard to the 45-min onset claims, the Federal Circuit held that the District Court properly found Actavis produced insufficient evidence for motivation to combine the single mean peak profile with "a liquid formulation with a 12-hour duration and 45-minute onset" as required in claim 20 of the '765 patent (what evidence there was being "at best, inconsistent").  The Federal Circuit agreed with the District Court's determination that the prior art taught away from "a liquid formulation with a single peak, 12-hour duration, and 45-minute onset," based on expert testimony inter alia that "[t]o achieve both an early onset and longer duration, a formulator would seek a concentration profile with two separate (bimodal) peaks" rather than the single mean peak profile recited in the asserted claims.  And the panel found that the District Court properly did not find the asserted claims obvious over the asserted prior art patent application disclosure because expert testimony characterized these teachings as "aspirational" and "hypothetical."

    Also unpersuasive was Actavis's assertion of U.S. Patent Pub. No. 2007/215511 (Mehta) for teaching "an artisan of ordinary skill how to develop a liquid formulation of MPH with a single mean peak [pharmacokinetic] profile, 12-hour duration, and 45-minute onset."  The District Court found these teachings to be at "too high a level of generality" to supply the requisite motivation to combine and reasonable expectation of success.  Finally, the panel agreed with the District Court's determination that there were secondary indicia factors that supported the District Court's non-obvious determination.

    Regarding claims that recited the "early Tmax" limitation, the District Court found and the Federal Circuit affirmed that the '844 published patent application "presented only a hypothetical formulation without any data or explanation for why or how a formulation with the claimed limitations could be accomplished."  Accordingly, the District Court did not err in finding that claim 6 of the '765 patent, claim 4 of the '033 patent, and claims 15, 16, and 20 of the '390 patent were not obvious because Actavis did not show by clear and convincing evidence a motivation to combine and reasonable expectation of success.

    On this record, the Federal Circuit affirmed.

    Tris Pharma, Inc. v. Actavis Laboratories FL, Inc. (Fed. Cir. 2022)
    Nonprecedential disposition
    Panel: Chief Judge Moore and Circuit Judges Chen and Hughes
    Opinion by Circuit Judge Chen

  • By Kevin E. Noonan —

    Federal Circuit SealThe Federal Circuit today affirmed a decision by the District Court denying defendant Thales DIS AIS Deutschland USA's motion for preliminary injunction to prevent patentee plaintiff Koninklijke Philips N.V. from obtaining an exclusion order from the International Trade Commission (ITC) under Section 337 of the Tariff Act of 1930 (19 U.S.C. § 1337), in Koninklijke Philips N.V. v. Thales DIS AIS Deutschland USA LLC.

    The ITC provides relief in the form of an exclusion order against importation of goods shown to infringe a U.S. patent.  After the Supreme Court's decision in eBay v. MercExchange, the more reliable access to injunctions available from the ITC became attractive to patent plaintiffs under circumstances where all or a portion of patented product was imported into the U.S.  Section 337 proceedings are also amenable to motions for preliminary injunction, with all the strategic benefits such motions can have (for a patent holder or, as here, an accused infringer wishing to avoid an injunction that would halt importation until district court litigation was completed or the case settled).  But such injunctions are not automatic, as Thales found in this case.

    The subject matter at issue was telecommunications equipment and technology, including wireless networks.  After fruitless negotiations regarding licensing of Philips' disputed standard essential patents (SEP) and Thales infringement thereof in its products, Philips filed an infringement suit in the U.S. District Court of Delaware and for an exclusion order before the ITC.  Thales counterclaimed for breach of contract and for a declaratory judgment on the FRAND rate determination, as well as a preliminary injunction to bar Philips from seeking a Section 337 exclusion order.  The District Court denied Thales' preliminary injunction motion and this appeal followed.

    The Federal Circuit affirmed, in an opinion by Chief Judge Moore joined by Judges Dyk and Chen.  As the Court explained, whether a preliminary injunction is granted is within the sound discretion of the district court and Federal Circuit review of the district court's decision was for an abuse of discretion.  The Court's assessment of this question was grounded on the sufficiency of Thales' satisfaction of the requirements for a preliminary injunction, which were that the moving party is "likely to succeed on the merits, that [it] is likely to suffer irreparable harm in the absence of preliminary relief, that the balance of equities tips in [its] favor, and that an injunction is in the public interest," citing Luminara Worldwide, LLC v. Liown Elecs. Co., 814 F.3d 1343, 1352 (Fed. Cir. 2016) (quoting Winter v. Nat. Res. Def. Council, Inc., 555 U.S. 7, 20 (2008)).  The basis for the panel's opinion that the District Court found Thales was properly not entitled to a preliminary injunction was the insufficiency of their allegations regarding the likelihood of irreparable harm.  "[M]ere possibility or speculation of harm is insufficient," according to the opinion, citing Winter.  Speculation (exemplified in the opinion as "customers merely expressing concern that a potential future ITC exclusion order could affect Thales' ability to deliver products down the road") does not evince a likelihood of irreparable harm, according to the Court (emphasis in opinion), citing Ferring Pharms., Inc. v. Watson Pharms., Inc., 765 F.3d 205, 219 (Fed. Cir. 2014), and Takeda Pharms. U.S.A., Inc. v. Mylan Pharms. Inc., 967 F.3d 1339, 1349 & n.8 (Fed. Cir. 2020).  The panel found no clear error by the District Court in finding Thales' evidence "conclusory" because it presented no evidence that "[Thales] lost customers, had customers delay purchases, or struggled to acquire new business because of the ongoing ITC proceedings."  Instead, Thales produced affidavits from clients that they had "voice[d] concerns" and "expressed doubt" regarding Thales' continued reliability as a supplier (expressed at oral argument as living under a "cloud on the business").  This is not sufficient evidence to satisfy the likelihood of irreparable prong of the test for obtaining a preliminary injunction, the Court held, citing Takeda Pharms and Celsis In Vitro, Inc. v. CellzDirect, Inc., 664 F.3d 922, 930 (Fed. Cir. 2012).

    Accordingly, the Court affirmed the District Court's denial of Thales' preliminary injunction motion; the opinion also ordered Thales to pay costs.

    Koninklijke Philips N.V. v. Thales DIS AIS Deutschland USA LLC (Fed. Cir. 2022)
    Panel: Chief Judge Moore and Circuit Judges Dyk and Chen
    Opinion by Chief Judge Moore

  • By Kevin E. Noonan —

    An international cadre of scientists* from almost 70 institutions worldwide recently reported their findings in the scientific journal Nature that the domesticated dog (Canis familiaris) arose from two populations of ancestral grey wolves (Canis lupus) (see "Grey wolf genomic history reveals a dual ancestry of dogs").

    The grey wolf is recognized as the species that survived the last Ice Age (the last glacial maximum or LGM occurring ~28,000-23,000 years ago) to give rise to domesticated dogs, but little is known about progenitor wolf populations.  Siberian grey wolves are known to have survived from this period but it is unknown the extent to which other grey wolf populations, which were widely distributed in the Northern Hemisphere for the last few hundred thousand years, either went extinct or responded to changes in climate by adaptation.  Archeological evidence places domesticated dog populations arising ~14,000 years ago, and divergence from grey wolf species occurring ~40,000-14,000 years ago.  But analyses of modern grey wolves and dogs have been unable to resolve domesticated dog origins due to, inter alia, dog genetic diversity and local extinction and gene flow occurring after domestication.

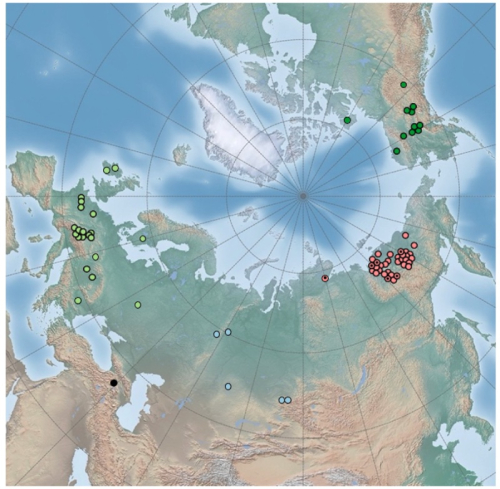
    To attempt to address these deficiencies, the researchers assessed 72 ancient wolf genomes over the past 100,000 years from Europe, Siberia, and North America:

    Image 1Sites of ancient wolf genomes reported in the study

    (consisting of 66 newly sequenced genomes, 5 previously sequenced specimens, and one ancient dhole genome from the Caucasus, dated to be more than 70,000 years old and used as an "outgroup" control).  These samples were ~69% males and through mitochondrial DNA analysis averaged 21,573 ± 5,133 years old.  As explained in the paper, the researchers "merged single-nucleotide polymorphism (SNP) genotypes called from these genomes with those from worldwide modern wolves (n = 68), modern (n = 369) and ancient (n = 33) dogs, and other canid species [and t]he total dataset spans the last 100,000 years."

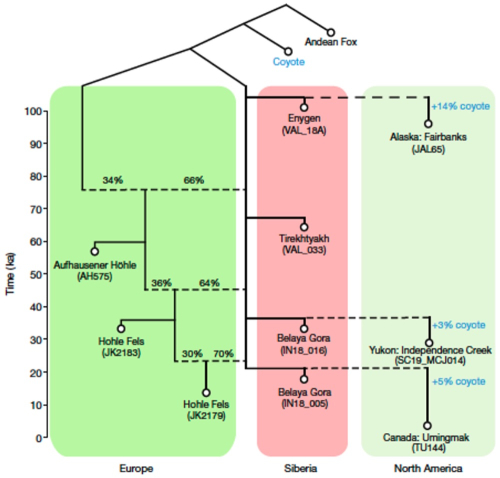
    The results of genetic analyses on these SNPs showed a directionality of gene flow in wolf populations, from the Siberian wolf populations to the European and Central Asian wolves and not vice versa, starting after a time less than ~23,000 years ago; these conclusions were supported by mitochondrial DNA assessments ("our results suggest that Siberia acted as a source and Europe as a sink for migration throughout the Late Pleistocene and show no evidence of gene flow in the other direction").  Paradoxically, these results also showed that there remains what the researchers termed a "minority fraction of deep European ancestry" that has persisted in modern wolf populations, having 10-40% of their ancestry that is more divergent than the oldest Siberian wolves studied.  Some of this diversity was identified as coming from the African grey wolf in the Near East and an unknown canid originating in Tibet.  This persistent evidence indicated to these researchers that grey wolves, unlike other megafauna (e.g., woolly mammoths) did not come close to extinction.  And some North American grey wolf populations show evidence of admixture with coyotes, species that began to diverge from one another ~700,000 years ago (no similar coyote-wolf admixture was seen in Eurasian grey wolf populations).  The genetic relationships between these populations were represented in the paper by this graph:

    Image 2
    And while there is some evidence of Siberian grey wolf genetic admixture in North American wolf populations the genetic evidence reported by these authors showed no reverse admixture of North American grey wolf populations into Siberian grey wolves.  The evidence also suggests local extinction in North American grey wolf populations during the LGM of the last Ice Age.

    In comparing the proportion of genetic variation between rather than within grey wolf populations, the researchers found that wolf populations showed low levels of variation even between populations in distant regions in the Pleistocene.  In the last ~10,000 years, however, (i.e., the Holocene) further gene flow from Siberia was not detected while gene flow from European populations was seen in modern wolves from China and Siberia, with population bottlenecks perhaps (the authors speculate) being due to human "persecution" in the past few centuries (supported by widespread effective population size declines).

    In further analyzing the observed connectivity in grey wolf populations throughout the past 100,000 years the authors report assessment of alleles at specific genetic loci, finding twenty-four regions in grey wolf genomes having some evidence for natural selection.  The observed connectivity between grey wolf populations resulted in mutations becoming fixed in these population between 40,000 and 30,000 years ago in the case of the IFT88 gene (a gene wherein disruption results in craniofacial abnormalities and cleft lips in mice and humans) on wolf chromosome 25, a characteristic shared with domesticated dogs.  (It might be recalled that changes in visage in domesticated dogs has been hypothesized as being relevant to their acceptance by humans; see "Selection for Facial Features in Domestic Dogs: The Evolution of Cuteness").  Additionally, these researchers reported that "[t]hree regions with evidence for selection overlap olfactory receptor genes, with variants on chromosome 15 increasing in frequency from close to 0% to 100%" between 45,000 and 25,000 years ago, "suggesting that olfaction was a recurrent target of adaptation in wolves" and that "[m]ost of the detected selection episodes occurred before the divergence of dogs, and dogs share the selected alleles."  Illustrating how natural selection and intentional breeding can operate on similar phenotypic determinants, the authors report that "a region on chromosome 10, where variation among dogs is associated with body size, drop ears and other traits [has been] under recent selection in specific dog breeds" was also found to have been selected in wolves in the last 20,000 years.

    While the impetus for the reported research was to better understand the origin of domesticated dog species from earlier wolf populations, the results of these studies showed that history is more complex than anticipated.  A closer relationship was observed between modern domesticated dogs from eastern Eurasian wolves than western Eurasian populations, but modern domesticated dogs in the Near East and Africa could trace their ancestral species from a population of grey wolves related to extant southwestern Eurasian populations.  Recent population genetic events, including admixture and population changes further complicate the picture, according to the paper.  As summarized, the authors state that "[t]hese results could be taken to support an eastern or central Eurasian dog origin outside of north-eastern Siberia, but we cannot draw firm geographical conclusions in the absence of ancient wolf genomes from these and other candidate regions."

    To further complicate matters, their results showed that "dogs have variable proportions of two distinct components of wolf ancestry" between Siberian wolf progenitors and European grey wolf populations, as illustrated by this diagram:

    Image 3The authors state that the data were not sufficiently robust to distinguish between there having been an independent domestication event from these population or from admixture of domesticated dogs with local wolf populations.  As a consequence, the researchers failed to find a direct match between either of these ancestral species and these two sources of modern domesticated dogs, concluding that "the exact progenitor populations remain to be located."

    *Anders Bergström, David W. G. Stanton, Ulrike H. Taron, Laurent Frantz, Mikkel-Holger S. Sinding, Erik Ersmark, Saskia Pfrengle, Molly Cassatt-Johnstone, Ophélie Lebrasseur, Linus Girdland-Flink, Daniel M. Fernandes, Morgane Ollivier, Leo Speidel, Shyam Gopalakrishnan, Michael V. Westbury, Jazmin Ramos-Madrigal, Tatiana R. Feuerborn, Ella Reiter, Joscha Gretzinger, Susanne C. Münzel, Pooja Swali, Nicholas J. Conard, Christian Carøe, James Haile, Anna Linderholm, Semyon Androsov, Ian Barnes, Chris Baumann, Norbert Benecke29, Hervé Bocherens, Selina Brace, Ruth F. Carden, Dorothée G. Drucker, Sergey Fedorov, Mihály Gasparik, Mietje Germonpré, Semyon Grigoriev, Pam Groves, Stefan T. Hertwig, Varvara V. Ivanova, Luc Janssens, Richard P. Jennings, Aleksei K. Kasparov, Irina V. Kirillova, Islam Kurmaniyazov, Yaroslav V. Kuzmin, Pavel A. Kosintsev, Martina Lázničková-Galetová, Charlotte Leduc, Pavel Nikolskiy, Marc Nussbaumer, Cóilín O'Drisceoil, Ludovic Orlando, Alan Outram, Elena Y. Pavlova, Angela R. Perri, Małgorzata Pilot, Vladimir V. Pitulko, Valerii V. Plotnikov, Albert V. Protopopov, André Rehazek, Mikhail Sablin, Andaine Seguin-Orlando, Jan Storå, Christian Verjux, Victor F. Zaibert, Grant Zazula, Philippe Crombé62, Anders J. Hansen, Eske Willerslev, Jennifer A. Leonard64, Anders Götherström, Ron Pinhasi, Verena J. Schuenemann, Michael Hofreiter, M. Thomas P. Gilbert, Beth Shapiro, Greger Larson, Johannes Krause, Love Dalén & Pontus Skoglund.

  • IPO #2The Intellectual Property Owners Association (IPO) will offer a one-hour webinar entitled "The Implementation of WIPO Standard ST.26" on July 14, 2022 from 2:00 pm to 3:00 pm (ET).  Anish Gupta, PCT Legal Examiner-International Patent Legal Administration, U.S. Patent and Trademark Office; Wayne Jaeschke of Johnson & Johnson; Hanna Kang, Legal Officer, PCT Legal and User Relations Division, World Intellectual Property Organization; and Mary Till, Senior Legal Advisor in the Office of Patent Legal Administration (OPLA), U.S. Patent Trademark Office will address the implementation of the World Intellectual Property Office (WIPO) Standard ST.26 and discuss the changes to sequence listings filed in patent applications that contain disclosures of nucleotide and/or amino acid sequences.  WIPO Standard ST.26, requiring XML formatting, went into effect worldwide on July 1, 2022.  The panel will provide guidance on how you will be able to prepare and submit a compliant sequence listing in U.S. patent applications and PCT applications filed on or after July 1, 2022.

    The registration fee for the webinar is $150 for non-members or free for IPO members (government and academic rates are available upon request).  Those interested in attending the webinar should register here.