• New York #2 American Conference Institute (ACI) will be holding its Boston edition of its Life Sciences Business Development & Acquisitions in Emerging Markets conference on September 26-27, 2011 in New York, NY.  ACI faculty will help attendees:

    • Implement strategies that provide commercial entry into lucrative emerging markets;
    • Replenish product pipelines via partnering and co-promotions;
    • Weigh when a partnership is the best entry vehicle vs. a full M&A;
    • Evaluate viable IP protections in developing regions;
    • Minimize risks through effective partner selection and due diligence;
    • Structure transactions to meet both parties' potential objectives;
    • Incorporate what worked in recent deals in India, China and other countries into your negotiations;
    • Integrate your company's product life cycle strategies into your business plan; and
    • Avoid traps that can undermine global collaborations and lead to alliance failures.
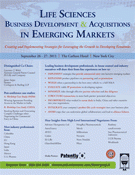
    ACI Brochure In particular, ACI's faculty will offer presentations on the following topics:

    • Emerging markets outlook: Surveying the fast-changing business landscape and learning from recent deal-making;
    • Identifying the right structure for new deals and negotiating the terms successfully;
    • Maximizing returns by finding the right partner in an emerging market;
    • Discerning what IP protection is available in an emerging market and conducting effective & strategic IP due diligence;
    • Focus session: Successfully implementing a growth strategy in Latin America;
    • Adjusting traditional M&A strategies to capitalize on opportunities in emerging markets;
    • Doing business with an emerging market company seeking access to the U.S. market: cultural, business, and regulatory challenges;
    • Integrating successful life cycle strategies with your business development approach to emerging markets;
    • Overcoming amplified barriers to effective alliance management; and
    • Demystifying the unique legal and regulatory challenges in entering into early stage research partnerships in emerging markets.

    Two post-conference case studies will be offered on September 27, 2011.  The first case study is entitled "Analyzing Winning Agreements Providing Access to the Market in India," and the second case study is entitled "Breaking Barriers and Overcoming Complex Challenges when Negotiating for Commercial Entry in China."
    ACI - American Conference Institute An agenda for the conference can be found here, and a complete brochure for this conference, including an agenda, detailed descriptions of conference sessions, list of speakers, and registration form can be obtained here.

    The registration fee is $2,295 (conference alone), $2,795 (conference and one workshop), or $3,295 (conference and both workshops).  Those registering by August 19, 2011 will receive a $200 discount.  Those interested in registering for the conference can do so here, by calling 1-888-224-2480, or by faxing a registration form to 1-877-927-1563.
  • By James DeGiulio —

    Drug Wholesalers Secure Injunction Blocking Braintree from Terminating MiraLAX Supply

    Braintree Laboratories Direct purchasers of MiraLAX, who have brought a class action suit headed by Rochester Drug Cooperative ("RDC") against drugmaker Braintree Laboratories Inc., were successful in obtaining a preliminary injunction blocking Braintree from terminating its sales to the drug wholesaler plaintiffs.  In the original action, Braintree filed a patent infringement suit against Schwarz Pharma in 2003, after the company submitted an ANDA to manufacture a generic version of MiraLAX.  The suit and prior Paragraph IV filing, which concerned U.S. Patent No. 5,710,183, triggered a 30-month stay on the FDA's approval of Schwarz's ANDA.  Braintree eventually dismissed its claims against Schwarz Pharma, thus waiving the remainder of the 30-month stay, and the ANDA was approved in July 2004.

    Rochester Drug Cooperative In March of 2007, RDC filed this putative class action suit on behalf of drug wholesalers who purchase MiraLAX directly.  RDC claimed that the original ANDA litigation was a ploy to extend Braintree's monopoly on MiraLAX because Braintree knew or should have known the patent it asserted against Schwarz Pharma was invalid.  The plaintiffs claim Braintree engaged in an anti-competitive scheme that allowed it to charge supracompetitive prices for MiraLAX, or polyethylene glycol 3350, and the delay of generic competition caused them to overpay for MiraLAX by millions of dollars.  In lieu of an answer, Braintree filed a motion to dismiss the action, which was denied.  In March 2011, Braintree sent letters to the plaintiffs, stating that they would no longer sell products to them due to their allegedly meritless antitrust litigation.  RDC then moved for a preliminary injunction to force Braintree to continue to supply MiraLAX to the plaintiffs.

    MiraLax On July 7, Judge Sue Robinson issued a memorandum opinion granting the request for a preliminary injunction from plaintiffs RDC, Louisiana Wholesale Drug Co., and Meijer Inc., forcing Braintree to stop its retaliatory action and resume its supplier relationships with the companies.  After Braintree cut off its supply, RDC said the move threatened the plaintiffs' ability to obtain and retain customers, and forced them to switch to other suppliers that charged them higher prices.  Judge Robinson analyzed the four eBay factors, and was persuaded by plaintiff's authority Bergen Drug Company Inc. v. Parke, Davis & Co., 307 F.2d 725 (3d Cir. 1962).  Judge Robinson agreed to issue the injunction, but she said the plaintiffs would have to agree to post a bond for the cost of the litigation.  The plaintiffs have shown only possible harm, not any specific harm, caused by Braintree's decision to terminate its business relationships with the plaintiffs.  This is because ultimate relief sought by the plaintiffs is to be reimbursed for the cost of Braintree's branded MiraLAX product.  Judge Robinson said the plaintiffs must bear the risk of costs going forward if the plaintiffs seek the extra relief of Braintree's continued business.


    Takeda Agrees to Drop Mylan Inc. from Prevacid Patent Litigation, But Mylan Subsidiary Remains

    Takeda Takeda Pharmaceutical Co. Ltd. has agreed to drop Mylan Inc. as a defendant in Takeda's lawsuit over Mylan's generic version of the acid reflux drug Prevacid, but subsidiary Mylan Pharmaceuticals Inc. remains in the suit to litigate the generic maker's position.  Takeda brought suit in May 2011, accusing Mylan of infringing its two patents covering Prevacid, U.S. Patent Nos. 6,328,994 and 5,464,632, by filing an ANDA for a generic version of the drug (see "Court Report – Part II," May 23, 2011).  Ethypharm SA, which owns the '632 patent and licenses it to Takeda, joined Takeda as a plaintiff.  According to the complaint, the plaintiffs claimed the generic's release would cause irreparable harm, and asked the District Court to block approval of Mylan's generic drug.  Takeda also requested that the case be declared exceptional so that Mylan would be forced to cover its attorneys' fees.  After Takeda filed its complaint, Mylan filed counterclaims seeking declaratory judgment that the two patents were invalid and that Mylan's drug did not infringe either patent.

    Mylan #1 On July 11, Judge Joel Pisano signed off on a stipulation dismissing Mylan Inc. as a defendant.  However, the dismissal may not have a significant impact, for Mylan Inc. subsidiary Mylan Pharmaceuticals Inc. remains in the suit.  In exchange for the dismissal, Mylan and Matrix Laboratories agreed to abide by the District Court's final judgment relating to any damages, as well as any court orders against it.  Mylan also agreed to make any documents or information relevant to the alleged infringement and validity of Takeda's patents available for discovery as if the generics maker were still party to the suit.


    Astellas Settles Adenoscan Patent Suit with Sagent and Strides Arcolab

    Astellas After just a few weeks, Astellas Pharma US Inc. has settled a lawsuit against Sagent Pharmaceuticals Inc. and Strides Arcolab Ltd., who were accused of infringing Astellas' patent covering the pharmacologic stress agent Adenoscan.  In June 2011, Astellas and Item Development AB brought suit against Sagent Strides, a joint venture between the two defendant companies, alleging infringement of U.S. Patent No. 5,731,296 (see "Court Report," June 26, 2011).  Astellas was notified in May that Sagent Strides had filed papers seeking FDA approval for an Adenoscan generic.  Sagent Strides asserted that the '296 patent was invalid and unenforceable.  Astellas has vigorously defended the '296 patent, which is set to expire in March 2015, bringing several suits against generic makers seeking to enter the market for its Adenonscan product.  However, Astellas entered into a settlement with Teva in 2007, which provides Teva with a license to market and sell its Adenoscan generic in September 2012.

    On July 11, Judge Mary Cooper signed off on an order dismissing the case against Sagent Strides without prejudice.  The action can be reopened if there is no settlement within 60 days, and the District Court retains jurisdiction over the matter to enforce the terms of the settlement, which were not disclosed.  It is not clear whether the settlement included a provision that would allow Sagent Strides to market and sell its generic Adenoscan product before the expiration of the patent.

  • By Kwame Mensah —

    MultiCell Technologies Recently, MultiCell Technologies, Inc. of Woonsocket, RI, a clinical-stage biopharmaceutical company focusing on unmet medical needs for the treatment of neurological disorders, hepatic disease, and cancer, announced that it had been issued U.S. Patent No. 7,935,528, which is related to the isolation and use of human liver stem cells to treat liver disease.  MultiCell Technologies is the worldwide exclusive licensee of the '528 patent, which is assigned to Rhode Island Hospital.

    The '528 patent describes methods to isolate and use human liver stem cells to treat degenerative liver diseases or inherited deficiencies of liver function.  MultiCell Technologies believes that one of the benefits of the claimed methods is that liver stem cells, which differentiate into functional mature hepatocytes, can be used to reconstitute a diseased liver.  This provides a useful alternative approach to whole organ transplants for certain liver diseases.

    W. Gerald Newmin, Chairman and Chief Executive Officer of MultiCell Technologies, stated that the liver stem cells of the '528 patent would help the company "better understand liver stem cell biology, and the role these cells play in chronic liver disease and primary hepatocellular carcinoma."

    Claim 1, which is the lone independent claim of the '528 patent, recites:

    A primary liver stem cell, wherein said stem cell (a) is obtained from normal liver tissue, (b) is isolated from an isolated liver cell cluster comprising a hepatocyte and said stem cell, wherein said hepatocyte and said stem cell are joined by a desmosomal junction, (c) expresses OV6 [an oval cell marker], and (d) does not express OC2 [an oval cell marker], wherein said stem cell comprises a DNA encoding a heterologous protein. 

  •     By James DeGiulio —

    FDA The Food and Drug Administration has been the subject of recent criticism from drugmaker advocates who allege that the FDA's tougher safety standards have slowed down the pace of drug approvals and have hurt the drug industry.  However, despite these allegedly repressive safety standards, the head of the FDA's drug division, Dr. Janet Woodcock, speculated in May that the FDA would approve more drugs in 2011 than the number of new drugs approved in 2010 ("FDA official sees drug approvals rising").  Barring some unforeseen event, it appears likely that this year, the FDA will easily surpass the number drug approvals for 2010. The FDA has already approved 20 new drugs so far, excluding approvals granted for vaccines or new formulations or uses of existing drugs, almost matching the 21 drugs approved in 2010, according to a recent article in the Wall Street Journal ("FDA's Drug-Approval Rate for 2011 on Pace to Exceed 2010"; subscription required for full access).

    These numbers should quiet critics of the safety standards for the time being.  Industry advocates and some lawmakers have criticized the FDA for instituting higher burdens on drugmakers to win approval.  The FDA has tightened safety regulations since the 2004 market withdrawal of the painkiller Vioxx.  After the Vioxx withdrawal, the FDA was criticized by the public for lack of vigilance, and the agency responded by re-evaluating its practices.

    Last week, The House Energy and Commerce's health subcommittee held a panel hearing to discuss the reauthorization of legislation that allows the FDA to charge drug companies user fees (which is expected to be renewed, as it is supported by both parties).  However, during the panel, Jonathan Leff, a managing director at Warburg Pincus LLC, a private-equity and venture-capital firm, provided written testimony asserting that the FDA's tougher focus on drug safety has hurt the drug industry and is a major contributor to the rising cost and length of drug development.  Mr. Leff stated that despite potential for producing a marketable and beneficial therapeutic, many promising scientific discoveries are not being developed into new treatments for disease due to lack of investment capital.  He cited figures showing venture investment in the U.S. life sciences industry declined by $2 billion from 2008 to 2010.

    Woodcock, Janet In response, Dr. Woodcock (at left) defended the FDA, insisting that the FDA has changed its requirements only because the new science required it.  She pointed instead to the high failure rates of drugs in the development stage, rather than the FDA approval stage, as presenting the main contributor to the rising cost and length of drug development.  Dr. Woodcock explained the agency meets more than 90 percent of deadlines that are part of the drug-review process.  She also said first cycle approvals are at a 20-year high, and more than two-thirds of new drugs are approved within the 6-to-10-month time frame given to new drug applications.  Dr. Woodcock admitted that there are certain cases where companies are asked to submit more information, which prompts additional reviews, and in other cases drugs are rejected.  She did note that expert advisory panels, which generally make determinations on new drugs, have been more difficult to assemble.  In 2007, the FDA tightened guidelines to minimize drugmaker industry ties that could sway a panelist's view.  Since many of the individuals with the most expertise are affiliated with drugmakers, they are ineligible to sit on these advisory panels due to conflicts.  Rather than relying on advisory panels for specific experimental drugs, Dr. Woodcock said she would prefer to see more panel discussions of general policy issues, such as approval standards for a specific area.

  • By Kwame Mensah —

    BIO International Convention Last month's BIO International Conference kicked off on June 27th with a presentation entitled "IP Issues Affecting Biomarker-Based Diagnostics."  The four panelists for the session were David Gass of Marshall, Gerstein & Borun, LLP; Dr. Thorlakur Jonsson, the Director of Intellectual Property at deCODE genetics ehf; Dr. Simon O'Brien of D. Young & Co. in the United Kingdom; and Dr. Jan Skouv, Director IPR at Exiqon A/S.

    The session began with two talks related to recent developments in technology related to biomarkers.  The first speaker was Dr. Jonsson, who focused on the use of the information provided by genome sequencing in personalized medicine.  Dr. Jonsson began with a discussion of genetic variation in the human genome.  He stated that the mere 1% genetic variation among humans can determine whether an individual may be at risk for a certain disease, may have certain drug metabolism, or may respond in a certain way to specific therapies.  He referred to these as high risk Mendelian phenotypes, and touted the use of high throughput and microarray assays as providing faster analysis of genomic sequences.  Dr. Jonsson concluded his presentation with a short summary of the work done by deCODE to investigate biomarkers for many diseases through correlation analyses of genealogy, genotype, phenotype, and genome databases.

    The second speaker of the session was Dr. Skouv who discussed detection of microRNAs (miRNAs; RNA molecules of 22 nucleotides or less in length) in blood samples as biomarkers.  These molecules have been found to be highly involved in the pathology of several types of cancer.  Dr. Skouv stated that although the miRNA molecules were generally found to be stable when associated with blood serum and its components after EDTA treatment, introduction of locked nucleic acids (LNAs) to the miRNAs via PCR further increased stability of the miRNAs.  LNAs are a class of nucleic acid analogues in which the ribose ring is "locked" by a methylene bridge connecting the 2'-O atom and the 4'-C atom of the ribose ring, which increases the molecule's affinity for other molecules.  Dr. Skouv concluded his presentation by noting that Exiqon had successfully used LNA-associated miRNAs as biomarkers for the early detection of colorectal cancer, and that preliminary work had been done on the early detection of melanoma.

    The next two speakers focused on patent-related issues.  Generally speaking, two types of intellectual property protection exist for detection and use of biomarkers:  (1) protection for the markers themselves (nucleic acid molecules, proteins, etc.) and (2) protection for use of the markers for diagnostics and therapeutics, including compositions, methods, systems, and kits.  The first of the two speakers, David Gass provided a general discussion of the legal requirements for receiving a patent, and then discussed recent decisions in the federal courts within the last year relating to the patent eligibility of nucleic acid molecules and processes of analysis.  He then moved to a discussion of two cases of particular relevance to biomarkers; Bilski v. Kappos and AMP v. USPTO (the "Myriad" case).  Mr. Gass highlighted the fact that the Supreme Court in Bilski held that the "machine or transformation" test is not the exclusive test for determining whether a process or method of analysis is patent eligible.  This is of particular relevance to the use of biomarkers because this holding indicates that determinative processes involved in the use of biomarkers do not necessarily have to be associated with a machine or be "transformative" to be patentable subject-matter.

    Mr. Gass then moved to a discussion of the Myriad case, highlighting that the claims to isolated DNA molecules (gene products) were found invalid under 35 U.S.C. § 101, but that the claims related to vectors, host cells, probes. i.e., man-engineered DNA-type molecules were left intact.  He specifically highlighted the amicus brief submitted during the appellate proceedings by the Department of Justice.  The DOJ brief focused on the difference in the character of the nucleic acid molecules themselves, specifically asserting that "naturally occurring" nucleic acid molecules were not patentable subject-matter, whereas engineered nucleic acid molecules (e.g, vectors, host cells, cDNA) were patentable subject-matter.  Mr. Gass then touched on the jurisdiction issues that were discussed during oral argument before the Federal Circuit.  The panel also inquired about what it means to isolate a gene, i.e., that the breaking of covalent bonds, under traditional chemistry, creates a new molecule.  Mr. Gass indicated that this line of inquiry suggests that the Federal Circuit could determine that isolated DNA is different from naturally-occurring DNA, therefore making it patentable subject-matter.  This no doubt would be a big victory for the biotech industry.  Mr. Gass concluded his discussion by focusing on diagnostic claims such as those in the Myriad patents.  As discussed in this space previously, Myriad has argued that its diagnostic claims are not abstract and that each laboratory step is transformative.

    The second patent law-related speaker of the afternoon was Dr. Simon O'Brien.  He began his talk by stating that there has been a drastic decrease in the number of patent applications for gene patents since 2000.  In contrast, applications for diagnostic methods have increased almost as dramatically, and the majority of these applications have been related to cancer diagnostics.  Dr. O'Brien then gave a brief summary of the requirements for receiving a diagnostic method patent in Europe.  The necessary steps in a claim are:  examination, comparison, finding of a derivation, and attribution to a disease.  Only one of these steps need be in vitro.  He then pointed out that Myriad's BRCA-1 patents were found to be valid in opposition proceedings in Europe due to the fact that because isolated DNA molecules are patentable subject-matter, use of these molecules in diagnostic steps was also patentable.

    Mr. Gass then returned to the microphone to discuss multi-party and multi-jurisdictional issues involved in enforcement.  These are related to actions by multiple parties, in different jurisdictions or countries in performing a claimed diagnostic method.  He focused on several cases that may provide some guidance as to what issues may arise regarding the enforcement of claims for biomarker-based diagnostic tests.  One case discussed was BMC Resources, Inc. v. Paymentech, L.P. 478 F.3d 1373 (Fed. Cir. 2007), in which the Court stated that all steps of a method must be performed by one party in order for the claimed method to be infringed, unless a single mastermind is orchestrating performance of the method.  Mr. Gass surmised that for a diagnostic method claim to be reasonably enforceable, one needs to draft a diagnostic method claim such that the method can only be performed by one party.  Another case Mr. Gass discussed was Centillion Data Systems, LLC v. Qwest Communications International, Inc. (Fed. Cir. 2011), where the claim at issue was to a system with various components.  The Court in Centillion stated that a party must put the system into service, control the system as a whole, and obtain a benefit from the system to infringe.  Regarding territoriality issues, Mr. Gass again provided some insight into how this could affect biomarker/diagnostic claims stating that one or more steps of a method performed offshore would probably not be infringing, whereas one or more steps of a system performed offshore would be.  Mr. Gass recommended using various types of diagnostic method claims within the same claim set to increase the likelihood of securing reasonably enforceable claims.

    Dr. O'Brien concluded the session with a discussion of issues related to performing certain steps of a diagnostic method in another country as a means to circumvent infringement.  He echoed Mr. Gass' suggestion that claims be drafted such that it is necessary for a method to be performed by the same person, such as by limiting the number of steps in the method.  He stated that in Europe, one court determined that a patent holder may only receive protection for a method related to the DNA when the DNA is performing its native function.  For example, using a cancer probe to diagnose a different disease would likely not be infringing.

    Overall the session's discussion of use of biomarkers in diagnostic methods provided guidance for patent prosecutors in drafting good claims, and provided an excellent summary of the state of the industry related to biomarkers.

  • By Donald Zuhn —

    Reports Suggest Senate Action on Patent Reform Bill Could Be Weeks Away

    IPO #2 In Monday's IPO Daily News, the Intellectual Property Owners Association (IPO) suggested that H.R. 1249, which the House passed on June 23 (see "House Passes H.R. 1249"), might not be brought up by the Senate until after Congress' August recess, which is scheduled to begin on August 6.  The IPO report also noted that while Sen. Patrick Leahy (D-VT) is seeking Senate passage of the bill passed by the House, other Senators are objecting to a few provisions in the bill, including § 37 (the so-called MedCo amendment; see "The Medicines Company Clearing Last Hurdles to Patent Term Extension for Angiomax®"), which would amend 35 U.S.C. § 156(d)(1) by adding the following text:

    For purposes of determining the date on which a product receives permission under the second sentence of this paragraph, if such permission is transmitted after 4:30 P.M., Eastern Time, on a business day, or is transmitted on a day that is not a business day, the product shall be deemed to receive such permission on the next business day.  For purposes of the preceding sentence, the term 'business day' means any Monday, Tuesday, Wednesday, Thursday, or Friday, excluding any legal holiday under section 6103 of title 5.

    Wegner, Harold On Saturday, Hal Wegner (at left) stated in his e-mail newsletter that "[a]n early Senate vote is now considered unlikely on the Leahy-Smith America Invents Act, with the strong likelihood that any full Senate consideration will take place at the earliest after Labor Day" (emphasis in original).

    A copy of the House bill as it was received in the Senate and placed on the Senate legislative calendar on June 27 can be found here.


    Economist Asks Congress to "Let the Patent Office Keep Its Money"

    Shane, Scott In an Op-Ed published last week by Bloomberg Businessweek, Economist Dr. Scott Shane contends that "[f]ee diversion is wreaking havoc on the patent system and undermining our national competitiveness" ("Let the Patent Office Keep Its Money").  Dr. Shane (at right), a Professor of Economics at Case Western Reserve University, should be familiar to Patent Docs readers for his contributions to the patent reform debate (particularly with respect to patent damages) in 2009 (see "Manufacturing Alliance on Patent Policy: Apportionment of Damages Provision Will Have Adverse Effects"; "MAPP Economist Advises Congress on Damages Provision"; and "Patent Reform Efforts Opposed by Small Business Group").

    Noting that between 2000 and 2010 the annual number of applications filed rose 65%, Dr. Shane argues in his article that "[t]o meet this demand, the USPTO needs more patent examiners to handle the greater volume of applications," and that "[b]y taking away a portion of the patent office's fees, Congress has prevented the USPTO from hiring enough examiners to evaluate patent applications in a timely manner."  He concludes that "[i]f the House adopts the Senate's version [of the USPTO funding provisions], fee diversion will end, but if the Senate adopts the House's version, then I'm much less optimistic that the USPTO will get the money it needs to operate successfully."


    American Action Forum Urges Senate to Retain USPTO Funding Provisions of S. 23

    American Action Forum The American Action Forum, an organization that describes itself as "a forward-looking policy institute dedicated to keeping America strong, free and prosperous," which "seeks to promote common-sense, innovative, and solutions-based policies that will reform government, challenge out-dated assumptions, and create a smaller, smarter government that will serve its citizens better," issued a statement today calling on Senate leadership to prevent fee diversion in the U.S. Patent and Trademark Office.  American Action Forum President Doug Holtz-Eakin, a former Director of the Congressional Budget Office, pointed out that "[e]ach year over $800 million in user fees are diverted away from the USPTO through the Congressional appropriations process."  He urged Senate Majority Leader Harry Reid (D-NV) and Minority Leader Mitch McConnell (R-KY) "to adopt the language in S.23, a bill that will effectively end this process."

    In a letter to Sen. Reid and Sen. McConnell, Mr. Holtz-Eakin stated that "[t]he establishment of the Patent and Trademark Fee Reserve Fund in H.R. 1249 would be ineffective in stopping the diversion of fees from USPTO."  In contrast, he explained that "the Senate bill [S. 23] creates a USPTO Public Enterprise Fund at the Treasury Department, which would receive all fees paid to USPTO and which would be available to USPTO when needed," stating that "[t]he funds would not be funneled through the Congressional appropriations process and would, therefore, not be exposed to the threat of being diverted to general government spending."  "If it is, in fact, the goal of Congress to end fee diversion and ensure that the USPTO is authorized to spend the fees it collects from its users," Mr. Holtz-Eakin contended that "the language contained in S.23 is clearly superior to the language of H.R.1249."

  •     By Donald Zuhn —

    Reckitt Benckiser Last week, in Reckitt Benckiser Inc. v. Watson Laboratories, Inc. – Florida, the Federal Circuit affirmed a decision by the District Court for the Southern District of Florida holding that Defendant-Appellee Watson Laboratories, Inc. – Florida did not infringe the asserted claims of U.S. Patent No. 6,372,252.  The Federal Circuit also determined that the District Court correctly construed the asserted claims of the '252 patent.

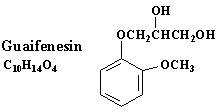
    Guaifene Plaintiff-Appellant Reckitt Benckiser Inc. markets bilayer tablets comprising guaifensin, an expectorant used to relieve congestion, as Mucinex®.  Reckitt's bilayer tablets contain guaifensin in both immediate release (IR) and sustained release (SR) formulations.  In obtaining FDA approval for its Mucinex® products, Reckitt listed the '252 patent in the Orange Book as covering the guaifensin tablets.

    Watson Pharmaceuticals Seeking approval to market guaifensin tablet formulations, Watson filed an Abbreviated New Drug Application (ANDA) with the FDA.  In response to Watson's ANDA filing, Reckitt brought suit against Watson for infringement of claims 24, 26-28, 31-34, 39, 57, and 58 of the '252 patent.  Representative claim 24 reads (emphasis added by panel):

    24.  A modified release product having two portions, wherein a first portion comprises a first quantity of guaifenesin in an immediate release form which becomes fully bioavailable in the subject's stomach and a second portion comprises a second quantity of guaifenesin in a sustained release form wherein the ratio of said first quantity to said second quantity provides a Cmax in a human subject equivalent to the Cmax obtained when the first of three doses of a standard immediate release formulation having one third the amount of guaifenesin is dosed every four hours over a 12 hour period and wherein said product also provides therapeutically effective bioavailability for at least twelve hours after a single dose in a human subject according to serum analysis.

    In construing the term "portion," which appears in each of the asserted independent claims, the District Court determined that the term means "a discrete part of the product."  Following a bench trial, the District Court found that Watson's products, which are non-layered polymer matrix tablets made from a single guaifenesin formulation, do not have separate IR and SR portions, and therefore do not literally infringe the asserted claims of the '252 patent.  The District Court also found no infringement under the doctrine of equivalents because Reckitt disclaimed products lacking two discrete structural portions during prosecution, and Watson's products lack two discrete structural portions.

    In its opinion, the Federal Circuit noted that the prosecution history of the '252 patent was relevant to the parties' arguments on appeal.  The Court pointed out that as originally filed, claims 1-11 were directed to a SR composition, claims 12-24 were limited to products with two portions, and claims 25-32 were directed to modified release tablets.  In response to a rejection of claims 1-32 for obviousness, applicants cancelled the claims and added new claims 33-55, stating that the new claims were directed to "[a] modified release tablet having two portions," and adding that "the sustained release claims have been cancelled to facilitate prosecution."  Applicants also distinguished the new claims over the cited prior art, stating that one reference "does not disclose a composition having both an immediate release portion . . . and a sustained release portion," and that the other reference, despite disclosing tablets with two separate portions, "specifically teaches away from employing any immediate realease portion."

    On appeal, Reckitt argued that the District Court erred in construing "portion" and in finding no infringement either literally or under the doctrine of equivalents.  In determining that the District Court did not err in construing the term "portion" to mean "a discrete part of the product," the Federal Circuit noted that:

    The applicants disavowed claim coverage of sustained release tablets by cancelling original claims 1-11 and remarking to the examiner that "[o]riginal claims 1-11 were directed to a sustained release formulation . . . . [T]he sustained release claims have been cancelled to facilitate prosecution."  The unmistakable effect of that disavowal, evident from the applicants' remarks distinguishing the prior art, was to limit the remaining claims to two-portion guaifenesin products [citations omitted].

    The panel determined that "[t]he file history thus demonstrates a 'clear and unambiguous' prosecution disclaimer."

    As for the District Court's finding of no infringement, the Federal Circuit stated that:

    As the court correctly found, Watson's products do not infringe because they are non-layered, single-formulation polymer matrix tablets that do not contain the claimed "first portion" or "second portion."  . . .  The district court correctly concluded that Watson's products do not have two structural portions and that guaifenesin granules on the surface of Watson’s tablets do not constitute the claimed first portion of guaifenesin in an IR form.

    With respect to infringement under the doctrine of equivalents, the panel concluded that:

    [O]n the facts of this case, prosecution history estoppel bars Reckitt from recapturing single-formulation SR guaifenesin tablets like those it disclaimed in obtaining the '252 patent.  As the district court correctly noted, Reckitt's narrowing claim amendments were made for reasons of patentability [citations omitted].

    The Federal Circuit therefore affirmed the District Court's finding of no infringement, either literally or under the doctrine of equivalents.

    Reckitt Benckiser Inc. v. Watson Laboratories, Inc. – Florida (Fed. Cir. 2011)
    Nonprecedential disposition
    Panel: Circuit Judges Lourie, Linn, and Dyk
    Opinion by Circuit Judge Lourie

  • By Sherri Oslick —

    Gavel About Court Report:  Each week we will report briefly on recently filed biotech and pharma cases.

    Ferring B.V. v. Watson Pharmaceuticals, Inc. et al.
    3:11-cv-00481; filed July 7, 2011 in the District Court of Nevada

    • Plaintiff:  Ferring B.V.
    • Defendants:  Watson Pharmaceuticals, Inc.; Watson Laboratories, Inc.; Watson Laboratories, Inc. – Florida; Watson Pharma, Inc.

    Infringement of U.S. Patent No. 7,947,739 ("Tranexamic Acid Formulations," issued May 24, 2011) following a Paragraph IV certification as part of Watson's filing of an ANDA to manufacture a generic version of Ferring's Lysteda® (tranexamic acid, used to treat heavy menstrual bleeding).  View the complaint here.


    Intervet Inc. et al. v. Boehringer Ingelheim Vetmedica Inc.

    1:11-cv-00595; filed July 6, 2911 in the District Court of Delaware

    • Plaintiffs:  Intervet Inc. d/b/a Merck Animal Health; Arizona Board of Regents
    • Defendant:  Boehringer Ingelheim Vetmedica Inc.

    Infringement of U.S. Patent No. 5,610,059 ("Etiological Agent for Porcine Enteritis," issued March 11, 1997) based on Boehringer's manufacture and sale of its Enterisol® Ileitis product.  View the complaint here.


    Shire LLC et al. v. Roxane Laboratiores, Inc.

    2:11-cv-03886; filed July 6, 2011 in the District Court of New Jersey

    • Plaintiffs:  Shire LLC; Shire Development Inc.
    • Defendant:  Roxane Laboratiores, Inc.

    Shire LLC et al. v. Sandoz Inc.
    3:11-cv-03787; filed June 30, 2011 in the District Court of New Jersey

    • Plaintiffs:  Shire LLC; Shire Development Inc.
    • Defendant:  Sandoz Inc.

    Shire LLC et al. v. Amneal Pharmaceuticals, LLC
    3:11-cv-03781; filed June 30, 2011 in the District Court of New Jersey

    • Plaintiffs:  Shire LLC; Shire Development Inc.
    • Defendant:  Amneal Pharmaceuticals, LLC

    The complaints in these cases are substantially identical.  Infringement of U.S. Patent Nos. 7,105,486 ("Abuse-resistant amphetamine compounds," issued  September 12, 2006), 7,223,735 ("Abuse resistant lysine amphetamine compounds," issued  May 29, 2007), 7,655,630  ("Abuse-resistant amphetamine prodrugs," issued  February 2, 2010), 7,659,253 ("Abuse-resistant amphetamine prodrugs," issued  February 9, 2010), 7,659,254 ("Abuse-resistant amphetamine prodrugs," issued  February 9, 2010), 7,662,787 ("Abuse resistant lysine amphetamine compounds," issued  February 16, 2010), 7,671,030 ("Abuse-resistant amphetamine prodrugs," issued  March 2, 2010), 7,671,031 ("Abuse-resistant amphetamine prodrugs," issued  March 2, 2010), 7,674,774 ("Abuse-resistant amphetamine prodrugs," issued  March 9, 2010), 7,678,770 ("Abuse-resistant amphetamine prodrugs," issued  March 16, 2010), 7,678,771 ("Abuse-resistant amphetamine prodrugs," issued  March 16, 2010), 7,687,466 ("Abuse-resistant amphetamine prodrugs," issued  March 30, 2010), 7,687,467 ("Abuse-resistant amphetamine prodrugs," issued  March 30, 2010), 7,700,561 ("Abuse-resistant amphetamine prodrugs," issued  April 20, 2010), 7,718,619 ("Abuse-resistant amphetamine prodrugs," issued  May 18, 2010), 7,723,305 ("Abuse-resistant amphetamine prodrugs," issued  May 25, 2010) following a Paragraph IV certification as part of Sandoz's filing of an ANDA to manufacture a generic version of Shire's Vyvanse® (lisdexamfetamine dimesylate, used to treat Attention Deficit Hyperactivty Disorder).  View the Sandoz complaint here.  [NB: The complaint against Amneal does not assert U.S. Patent No. 7,659,254.]


    Alcon Pharmaceuticals Ltd. et al. v. Lupin Ltd. et al.

    1:11-cv-00587; filed July 1, 2011 in the District Court of Delaware

    • Plaintiffs:  Alcon Pharmaceuticals Ltd.; Alcon Research Ltd.
    • Defendants:  Lupin Ltd.; Lupin Pharmaceuticals Inc.

    Infringement of U.S. Patent Nos. 6,716,830 ("Ophthalmic Antibiotic Compositions Containing Moxifloxacin," issued April 6, 2004) and 7,671,070 ("Method of Treating Ophthalmic Infections with Moxifloxacin Compositions," issued March 2, 2010) following a Paragraph IV certification as part of Lupin's filing of an ANDA to manufacture a generic version of Alcon's Vigamox® (moxifloxacin, used to treat bacterial conjunctivitis).  View the complaint here.


    Orexo AB v. Mylan Pharmaceuticals Inc. et al.

    3:11-cv-03788; filed June 30, 2011 in the District Court of New Jersey

    • Plaintiff:  Orexo AB
    • Defendants:  Mylan Pharmaceuticals Inc.; Mylan, Inc.

    Infringement of U.S. Patent No. 6,761,910 ("Pharmaceutical Composition for the Treatment of Acute Disorders," issued July 13, 2004), licensed to Meda Pharmaceuticals, following a Paragraph IV certification as part of Mylan's filing of an ANDA to manufacture a generic version of Meda's Edluar (zolpidem sublingual tablets, used to treat short-term insomnia).  View the complaint here.

  • Calendar

    July 16-19, 2011 – 2011 Annual Meeting; The Nuts & Bolts of Patent Prosecution Practice; and 2011 Patent Practice Update (National Association of Patent Practitioners) – Las Vegas, NV

    July 18-19, 2011 – Hatch-Waxman Boot Camp*** (American Conference Institute) – San Diego, CA

    July 20-21, 2011 – Advanced Forum on Life Sciences Collaborative Agreements and Acquisitions*** (American Conference Institute) – San Francisco, CA

    July 25-27, 2011 – Intensive Patent Law Seminar (Chisum Patent Academy) – Seattle, WA

    July 26, 2011 – The New Inequitable Conduct Standard in Patent Litigation: Asserting and Defending Inequitable Conduct Challenges After the Landmark Therasense Decision (Strafford) – 1:00 – 2:30 PM (EDT)

    August 4-7, 2011 – 2011 Annual Meeting (American Bar Association) – Toronto, Ontario

    August 15-19, 2011 – Intellectual Property Law Summer School 2011 (IBC Legal) – Cambridge, UK

    August 16-18, 2011 – The Bilski Impact: Procuring & Enforcing Software, Business Methods & Bioinformatics Patents (Patent Resources Group) – Alexandria, VA and Southfield, MI

    September 6-7, 2011 – Pharmaceutical Law Academy (IBC Legal Conferences) – London, UK

    September 11-13, 2011 – 2011 Annual Meeting (Intellectual Property Owners Association) – Los Angeles, CA

    September 14-16, 2011 – 85th Annual Meeting (Intellectual Property Institute of Canada) – Chicago, IL

    September 18-20, 2011 – Accelerating Intellectual Property and Innovation in South Africa*** (South African Department of Science and Technology) – Cape Town, South Africa

    September 22-23, 2011 – FDA Boot Camp*** (American Conference Institute) – Boston, MA

    September 26-27, 2011 – Biosimilars and Biobetters*** (SMi Group) – London, UK

    ***Patent Docs is a media partner of this conference or CLE

  • London The SMi Group will be holding its 2nd conference on Biosimilars and Biobetters from September 26-27, 2011 in London, UK.  The conference will offer presentations on the following topics:

    • The market for biosimilars: Unraveling the mystery;
    • The EMA scientific advice compared with EMA guidelines;
    • Industry interpretation the new guideline on biosimilar mAbs;
    • Next generation monoclonal antibody development and clinical challenges with biosimilar drug development;
    • EU litigation considerations and update;
    • The ins and outs of the biosimilar regulatory pathway;
    • The developing biosimilars regulatory framework in the U.S.;
    • U.S. patent issues in biosimilars;
    • Biosimilars: Patent protection issues;
    • Maintaining a consistent quality standard of biosimilars;
    • From imitation to innovation — Biosimilars are not generics;
    • Making things last longer for less;
    • Pharmacovigilance & biosimilar monoclonal antibodies: Practical considerations;
    • Factors to be considered for biosimilar CMC development;
    • GCSF biosimilar case study;
    • Biosimilar sales forecast in seven unregulated markets in 2016: How large is the opportunity?
    • Opportunities to drive your longer term biosimilars strategy — India and emerging market perspective.

    Banner Two post-conference workshops will be offered on September 28, 2011.  The first workshop, entitled "How similar is similar?" will cover a variety of factors that dictate and determine exactly how similar is similar.  The second workshop, entitled " A Global View of Biosimilars: Regulatory Analysis and Pricing Structure Case Studies," will examine the potential of companies with a strong biosimilars portfolio in emerging markets to expand into highly-regulated markets via alliances, M&A deals and utilization of post-marketing data.

    The programme for the Biosimilars and Biobetters conference can be found here (Day 1), here (Day 2), and here (workshops).  A complete brochure for this conference can be obtained here.

    SMi Group The registration fee for the conference is £1,399 and for each workshop is £599.  Those registering for the conference by July 15, 2011 will receive a £200 discount.  Those interested in registering for the conference and workshops can do so here.

    Patent Docs is a supporting publication of the Biosimilars and Biobetters conference.