• Chicago #5 American Conference Institute (ACI) will be holding its FDA Boot Camp Devices Edition conference on October 25-26, 2011 in Chicago, IL.  ACI faculty will help attendees:

    • Master the basics of the application and approval processes, including 510(k) clearance and PMAs;
    • Navigate the complexities of device regulations;
    • Comprehend the structure of the FDA and the roles of the three major agency centers:  CDER, CBER, and CDRH;
    • Develop a practical working knowledge of clinical trials and IDEs;
    • Learn how devices are classified, monitored, and regulated;
    • Recognize the pivotal role of labeling and learn how to avoid misbranding and off label promotion;
    • See the importance of cGMPs and QSRs to the post-approval regulatory process; and
    • Navigate the protocols of adverse events monitoring, product withdrawals, and recalls.

    Brochure FDA Boot In particular, ACI's faculty will offer presentations on the following topics:

    • The basics: Understanding and working with the FDA — jurisdiction, functions, organization, and operations;
    • Overview of device regulation;
    • Understanding clinical trials and the investigational device exemption (IDE);
    • Navigating the 510(k) clearance process;
    • Comprehending the premarket approval process (PMA);
    • Complying with general post-market controls;
    • Understanding FDA enforcement tools, policies, practices, and trends;
    • Labeling and promotion;
    • Medical device reporting;
    • Guidance for navigating recalls and withdrawals; and
    • Quality system regulation.

    A pre-conference training session on the "Fundamentals of FDA Regulatory Law," will be offered on October 24, 2011 from 1:00 – 5:00 pm.  The workshop will provide a basic overview of FDA regulations and will prepare attendees for the in-depth discussions that will take place throughout the conference.

    Two concurrent post-conference master classes will be offered from 2:00 – 5:00 pm on October 26, 2011.  The first master class, entitled "Staying Ahead of the Curve: FDA's Regulation of In Vitro Diagnostics and Laboratory Developed Tests," will provide an in-depth overview of the FDA regulatory structure governing IVDs, and the coming changes that will regulate LDTs.  The second master class, entitled "Software as a Medical Device: Understanding the Nuances of FDA Regulation," will address issues that are encountered when dealing with software classified as medical devices.

    A complete brochure for this conference, including an agenda, detailed descriptions of conference sessions, list of speakers, and registration form can be obtained here.

    ACI - American Conference Institute The registration fee is $2,295 (conference alone), $2,895 (conference and training session or conference and one master class), or $3,295 (conference, training session, and one master class).  Those registering by August 25, 2011 will receive a $300 discount and those registering by September 22, 2011 will receive a $200 discount.  In addition, Patent Docs readers who reference the discount code "PD 200" will receive $200 off the current price tier when registering.  Those interested in registering for the conference can do so here, by calling 1-888-224-2480, or by faxing a registration form to 1-877-927-1563.

    Patent Docs is a media partner of ACI's FDA Boot Camp Devices Edition conference.

  • Brochure EU Pharma C5 will be holding its 2nd Annual Forum on European Pharmaceutical Regulatory Law on September 21-22, 2011 in Brussels, Belgium.  The conference will provide attendees with practical and strategic guidance on the most current regulatory developments impacting on the pharma sector, including presentations on the following topics:

    • Interpreting the pharmacovigilance rules and the impact on reporting obligations;
    • Reviewing pharma regulatory requirements and the interaction with competition law;
    • The inclusion of environmental aspects into the EU legislation on good manufacturing practice (GMP);
    • Obtaining and maintaining an effective pricing and reimbursement structure;
    • The evolution of regulation and the drug approval system in the 21st century;
    • The do's and don'ts in multi-jurisdiction clinical trials: Where are we and where are we going?
    • Adopting an effective product labeling model to combat anti-counterfeiting;
    • Implementation of the UK bribery act and how pharma company practices will need to change;
    • The rise in social media: Balancing information to patients and advertising to ensure compliance;
    • Avoiding product liability litigation by reviewing global labeling practices;
    • The current U.S. approval process in light of recent regulatory and legislative developments;
    • Overcoming obstacles to ehealth and personalized medicine;
    • Successfully obtaining SPCs and extensions of regulatory data protection: Extending market exclusivity (panel session);
    • Guidance on the regulations applicable to advanced therapy medicinal products; and
    • Privilege and discovery in pharma litigation.

    A post-conference workshop on "Adopting Effective Risk Management Strategies to Avoid Product Liability Claims," will be offered from 9:00 am to 12:30 pm on September 23, 2011.  The workshop will provide in-depth guidance on devising strategic risk management plans to ensure that internal processes and drug standards comply with drug safety requirements across Europe and the U.S., and equip attendees with the practical and strategic tools needed to obtain sustainable market access in Europe and the U.S. to avoid costly product liability claims.

    The agenda for the European Pharmaceutical Regulatory Law Boot Camp can be found here.  A complete brochure for this conference, including an agenda, description of the sessions, list of speakers, and registration form can be downloaded link.

    C5 The registration fee for this conference is €1899 (conference alone) or €2398 (conference and workshop).  Patent Docs readers who reference the discount code "PD 200" will receive €200 off the current price tier when registering.  Those interested in registering for the conference can do so here, by calling +44 20 7878 6888, or by faxing a registration form to +44 20 7878 6885.

    Patent Docs is a media partner of ACI's Forum on European Pharmaceutical Regulatory Law.

  • By Donald Zuhn —

    Waxman, Henry Last week, a group of seven House Democrats, led by Rep. Henry Waxman (D-CA), sought to have the 12-year data exclusivity period provided by the biosimilar approval pathway of the Patient Protection and Affordable Care Act (PPACA) excluded from the Trans-Pacific Partnership (TPP) agreement.  According to a report in The Hill, the legislators sent a letter to President Obama, asking him to exclude the 12-year period from the TTP's intellectual property provisions.

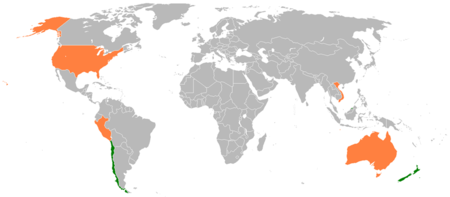
    The TTP, or Trans-Pacific Strategic Economic Partnership Agreement, is a multilateral free trade agreement currently being negotiated by Australia, Brunei, Chile, Malaysia, New Zealand, Peru, Singapore, the United States, and Vietnam (Canada, Japan, the Philippines, South Korea, and Taiwan have also expressed interest in participating in the agreement).  The seventh round of TTP negotiations was held in Ho Chi Minh City, Vietnam, in June, and two more rounds of negotiations are scheduled to take place in September (in the United States) and in October (in Peru).  The nine TPP members have set a goal of reaching the outlines of an agreement by November of this year.

    In the letter sent to the President, the House members stated that "[w]ere the TPP to ultimately contain a 12 year biologics exclusivity provision, it would impede the ability of Congress to achieve the Administration's proposed 7 year change without running afoul of U.S. trade obligations."  The legislators added that "[w]e see no reason for the United States to agree to such a provision, much less propose it."  Prior to passage of the PPACA in March of last year, Rep. Waxman had introduced biosimilar legislation in the House that would, like the Hatch-Waxman regulatory pathway for small molecule generic drugs, provide 5 years of exclusivity for generic biologics (see "Waxman Introduces Follow-on Biologics Bill").  In addition, the Obama Administration, via the Office of Management and Budget (OMB), released a letter in June of 2009 that advocated for a 7 year exclusivity period (see "White House Recommends 7-Year Data Exclusivity Period for Follow-on Biologics").  Earlier this year, the President unveiled his 2012 budget proposal, which if approved unchanged, would trim the data exclusivity provision of the PPACA from 12 years to 7 years (see "President's Budget Proposal Increases Funding for Basic Research But Seeks to "Trim" Data Exclusivity Period and Pay-for-Delay Agreements").

    TTP Members
    TTP members – original four members in orange and additional five members in green.

  • By Kevin E. Noonan —

    Lilly On the same day the Federal Circuit rendered its opinion in AMP v. USPTO (the Myriad case), the Court also handed down its opinion in Eli Lilly & Co. v. Activis Elizabeth LLC.  In an opinion by Judge Newman, joined by Judge Lourie (Judge Friedman was on the panel, but died before the court reached its decision), the Federal Circuit reversed a District Court finding of invalidity based on the patentee's failure to satisfy the utility aspect of the enablement requirement of § 112, 1st paragraph ("The specification shall contain a written description of the invention . . . in such full, clear, concise, and exact terms as to enable any person skilled in the art to which it pertains, or with which it is most nearly connected, to make and use the same. . ."), reversed the District Court's holding that, if held valid, defendants would be liable for inducing infringement but not contributory infringement, and affirmed in all other respects.

    The defendants (Actavis Elizabeth LLC, Sun Pharmaceuticals Ltd., Sandoz Inc., Mylan Pharmaceuticals Inc., Apotex, Inc., Aurobindo Pharma, Ltd., and  Teva Pharmaceuticals USA Inc.) each filed an Abbreviated New Drug Application (ANDA) on atomoxetine ((R)-(–)-N-methyl-3- (2-methylphenoxy)-3-phenylpropylamine), sold by Eli Lilly as Strattera® to treat attention deficit/hyperactivity disorder (ADHD).  The patent-in-suit was U.S. Patent No. 5,658,590; specifically at issue was claim 1:

    A method of treating attention- deficit/hyperactivity disorder comprising administering to a patient in need of such treatment an effective amount of tomoxetine.
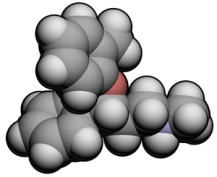
    Atomoxetine Atomoxetine itself was known and patented by Eli Lilly & Co. in U.S. Patent No. 4,314,081, which expired in 1999.  That patent disclosed urinary incontinence and depression as two uses for the drug; it was only in 1993 that Eli Lilly scientists suggested that atomoxetine could be used to treat ADHD.  In this regard, the District Court found that "occurrence of ADHD is wide, the cause is unknown, and the mechanism of drug treatment unclear," all of which is exacerbated by the lack of an animal model.  In addition, the '590 patent specification taught that treatments current at the time the invention was made were inadequate and "exhibited deficiencies" (including sudden death in children).  Despite these deficiencies, testimony established that the Lilly scientists' suggestion that the drug be used to treat ADHD was "met with skepticism" and FDA approval took approximately 7 years.

    The ANDAs filed by the defendants contained Paragraph IV certifications of invalidity (for obviousness, and lack of enablement based on scope and utility), unenforceability, and non-infringement (under both inducement to infringe and contributory infringement).  The District Court held that none of the defendants had established invalidity for obviousness or lack of enablement based on scope, but that claim 1 of the '590 patent was invalid for failure to provide evidence that the claim was operable, i.e., the specification failed to satisfy the utility aspect of the enablement requirement.  The basis for the District Court's decision was that the "utility [of the claimed method] was not established because experimental data showing the results of treatment of ADHD were not included in the specification."  The question addressed by the Federal Circuit was the extent of disclosure required to satisfy this requirement of the statute.  Lilly contended that disclosing the use of atomoxetine to treat ADHD was enough, while defendants contended (and the District Court agreed) that the claimed utility must be established by experimental evidence in the application as filed.
    Federal Circuit Seal The Federal Circuit reversed, based on its determination that the District Court's finding that experimental data was required to be in the application as filed was error.  Provided priority between inventors is not involved, the Court's opinion said that applicants are able to submit data "either before or after the patent application was filed," where after-filed evidence "can be used to substantiate any doubts as to the asserted utility since this pertains to the accuracy of a statement already in the specification."  The Court's opinion reversing the District Court's finding of invalidity was based on the requirement that an assertion of utility must be accepted as sufficient to satisfy the utility requirement unless there is "some reason" for the skilled worker to question the "objective truth" of the asserted utility, citing In re Langer, 503 F.2d 1380, 1391 (CCPA 1974), In re Brana, 51 F.3d 1560 (Fed. Cir. 1995), and In re Marzocchi, 439 F.2d 220, 223 (CCPA 1971).  From Brana, the Court quoted the requirement that "[o]nly after the PTO provides evidence showing that one of ordinary skill in the art would reasonably doubt the asserted utility does the burden shift to the applicant to provide rebuttal evidence sufficient to convince such a person of the invention's asserted utility," citing In re Bundy, 642 F.2d 430, 433 (CCPA 1981).

    The Federal Circuit noted that the accuracy of the disclosed utility had not been questioned by the defendants, and that it was also undisputed that Lilly performed successful clinical trials after the patent application that resulted from the '590 patent was filed.  And during prosecution, the examiner "did not require the submission of data" that ADHD could be treated with atomoxetine.

    In assessing the adequacy of the disclosure of in the specification, the Federal Circuit opinion also noted that "[t]he utility of this product to treat ADHD is not so incredible as to warrant the special procedures that are authorized for use when the examiner doubts the described utility" or "for subject matter in once notoriously intractable areas such as cures for baldness or cancer."  Here, "deference is owed to the 'qualified agency presumed to have done its job,'" citing Am. Hoist & Derrick Co. v. Sowa & Sons, Inc., 725 F.2d 1350, 1359 (Fed. Cir. 1984).

    The opinion appears to restore to its traditional moorings the requirements for enablement that were called into question by the Court's decision in Janssen Pharmaceutica N.V. v. Teva Pharmaceuticals USA, Inc., 583 F.3d 1317 (Fed. Cir. 2009) (aka In re '318 Patent Litigation).  In that case, the teachings that a specific compound, galantamine, could be used to treat Alzheimer's disease were found insufficient because "animal tests were 'not finished . . . by the time the '318 patent was allowed.'"  The panel here distinguished the purported lack of an enabling disclosure in Janssen with the instant case, where the specification "contained a full description of the utility, experimental verification had been obtained before the patent was granted, and the examiner had not requested additional information."  Here, the Court's opinion seems to be based on the existence of after-filing evidence sufficient for Lilly to rebut an assertion that the "objective truth of the asserted utility" would be doubted by the skilled worker.

    Regarding the other asserted grounds of invalidity, the Federal Circuit affirmed the District Court finding that defendants had not established that the claimed invention was obvious.  Defendants' obviousness argument was based on prior art that another compound — desipramine, a norepinephrine inhibitor — was used to treat ADHD and that atomoxetine was also to be a norepinephrine inhibitor.  Thus, defendants argued that it would have been obvious to test atomoxetine for the treatment of ADHD.  The District Court found this argument to be insufficient, because it is not permissible to "pick and choose" from references to find evidence supporting obviousness, while excluding other evidence (including serious side-effects from despiramine use) that would be necessary to appreciate what a reference "fairly suggests," citing In re Wesslau, 353 F.2d 238, 241 (CCPA 1965).  The District Court also rejected defendants' argument that atomaxetine would have been "obvious to try" for treating ADHD; this determination was based on the Supreme Court's decision in KSR Int'l Co. v. Teleflex Inc. to the effect that "there was no evidence . . . the exercise of common sense would have led a person of ordinary skill to test atomoxetine for treatment of ADHD" (a conclusion supported by testimony from experts on both sides).

    Defendants also argued that the specification did not enable the claimed method throughout its full scope, relating to the claim limitation directed to "administering to the patient an effective amount" of the drug for treating ADHD, because this limitation was not directed to the expressly disclosed formulations.  Defendants presented expert testimony that undue experimentation would be required to determine particular formulations and effective amounts to be administered.  The specification disclosed that oral formulations were preferred and contained generic disclosure of oral formulations, as well as ranges of preferred doses.  Applying the rubrics of In re Wands, the District Court found that a person skilled in the art would be able to make formulations throughout the full scope of the claimed limitation through the exercise of no more than routine experimentation.  In affirming this result, the Federal Circuit rejected defendants' argument based on ALZA Corp. v. Andrx Pharmaceuticals, LLC, 603 F.3d 935 (Fed. Cir. 2010) (where the claimed formulations were "a breakaway from the prior art"), and asserted the principle from Spectra-Physics, Inc. v. Coherent, Inc., 827 F.2d 1524, 1534 (Fed. Cir. 1987) that "a patent need not teach, and preferably omits, what is well known in the art."

    Finally, the Federal Circuit reversed the District Court's finding that if claim 1 of the '590 patent was valid, the defendants were liable for inducing infringement under 35 U.S.C. § 271(b), but not contributory infringement under § 271(c).  According to the Court, the possibility of "off-label" uses were irrelevant to the issue, since each defendant would be selling the generic product with an FDA-approved label directed to Lilly's claimed method.  Moreover, since the only use for which the ANDA filers could sell the drug was the FDA-approved use, atomoxetine had no "substantial non-infringing use" and thus its sale also constituted contributory infringement under § 271(c).

    Eli Lilly & Co. v. Actavis Elizabeth LLC (Fed. Cir. 2011)
    Nonprecedential disposition
    Panel:  Circuit Judges Newman, Friedman, and Lourie
    Opinion by Circuit Judge Newman
    (The opinion notes that Circuit Judge Friedman heard oral argument in the appeal, but died on July 6, 2011, and did not participate in the final decision.  The case was decided by the remaining judges on the panel, in accordance with Fed. Cir. Rule 47.11.)

  • By James DeGiulio —

    New England Journal of Medicine In an article published in the New England Journal of Medicine on August 4, U.S. Food and Drug Administration (FDA) executives presented a sneak preview of what to expect when the FDA issues its biosimilar standards under the Biologics Price Competition and Innovation Act (BPCIA) later this year.  The article, entitled "Developing the Nation's Biosimilars Program," was authored by Dr. Steven Kozlowski (Director, Office of Biotechnology Products), Dr. Janet Woodcock (Director, Center for Drug Evaluation and Research), Dr. Karen Midthun (Director, Center for Biologics Evaluation and Research), and Dr. Rachel Behrman Sherman (Associate Director for Medical Policy).

    FDA If the article was intended to indicate how far the FDA has progressed in formulating biosimilar standards, it appears we may be waiting a while longer, as the article provides minimal substantive guidance.  However, if a preview of substantive biosimilarity standards is desired, the article does indicate that the FDA is evaluating the adoption of some of the guidelines previously developed by the European Medicines Agency (EMA).  The article makes several references to the EMA's Guideline on Similar Biological Medicinal Products, published in 2005, as well as the EMA's 2010 Guideline on Similar Biological Medicinal Products containing Monoclonal Antibodies (see "EMA Publishes Guidelines for Biosimilar Antibodies – Part I" and "Part II"), answering the question of how much the FDA would seek to harmonize its biosimilar guidelines with those of the EMA, and suggesting that the EMA Guideline documents could provide a substantive sneak preview of the FDA's biosimilar standards.

    Rather than provide any details on the actual scientific standards, the authors instead categorize the FDA's tentative approach to formulating these standards using a series of buzzwords:  (1) "totality of the evidence," including sponsor input on whether further clinical studies are needed; (2) no chance of a "one size fits all" standard akin to bioequivalence/bioavailability under the Hatch-Waxman ANDA scheme; and (3) depending on how well the mechanism of the biologics are characterized, "fingerprint identification" studies, rather than full scale clinical trials, may be permitted to establish biosimilarity with a reference product.

    Biologics vary in complexity from cellular therapies to small, highly purified proteins, although even the simplest biologics typically are more structurally complex than the small-molecule drugs in today's market.  Partly due to this complexity, new biologics require a substantially greater investment from BLA filers (or "sponsors") for their development and regulatory approval.  As a result, biologics tend to be more expensive than small-molecule drugs for patients, and the article notes that these cost obstacles currently limit patient access to these biological products.  The Hatch-Waxman framework has proved very successful in overcoming cost barriers and improving patient access to small-molecule drugs.  In 2009, almost 75% of small-molecule prescriptions dispensed in the United States were for generics, and the approval of a generic drug resulted in average savings of 77% of the product's cost within 1 year.  The article acknowledges that cost reductions for biosimilars probably won't be as large, at least in the early going, but points to the Federal Trade Commission report ("Emerging Health Care Issues: Follow-on Biologic Drug Competition"; see "No One Seems Happy with Follow-on Biologics According to the FTC"), which predicts that the availability of biosimilar products will significantly reduce the cost of biologics and increase their accessibility.

    The article outlines the familiar challenges facing the FDA in formulating the standards that an applicant must meet to secure approval for a follow-on biologic product.  The agency must establish scientific criteria that address the key question:  how similar is similar enough when it comes to the substitution of complex biologic drug products in clinical practice?  The complex structures of biologic products are not as easily characterized as small-molecule drugs, with biologics generally requiring more advanced and expensive technology.  Further, therapeutic proteins must have a specific set of structural features, such as glycosylation and protein folding, essential to their intended effect, and slight modifications in these features can affect the performance of these drugs in humans.  This additional secondary structural layer is required to fully characterize biologics, and this layer is more difficult to detect using conventional technological means.  This secondary structural layer, often represented by chemical modifications of the amino acid sequence of the protein, has proven to be just as important as the primary amino acid sequence and structure, for changes in the chemical modifications of biologics affect their immunogenicity and safety profiles.

    The authors quickly turn to reassuring the reader that the FDA is fully equipped to deal with these challenges, discussing the agency's expertise in this area.  The article notes that the FDA has worked with biologics since the mid-1990s, evaluating the physiochemical and functional assays used to characterize biologics and test changes in manufacturing processes.  The FDA has made decisions on when animal or clinical studies are required to resolve any remaining uncertainties about the comparability of the products created before and after such changes, these decisions being extremely relevant in setting the biosimilar standards.  The FDA must consider the following balance:  the biosimilar applicant must be required to establish with sufficient confidence that safety and efficacy are not diminished with their product.  However, requiring the biosimilar applicant to repeat all of the clinical trials conducted by the BLA holder would defeat the purpose of the regulatory framework.  The FDA acknowledges that simply repeating all of the sponsor's trials is not feasible and is too costly.  Utilizing knowledge from the reference products, the abbreviated pathway must eliminate unnecessary (and therefore unethical) testing of biosimilars in animals and humans.

    EMA Despite the FDA's experience in deciding when additional clinical trials are required to establish similarity between two products, the authors prudently acknowledge that the EMA has even more experience in this area.  The EMA biosimilar guidelines have been in place since 2005, and the EMA first approved a biosimilar in 2006.  The FDA seems to be fashioning its approach after the EMA, relying on the EMA's more than five years of experience in implementing a biosimilar framework, and latching onto the EMA's standards and ideas for determining exactly how many studies will be required.

    Continuing with the theme of acknowledging others with more experience than the FDA, the article suggests that BLA holders should expect to be heavily involved with setting biosimilar standards and approving biosimilar products than under the Hatch-Waxman framework.  Sponsor input will be more extensive and will carry more weight, and extensive product review will be required early on.  The new pathway will require a new paradigm for sponsor–FDA interactions.  Although the agency frequently meets with sponsors before they submit investigational new drug (IND) applications, a more extensive product review will be required to determine how much additional data and studies are needed for a biosimilar.  The first step will be taken by the FDA, which will conduct an in-depth review of the biosimilar applicant's comparative analytic characterization and in vitro data.  Once the review of this data is completed, the FDA will work with the BLA holder/sponsor to determine the scope of any required animal and human studies.  Since the BLA holder/sponsor has an interest in keeping the biosimilar product off of the market, the agency is currently considering how to structure this interaction so that neutral and more reliable input can be acquired.

    The FDA has traditionally relied on integrating various kinds of evidence in making regulatory decisions, and drafting the biosimilar guidelines will be no exception, as seen by importing the EMA and BLA holder standards.  The FDA authors prefer a flexible "totality of the evidence" approach for assessing biosimilars, since multiple assay technologies are currently used to characterize the structural attributes of biosimilars, and more likely to be developed going forward.  For small molecules, a generic's structure is readily determined by nuclear magnetic resonance, mass spectrometry, infrared spectrometry, X-ray crystallography, or other well-known physical methods.  However, accepted methods such as these do not yet exist for biosimilar products.  Developers of biosimilars who intend to match reference products more closely will need to select appropriate assays and source materials, and tune their processes carefully.  Current approaches to manufacturing process design and improvements in process analytics will become more useful as the technology continues to progress.

    Even if certain technological assays become recognized as the gold-standard for determining a biological structure, given the complex nature of biologics, it's unlikely that a "one size fits all" systematic assessment of biosimilarity will ever be developed.  Indeed, the EMA biosimilar guidelines are highly product-specific, with the EMA guidance providing structural, animal, and clinical study requirements for each biosimilar product.  This is in sharp contrast to the Hatch-Waxman small molecule scheme, where often an ANDA filer must show only bioavailability and bioequivalence for approval.  In the biosimilar setting, FDA scientists will need to integrate multiple types of information from multiple assays to provide an overall assessment that a biologic is biosimilar to an approved reference product.  The authors also suggest that implementing a more rigorous pharmacovigilance program will also be significant information source, with an emphasis on tracking adverse events and even the smallest change in manufacturing.  This will inform future decisions not only on that particular product, but also on products that can be grouped in the same class, such as monoclonal antibodies.

    Importantly, the development of the mechanism of the biologic's activity will also factor in determining the product-specific standards for the particular biosimilar.  If the mechanism is well known, there might be more flexibility to deviate from the structure of the approved reference product.  For example, if the literature shows that all known glycosylation patterns of a biologic molecule show comparable activity and immunogenicity, a glycosylation pattern identical to the reference product may not be required for a finding of biosimilarity.  Understanding the mechanism of biologic activity, including any clinical information available, will help evaluate the risks of changes in manufacturing by the applicant and the resulting potential to introduce harmful impurities.  This factor is significant to the basic science research community, including university research labs, which could be in a position to secure funding from biosimilar applicant companies that need to establish the safety profile of new manufacturing protocols.

    If the mechanism of a biosimilar product is well-characterized, the FDA may accept a "fingerprint"-like identification of similar patterns in two different products.  The EMA monoclonal antibody guidelines introduce an outline to consider when designing biosimilarity studies, including the use of populations, pharmacodynamic markers, and end points that are sensitive to the potential differences between products.  The EMA guidelines allow a biosimilar monoclonal antibody to differ structurally from the reference product, providing a specific set of "fingerprint" metrics are satisfied.  These "fingerprint" strategies were used to secure the approval of the generic low-molecular-weight heparin product enoxaparin, which is structurally different from the reference product.  The authors admit that although animal and clinical studies will generally be needed for protein biosimilars for the foreseeable future, the scope and extent of such studies may be reduced further if more extensive "fingerprint"-like characterization is used.  Therefore, it is conceivable that at some future date, there may be a set of minimum "fingerprint" biosimilar characteristics, akin to the familiar bioequivalence and bioavailability minimum requirements seen in Hatch-Waxman ANDA filings.  Of course, that future date is likely decades away.

    Finally, the authors seemingly feel obligated to address the "interchangeability" standard of the BPCIA, which provides for when a biosimilar may be substituted for the reference product without requiring the prescriber's intervention (as in the classic generic drug scenario).  The language in the BPCIA focuses on alternating or switching between the two products, requiring the biosimilar to provide the same risk and clinical result as using the reference product throughout.  The article presents little reason for optimism in meeting such a heightened standard, instead focusing on the FDA's dedication to policing the marketplace to ensure that merely biosimilar products are not substituted for a reference product without the prescriber's consent.  Even equipped with the EMA guidelines as a template, the FDA has its hands full in determining what should be the standards for biosimilarity.  Until the FDA's guidelines are established, the standard for interchangeability is likely to be left to statutory interpretation according to the language in the BPCIA.  This (perhaps purposefully) vague standard is extremely difficult to meet as it reads textually, and it would be surprising if the FDA approves any biosimilar as interchangeable before the year 2020.

  • By Donald Zuhn —

    USPTO Seal The U.S. Patent and Trademark Office recently established a website to provide information and seek input on the implementation of the Leahy-Smith American Invents Act, which the House of Representatives passed on June 23.  Although the Office notes that the legislation is still pending before the Senate and "neither its passage nor enactment can be presumed," the Office has decided "to proactively engage with the public to best prepare for the timely and effective implementation of the legislation should it be enacted."

    Noting that implementation of the legislation will require "a series of . . . expeditious rulemaking proceedings," the new website indicates that the process of implementing the legislation might be facilitated by collecting input and comments from stakeholders and the public prior to enactment of the legislation.  The Office points out, however, that any input received will not be treated as a formal request for comments or a rulemaking, and that if the legislation is enacted, the Office will issue formal notices of proposed rulemaking.

    The website provides five categories of "implementation documents":

    • Patents
    • Board of Patent Appeals and Interferences (BPAI)
    • Fees and Budgetary Issues
    • Congressionally-Directed Studies and Reports
    • Miscellaneous

    For the "Patents" category, the Office provides a 6-page list of potential items to aid the public in focusing its comments with respect to that category, and a single page list of changes having effective dates of 60 days or less.  Under the "Board of Patent Appeals and Interferences (BPAI)" category, the Office provides a 3-page list of potential items to aid the public in focusing its comments with respect to that category.  The "Fees and Budgetary Issues" category includes a fee setting flow chart and a timeline for setting fees under § 10 of the legislation.  Under the "Congressionally-Directed Studies and Reports" category, the Office provides a table listing the seven reports, two studies, and single program that are mandated by the legislation.  The "Miscellaneous" category currently lists no implementation documents.

    The website also provides a list of resources, including a useful 6-page section-by-section summary of the changes to be implemented by the Leahy-Smith American Invents Act.

    Comments should be submitted to the attention Hiram Bernstein, Senior Legal Advisor, Office of Patent Legal Administration (Patents and Miscellaneous); Michael P. Tierney, Administrative Patent Judge, Board of Patent Appeals and Interferences (Board of Patent Appeals and Interferences); Michelle Picard, Senior Advisor for Financial Management, Office of the Chief Financial Officer (Fees and Budgetary Issues); or Janet Gongola, Associate Solicitor, Office of the Solicitor (Congressionally-Directed Studies and Reports), and should also reference one of the five categories and the specific issue being addressed in the subject line.  Comments can be submitted by e-mail to:  aia_implementation@uspto.gov, or by regular mail to:  Mail Stop Comments-Patents, Commissioner for Patents, P.O. Box 1450, Alexandria, VA 22313–1450, marked to the attention of Hiram Bernstein.

  • By Sherri Oslick —

    Gavel About Court Report:  Each week we will report briefly on recently filed biotech and pharma cases.

    Astrazeneca AB et al. v. Hetero Drugs, Ltd., Unit III et al.
    3:11-cv-04468; filed August 2, 2011 in the District Court of New Jersey

    • Plaintiffs:  Astrazeneca AB; Aktiebolaget Hassle; Astrazeneca LP; KBI Inc.; KBI-E Inc.
    • Defendants:  Hetero Drugs, Ltd., Unit III; Hetero USA Inc.

    Infringement of U.S. Patent Nos. 5,714,504 ("Compositions," issued February 3, 1988), 5,877,192 ("Method for the Treatment of Gastric Acid-Related Diseases and Production of Medication Using (-)Enantiomer of Omeprazole," issued March 2, 1999), and 6,875,872 ("Compounds," issued April 5, 2005), 6,369,085 ("Form of S-omeprazole," issued April 9, 2002) following a Paragraph IV certification as part of Hetero's filing of an ANDA to manufacture a generic version of AstraZeneca's Nexium® (esomeprazole magnesium, used for the treatment of gastroesophageal reflux disease).  View the complaint here.


    Tibotec Inc. et al. v. Lupin Ltd. et al.

    2:11-cv-04437; filed August 1, 2011 in the District Court of New Jersey

    • Plaintiffs:  Tibotec Inc.; Tibotec Pharmaceuticals; G.D. Searle, LLC
    • Defendants:  Lupin Ltd.; Lupin Pharmaceuticals Inc.; Mylan Pharmaceuticals Inc.; Mylan Inc.

    Infringement of U.S. Patent Nos. 5,843,946 ("α- and β-Amino Acid Hydroxyethylamino Sulfonamides Useful as Retroviral Protease Inhibitors," issued December 1, 1998) and 6,248,775 (same title, issued June 19, 2001) following a Paragraph IV certification as part of Lupin's filing of an ANDA to manufacture a generic version of Tibotec's Prezista® (darunavir, used to treat human immunodeficiency virus (HIV-1) infection).  View the complaint here.


    Pfizer Inc. et al. v. Hetero Drugs Ltd. et al.

    2:11-cv-04372; filed July 28, 2011 in the District Court of New Jersey

    • Plaintiffs:  Pfizer Inc.; Pfizer Ltd.; Pfizer Ireland Pharmaceuticals
    • Defendants:  Hetero Drugs Ltd.; Hetero USA Inc.; Invagen Pharmaceuticals Inc.; Hetero Labs Ltd., Unit V.; Camber Pharmaceuticals Inc.

    Infringement of U.S. Patent No. 6,469,012 ("Pyrazolopyrimidinones for the Treatment of Impotence," issued October 22, 2002) following a Paragraph IV certification as part of Hetero's filing of an ANDA to manufacture a generic version of Pfizer's Viagra® (sildenafil citrate, used to treat erectile dysfunction).  View the complaint here.

  • Calendar

    August 15-19, 2011 – Intellectual Property Law Summer School 2011 (IBC Legal) – Cambridge, UK

    August 16-18, 2011 – The Bilski Impact: Procuring & Enforcing Software, Business Methods & Bioinformatics Patents (Patent Resources Group) – Alexandria, VA and Southfield, MI

    September 6-7, 2011 – Pharmaceutical Law Academy (IBC Legal Conferences) – London, UK

    September 8, 2011 – Patents and the Written Description Requirement: Lessons Learned Since Ariad v. Lilly (Strafford) – 1:00 – 2:30 PM (EDT)

    September 11-13, 2011 – 2011 Annual Meeting (Intellectual Property Owners Association) – Los Angeles, CA

    September 14-16, 2011 – 85th Annual Meeting (Intellectual Property Institute of Canada) – Chicago, IL

    September 18-20, 2011 – Accelerating Intellectual Property and Innovation in South Africa*** (South African Department of Science and Technology) – Cape Town, South Africa

    September 19-21, 2011 – Business of Biosimilars & Biobetters*** (Institute for International Research) – Boston, MA

    September 22-23, 2011 – FDA Boot Camp*** (American Conference Institute) – Boston, MA

    September 26-27, 2011 – Biosimilars and Biobetters*** (SMi Group) – London, UK

    September 26-27, 2011 – Life Sciences Business Development & Acquisitions in Emerging Markets (American Conference Institute) – New York, NYNew York, NY

    October 4-5, 2011 – Maximizing Pharmaceutical Patent Lifecycles*** (ACI) –

    October 5-6, 2011 – Biotech Patenting*** (C5) – London, UK

    ***Patent Docs is a media partner of this conference or CLE

  • AM_Banner
    The Intellectual Property Owners Association (IPO) will be holding its 2011 Annual Meeting on September 11-13, 2011 in Los Angeles, CA.  Among the presentations being offered at the annual meeting are:

    • Recent case law developments;
    • A look at patent landscaping and analysis;
    • Legal issues surrounding genetic information research and development — Patent Docs author Dr. Kevin Noonan of McDonnell Boehnen Hulbert & Berghoff LLP will be a panelist on the first of this session's two panels;
    • Damages;
    • Current state of the ITC;
    • Patentability issues;
    • Recent developments in patent offices around the world; and
    • Ethical questions arising from en banc review of Therasense v. Becton Dickinson — Paul Berghoff of McDonnell Boehnen Hulbert & Berghoff LLP will moderate this panel.

    In addition, James Pooley, Deputy Director General for Patents for the World Intellectual Property Organization, will present the luncheon keynote on September 13.

    A program for the meeting, including an agenda, descriptions of the scheduled sessions, and list of speakers, can be obtained here, and a brochure for the meeting can be obtained here.

    The registration fee for the meeting is $950 for IPO members or $1,450 for attendees who are not IPO members.  Those interested in registering for the meeting can do so here.

  • Strafford #1 Strafford will be offering a webinar entitled "Patents and the Written Description Requirement: Lessons Learned Since Ariad v. Lilly" on September 8, 2011 from 1:00 – 2:30 PM (EDT).  Michael Dzwonczyk of Sughrue Mion, PLLC will provide guidance for dealing with the written description requirement of 35 U.S.C, § 112, and discuss the lessons learned since the Federal Circuit's decision in Ariad v. Eli Lilly for compliance and patent challenges.  The webinar will review the following questions:

    • How is the Ariad decision being applied by the courts — and what has its impact been on patent prosecutions?
    • What are the implications of Ariad for the predictable and non-predictable arts?
    • What steps can patentees and counsel take to meet the written description requirement and withstand invalidity challenges based on the written description?

    An interactive Q&A session will follow the presentation.

    The registration fee for the webinar is $297 ($362 for registration and CLE processing).  Those registering by August 12, 2011 will receive a $50 discount.  Those interested in registering for the webinar, can do so here.