• Will I lose my dignity?
    Will someone care?
    Will I wake tomorrow
    From this nightmare?

    "Will I," Rent

    By Kevin E. Noonan —
    MyriadThe Supreme Court's grant of certiorari over the question "Are human genes patentable"
    had raised for many the specter of an uninformed generalist court rendering a
    decision containing dicta that would negatively affect biotechnology.  This possibility is real, in view of the
    voices raised against patenting human genes based on moral, policy, or
    ideological grounds.  While strongly
    felt, these sentiments are based, in large part, on a misunderstanding or
    mischaracterization of either the facts or the law, specifically patent
    law.  These misstatements include
    arguments based on patent claims somehow exerting an ownership interest in
    individual's DNA, or that isolated human nucleic acid claims preempt future
    research (in the face of thousands of published scientific journal articles
    published since the BRCA gene patents were granted), or that isolated DNA
    patents inhibit future technologies like personalized medicine.

    This is an argument that might have been relevant
    30 years ago, but at least two facts mitigate against resolving this issue
    now.  The first is that the day of the
    DNA patent claim is rapidly coming to a close.  These types of patents were first filed directed to specific genes
    encoding biologically relevant proteins having therapeutic benefits.  These include erythropoietin, tissue plasminogen
    activator, interferon, blood clotting Factors VIII and IX, and others.  (Indeed, one of the most compelling types of
    evidence not considered by any court in the Myriad
    case is the testimony of the tens or hundreds of thousands of patients whose lives
    have been saved or bettered by biotechnology drugs protected, in part, by
    so-called "gene patents.")  Myriad's BRCA gene patent claims are in this form, one of the reasons
    why these claims are not infringed by genetic diagnostic methods used
    today.

    The vast majority of gene patent applications,
    however, were filled as the result of the Human Genome Project.  These patents are limited in at least three
    ways.  First, the U.S. Patent and Trademark Office has established a
    policy that patents will be granted only if an applicant provided on the filing date specific, substantial, and credible utility for the protein encoded by the claimed DNA.  This provision prevents an applicant from
    obtaining patent exclusivity unless the applicant shows that she establishes utility,
    a requirement of the patent statute, of the encoded protein.  (This requirement is related to the policy
    enunciated in Brenner v. Manson,
    where claims to methods for making steroid molecules were invalidated based on
    failure to disclose a utility for the steroid molecules made using the
    methods.)  Second, certain researchers
    (notably, Sir John Shulston in England) made certain that the sequence results
    of the HGP were published in public databases as soon as possible; doing so
    placed the sequences in the prior art and precluded patentability for any
    application filed after the sequences were made public.  Finally, because almost all of these patents
    were filed at the turn of the century, and the term of these patents will expire
    no later than 20 years after application filing, the "problem" of
    gene patents will disappear no later than 2020 (and in many instances much
    earlier even if a patent can be obtained under the enhanced utility
    requirements).

    Another reason why the ACLU's challenge is
    anachronistic (except for those meaning to make an ideological point) is that present
    and future technology, such as genetic diagnostic testing or personalized
    medicine, is not prevented or inhibited by claims to isolated DNA molecules
    encoding a specific human protein.  This
    is a consequence of the history of these type of claims for providing means for
    producing biologically relevant proteins like EPO and TPA.  The claims require that the DNA is isolated and that it encodes the full-length protein,
    as defined by specific amino acid sequence.  (For these reasons these claims are
    exceedingly narrow, and do not broadly preempt:  for example, such claims are
    not literally infringed if the encoded protein differs by a single amino acid,
    meaning that a difference of a single methylene group (the difference between a
    valine and an isoleucine residue) is enough to take a species outside the scope
    of the claim.)  Modern gene sequencing methods used for
    diagnostics neither isolate DNA nor produce a full-length DNA molecule
    encoding the protein.  Rather, sequencing
    is performed in such a way that fragments of the DNA are produced in a reaction
    mixture and sequenced without isolation; what is produced is the sequence, a
    characteristic of the DNA molecule that is not
    protected by the patent.  (This is why
    infringement of isolated DNA claims does not lie by using the sequence
    information to, inter alia,
    interrogate a DNA database.)  And the Myriad
    case has established that method claims directed to merely comparing an
    individual's gene sequence with a reference sequence and disclosing mutations
    identified as indicating a risk of disease, without more, are not patent
    eligible (under either Bilski v Kappos
    or Mayo v. Prometheus).

    So there appears to be no sound policy reason for
    having the Supreme Court render a decision on the patent eligibility of human
    genes.  Sadly, there is seemingly no
    sound business reason for Myriad Genetics to have the Court decide this question
    either.  Myriad's counsel, Greg Castanias,
    argued in the first Federal Circuit oral argument that the plaintiffs did not have
    standing because there was no redressability to their claim.  Specifically, Mr. Castanias argued that even
    if the Court ruled in plaintiffs' favor, Myriad had other patent claims that
    would prevent Dr. Harry Ostrer and other plaintiffs from providing BRCA gene
    diagnostic tests.  In addition, while
    these patents were undoubtedly important when Myriad was establishing its
    diagnostics business they are less important now.  Fifteen years and thousands of genetic tests
    (apparently without any failures) later, the company's reputation, experience and
    extensive database is far more valuable than the claims at issue.

    Under these circumstances, the question must be
    asked whether the prudent thing for Myriad to do is grant Dr. Ostrer a covenant
    not to sue on all the patents and claims involved in this litigation.  The result would be to render the issues
    before the Court moot; regardless of the perceived importance of the Question
    Presented, even the Supreme Court is limited by Article III of the Constitution
    and cannot render an advisory opinion on this issue.  Such an action by Myriad would not only
    prevent this case from being decided, it would preclude the Court from
    including in its opinion dicta on the patent eligibility of isolated DNA from
    other species (which may have importance in fields like agriculture or
    biofuels) or other "products of nature" that may provide the basis
    for biologic and other types of drugs.  In view of the seemingly complete lack of any practical reason for
    having the Supreme Court decide this issue, and the risks to the progress
    provided for a generation by the biotechnology industry, nothing other than a
    desire to be vindicated remains to support seeing this case through to its
    conclusion.  Frankly, in view of what is at stake, such vindication is simply not enough.
  • By Donald Zuhn —

    FDA ReportLast month, the U.S. Food
    and Drug Administration released its annual report on innovative drug
    approvals, noting that the agency approved 35 novel medicines in FY 2012.  The 35 approvals equaled the number of approvals for FY 2011, and marked the highest number of
    approvals since the FDA approved 36 new drugs in FY 2004.  According to the report on FY 2012 Innovative
    Drug Approvals
    ,
    ten of the 35 new drugs were approved for the treatment of cancer, two represent
    advances in personalized medicine, and nine new drugs were approved for the
    treatment of orphan diseases.  The report
    notes that the term "drugs" is intended to include products approved
    by both the FDA's Center for Drug Evaluation and Research (CDER), which reviews New
    Drug Applications (NDAs) for pharmaceuticals, and the agency's Center for
    Biologics Evaluation and Research (CBER), which reviews biological therapeutic
    agents.

    The report indicated that most
    of the new drugs were approved for U.S. patients before the drugs were
    available in other countries.  In
    particular, of the 32 new drugs for which comparisons could made with approvals
    in other countries, 75% (24) were first approved in the U.S.  The FDA was able to achieve such results by
    approving 77% of the 35 new drugs on the first cycle of review, without
    requiring additional information that would have lengthened the approval time.

    The 35 new drugs that were
    approved in FY 2012 include the following:

    •
    AMYVID (florbetapir
    F18) – Avid Radio-pharmaceuticals, Inc.
    • AUBAGIO (teriflunomide) – Sanofi-Aventis,
    U.S., LLC
    • BElVIQ (lorcaserin hydrochloride) –
    Arena Pharmaceuticals, Inc.
    • BOSULIF (bosutinib) – Pfizer Inc.
    •
    CHOLINE C 11 –
    Mayo Clinic PET Radiochemistry Facility
    • ELELYSO (taliglucerase alfa) –
    Protalix Biotherapeutics Inc.
    •
    ERIVEDGE (vismodegib)
    – Genentech, Inc.
    •
    ERWINAZE (asparaginase
    erwinia chrysanthemi) – EUSA Pharma (USA), Inc.
    •
    EYLEA (aflibercept)
    – Regeneron Pharmaceuticals, Inc.
    • FERRIPROX (deferiprone) – Apopharma,
    Inc.
    • GINTUIT (allogeneic cultured
    keratinocytes and fibroblasts in bovine collagen) – Organogenesis, Inc.
    • HEMACORD (hemapoietic progenitor
    cells, cord (HPC-C) – New York Blood Center, Inc.
    • HPC, Cord Blood (hemapoietic progenitor
    cells, cord (HPC-C) – Clinimmune Labs
    •
    INLYTA (axitinib)
    – Pfizer, Inc.
    •
    JAKAFI (ruxolitinib)
    – Incyte Corp.
    •
    KALYDECO (ivacaftor)
    – Vertex Pharmaceuticals, Inc.
    •
    KYPROLIS (carfilzomib)
    – Onyx Pharmaceuticals, Inc.
    •
    LINZESS (linaclotide)
    – Forest Laboratories, Inc.
    •
    MENHIBRIX (Meningococcal
    Groups C and Y and Haemophilus b Tetanus Toxoid Conjugate Vaccine) – GlaxoSmithKline
    Biologicals
    •
    MYRBERTIQ (mirabegron)
    – Astellas Pharma Global Development, Inc.
    •
    NEUTROVAL (tbo-filgrastim)
    – Sicor Biotech UAB
    •
    OMONTYS (peginesatide)
    – Affymax, Inc.
    •
    ONFI (clobazam)
    – Lundbeck, Inc.
    •
    PERJETA (pertuzumab)
    – Genentech, Inc.
    •
    PICATO (ingenol
    mebutate) – Leo Pharma AS
    •
    PREPOPIK (sodium
    picosulfate, magnesium oxide, citric acid) – Ferring Pharmaceuticals
    •
    STIVARGA (regorafenib)
    – Bayer Healthcare Pharmaceuticals, Inc.
    •
    STENDRA (avanafil)
    – Vivus Inc.
    •
    STRIBILD (elvitegravir,
    cobicistat, emtricitabine, tenofovir disoproxil fumarate) – Gilead Sciences, Inc.
    •
    SURFAXIN (lucinactant) – Discovery Laboratories, Inc.
    •
    TUDORZA PRESSAIR
    (aclidinium bromide) – Forest Laboratories, Inc.
    • VORAXAZE (glucarpidase) – BTG
    international, Inc.
    • XTANDI (enzalutamide) –
    Medivation, Inc.
    • ZALTRAP (ziv-aflibercept) –
    Sanofi-Aventis U.S., LLC
    •
    ZIOPTAN (tafluprost)
    – Merck Sharp and Dohme Corp.

  • By Donald Zuhn —

    USPTO SealLast Friday, the U.S.
    Patent and Trademark Office published a final rule in the Federal Register (78
    Fed. Reg. 4212
    )
    setting and adjusting certain patent fees pursuant to the Office's fee-setting
    authority under § 10 of the Leahy-Smith America Invents Act.  The publication of the 80-page final rule
    (nearly) culminates a process that began last February when the Office
    submitted a proposed fee schedule to the Patent Public Advisory Committee
    (PPAC).  PPAC the held two public
    hearings on the proposed schedule, and in September issued a report on that
    schedule.  Following consideration of
    PPAC's report, the published a notice of proposed rulemaking setting and adjusting
    certain patent fees, which initiated a 45-day comment period that ended in
    November.  Although the AIA allows the
    new fee schedule to take effect 45 days after publication (i.e., March 4, 2013), in order to give Congress an opportunity to
    review the schedule, the notice indicates that the final rule will take effect
    on March 19, 2013, except for the following amendments, which will take effect
    on January 1, 2014:

    § 1.18(a)(1), (b)(1),
    (c)(1), and (d)(1) – patent issue and publication fees
    § 1.21(h)(1) – fee for
    recording a patent assignment electronically
    § 1.482(a)(1)(i)(A),
    (a)(1)(ii)(A), and (a)(2)(i) – international application filing, processing and
    search fees)
    § 1.445(a)(1)(i)(A),
    (a)(2)(i), (a)(3)(i), and (a)(4)(i) – international application transmittal and search fees

    In September, we posted
    several tables listing the fees for large entities that were proposed to change
    (see "More on USPTO's Proposed New Fees – Part II").  One table sorted the fees as they might be
    paid during the course of prosecution, another sorted the fees by amount, and a
    third table sorted the fees by dollar and percent change.  Because only seven of the patent fees for
    large entities changed from the notice of proposed rulemaking to the final
    rule, the table below lists the fees as they might be paid during the course of
    prosecution, with the current fee, proposed fee, final fee, and dollar change
    from the proposed fee to the final fee (click on table to enlarge):

    Final Fees

  • By Sherri Oslick —

    Gavel About Court Report:  Each week we will report briefly on recently filed biotech and pharma cases.

    Cadence
    Pharmaceuticals, Inc. et al. v. Fresenius Kabi USA, LLC

    3:13-cv-00139;
    filed January 17, 2013 in the Southern District of California

    • Plaintiffs: 
    Cadence Pharmaceuticals, Inc.; SCR Pharmatop
    • Defendant: 
    Fresenius Kabi USA, LLC

    Infringement
    of U.S. Patent Nos. 6,028,222 ("Stable Liquid Paracetamol Compositions,
    and Method for Preparing the Same," issued February 22, 2000) and
    6,992,218 ("Method for Obtaining Aqueous Formulations of
    Oxidation-Sensitive Active Principles," issued January 31, 2006), both
    licensed to Cadence, following a Paragraph IV certification as part of
    Fresenius' filing of an ANDA to manufacture a generic version of Cadence's
    Ofirmev® (acetaminophen injection, used for the management of mild to moderate
    pain, the management of moderate to severe pain with adjunctive opioid
    analgesics, and the reduction of fevers). 
    View the complaint here.


    Bioprojet SCR
    v. Kappos

    1:13-cv-00055;
    filed January 14, 2013 in the Eastern District of Virginia

    Review and
    correction of the patent term adjustment calculation made by the U.S. Patent
    and Trademark Office for U.S. Patent No. 8,222,294 ("Combinations of an
    Anti Emetic Agent and an Enkephalinase Inhibitor," issued July 17, 2012).  View the complaint here.


    Wyeth LLC et
    al. v. Kappos

    1:13-cv-00049;
    filed January 10, 2013 in the Eastern District of Virginia

    • Plaintiffs:
    Wyeth LLC; Janssen Alzheimer Immunotherapy
    • Defendant:
    Hon. David J. Kappos

    Review and
    correction of the patent term adjustment calculation made by the U.S. Patent
    and Trademark Office for U.S. Patent No. 8,227,403 ("A-β Immunogenic
    Peptide Carrier Conjugates and Methods of Producing Same," issued July 24,
    2012).  View the complaint here.

  • Calendar

    January 22, 2013 – Top Patent Law Stories of 2012 (McDonnell
    Boehnen Hulbert & Berghoff LLP) – 10:00 to
    11:15 am (CT)

    January 22, 2013 – Patent Term Adjustments and Extensions: Recent Developments —
    Leveraging Exelixis, Other Court
    Decisions and Recent USPTO Rule Changes to Maximize PTAs and PTEs
    (Strafford) – 1:00 – 2:30 pm (EST)

    January 23-24, 2013 – The Comprehensive Guide to Patent Reform*** (American Conference
    Institute) – New York, NY

    January 24, 2013 – Patent Term Adjustment: An Update (Intellectual Property Owners Association) – 2:00 – 3:00 pm (ET)

    January 24, 2013 – Canadian
    IP Law Update — Practical Tips and Information for U.S. Attorneys
    (Smart & Biggar) – Chicago, IL

    January
    25, 2013 – Coping With The New First-Inventor-to-File Prior Art Law: What Patent
    Prosecutors Need to Know and Do Before and After the Effective Date
    (Intellectual Property Law Association of Chicago) – Chicago, IL

    January 27, 2013 – Canadian
    IP Law Update — Practical Tips and Information for U.S. Attorneys
    (Smart & Biggar) – Minneapolis, MN

    January 29, 2013 – Combined Customer Partnership Meeting
    of TC3700 and TC1600
    (U.S. Patent and Trademark Office) – Alexandria, VA

    January
    29-30, 2013 – Biotech & Pharma Patent Litigation*** (C5) – Amsterdam, Netherlands

    February
    4-5, 2013 – Naples Midwinter Patent Law Experts Conference (Akron University Law School) – Naples, FL

    February 5, 2013 – Single Claim Restriction Requirements: 
The Interplay Between 35 USC
    Section 112 and 35 USC Section 121
    (American Bar
    Association) – 12:00
    – 1:30 pm (Eastern)

    February 13, 2013 – Corporate Intellectual
    Property Law Conference
    (Law Bulletin
    Publishing Company) – Chicago, IL

    February 14, 2013 – Last Chance! The Top Ten Things
    In-House Counsel Need to Do to Prepare for The AIA’s March 2013 Deadline
    (McDonnell
    Boehnen Hulbert & Berghoff LLP) – 10:00 am to 11:15 am (CST)

    February 20, 2013 – AIA Strategy Considerations for Patent Filings in
    2013: Should You File on March 15 or March 16 (or Both)?
    (Intellectual Property Owners Association) – Washington, DC

    February 20-22, 2013 – Intensive Patent Law Seminar (Chisum Patent Academy) – New York, NY

    February
    26-27, 2013 – Biotech & Pharmaceutical Patenting (IBC Legal) – Munich, Germany

    February 27-28, 2013 – Life Sciences
    Collaborative Agreements and Acquisitions
    *** (American Conference
    Institute) – New
    York, NY

    March 5-6, 2013 – Medical Device
    Patents
    *** (American Conference
    Institute) – Chicago, IL

    March 19-20, 2013 – FDA Boot Camp*** (American Conference
    Institute) – New York, NY

    ***Patent Docs is a media partner of this conference or CLE

  • New York #1American Conference
    Institute (ACI) will be holding the next session of its FDA Boot Camp
    conference on March 19-20, 2013 in New York, NY.  ACI faculty will help attendees:

    • Master the basics
    of the application and approval processes for drugs, biologics, and devices;
    • Comprehend the
    structure of the FDA and the roles of the three major agency centers:  CDER, CBER, and CDHR;
    • Develop a
    practical working knowledge of clinical trials for drugs and biologics and the
    clearance process for devices;
    • Learn how devices
    are classified, monitored, and regulated;
    • Appreciate the
    complexities of pharmaceutical IP and the regulatory balance between brand name
    and generic products;
    • Recognize the
    pivotal role of labeling in the drug and biologics approval process;
    • See the
    importance of cGMPs to the post-approval regulatory process; and
    • Navigate the
    protocols of adverse events monitoring, signal detection, product withdrawals,
    and recalls.

    BrochureIn particular,
    ACI's faculty will offer presentations on the following topics:

    • The basics:  Understanding and working with the FDA —
    Jurisdiction, functions, organization, and operations;
    • The nature of the
    approval process;
    • Understanding the
    clinical trial process for drugs and biologics;
    • IP overview for
    drugs and biologics:  Hatch-Waxman,
    BPCIA, trade dress, and more:
    • Part 1 —
    Overview of patent and trademark issues;
    • Part 2 —
    Hatch-Waxman and BPCIA overview;
    • Drugs and
    biologics:  Labeling;
    • cGMPs:  Drugs and biologics (current good manufacturing
    practices);
    • Medical
    devices:  Classifications, the essentials
    of the premarket review process, and post-market requirements and concerns;
    • Adverse events
    monitoring, pharmacovigilance and risk management; and
    • Recall guidance
    for drugs, biologics, and medical devices: 
    What you need to know.

    A pre-conference
    workshop on the "Fundamentals of FDA Regulatory Law" and
    "Resolving Ethical Challenges Encountered During the Drug Approval
    Process" will be offered on March 18, 2013 from 1:00 to 5:00 pm.  The workshop will provide a basic overview of
    FDA regulations and will prepare attendees for the in-depth discussions that
    will take place throughout the conference, and explore ethical issues that may
    arise in the context of communications with FDA on behalf of clients.

    Two post-conference
    master classes will be offered on March 20, 2013.  The first master class, entitled
    "Hatch-Waxman and BPCIA: Overview of Biosimilars and Life Cycle Planning
    for Drugs and Biologics," will provide an in-depth overview of biosimilars
    as well as analyses of bioequivalence and exclusivities and their role in
    patent and product life cycle management. 
    The second master class, entitled "Post-Approval Marketing Guidance
    and Preemption Protocols," will address issues that arise post-approval,
    including advertising, promotion, and off-label promotion and enforcement, as
    well as preemption fundamentals.

    An agenda for the
    conference can be found here.  A complete brochure for this conference,
    including an agenda, detailed descriptions of conference sessions, list of
    speakers, and registration form can be obtained here.

    ACI - American Conference InstituteThe registration
    fee is $2,295 (conference alone), $2,895 (conference and workshop or conference
    and one master class), or $3,295 (conference, workshop, and one master
    class).  Those registering by January 23,
    2013 will receive a $300 discount and those registering by February 20, 2013
    will receive a $200 discount.  Patent Docs readers who reference the
    discount code "PD 200" will receive $200 off the current price tier
    when registering.  Those interested in
    registering for the conference can do so here, by
    e-mailing CustomerService@AmericanConference.com, by calling 1-888-224-2480, or
    by faxing a registration form to 1-877-927-1563.

    Patent Docs is a media partner of ACI's FDA Boot Camp conference.

  • IPO #2The
    Intellectual Property Owners Association (IPO) will offer a one-hour webinar entitled
    "Patent Term Adjustment: An Update"
    on January 24, 2013 beginning at 2:00 pm (ET). 
    A panel consisting of Patricia
    Carson of Kirkland & Ellis, LLP; Adrian Looney of Pfizer, Inc.; and Eli Loots of Knobbe Martens Olson & Bear, LLP will consider:

    •
    What steps patent owners should take in light of recent court decisions such as
    Exelixis v. Kappos;
    •
    Recent USPTO rule changes including revisions in the rules of practice
    regarding the period of appellate review; and
    •
    The impact of the technical amendments to the AIA that became law this month.

    The
    registration fee for the webinar is $120 (government and academic rates are
    available upon request).  Those
    interested in registering for the webinar can do so here.

  • ABAThe American Bar
    Association (ABA) Section of Intellectual Property Law, Section of Science and
    Technology, and Center for Professional Development will be offering a live
    webinar and teleconference entitled "Single Claim Restriction Requirements: 
The Interplay Between 35 USC
    Section 112 and 35 USC Section 121" on February 5, 2013 from 12:00
    – 1:30 pm (Eastern).  Keisha Hylton-Rodic of Drinker Biddle & Reath LLP will
    moderate a panel including Thomas L.
    Irving of Finnegan, Henderson,
    Farabow, Garrett & Dunner, LLP; Orlando Lopez of Burns
    & Levinson LLP; Daniel M.
    Sullivan of the U.S. Patent and
    Trademark Office; and Robert D.
    Titus of Eli Lilly.  The panel will examine the balance between 35
    U.S.C. § 112 and 35 U.S.C. § 121, and their interplay in the context of single
    claim restriction requirements and restriction practice as it relates to Markush claims, as well as address
    tactics that can be used during prosecution to avoid the pitfall of single
    claim restrictions in chemical and biotech cases in contravention of current
    case law on the practice.

    The registration fee for
    the webcast is $95 for members of any of the sections sponsoring the webinar,
    $99 for government attorneys, $150 for ABA members, and $195 for the general
    public.  Those interested in registering
    for the webinar, can do so here
    or by calling 800-285-2221.

  • IPLACThe
    Intellectual Property Law Association of Chicago (IPLAC) will be offering a program
    on "Coping With The New First-Inventor-to-File Prior Art Law: What Patent
    Prosecutors Need to Know and Do Before and After the Effective Date" on January
    25, 2013 from 12:00 to 2:30 pm (Central) at the DePaul University Law School in
    Chicago, IL.  Edward Manzo of Husch
    Blackwell LLP, Joseph Hetz of Brinks Hofer Gilson & Lione, John Paniaguas
    of Katten Munchin Rosenman LLP, and George Wheeler of McAndrews, Held &
    Malloy, Ltd. will provide patent prosecutors with practical tips and strategies
    to file and prosecute patent applications in light of the imminent new
    definition of prior art under the Leahy-Smith America Invents Act (AIA),
    enacted on September 16, 2011, and soon to be effective.  The registration fee for the program is $35
    for IPLAC members, $45 for non-members, $25 for students (other than DePaul
    students, for whom there is no fee).  Those
    interested in registering for event can do so here.

  • By Kevin E. Noonan —

    Federal Trade Commission (FTC) SealAs it has frequently in
    the past decade, the Federal Trade Commission on Thursday released a Report on
    the frequency of reverse payment settlement agreements in ANDA litigation
    between generic and branded drug makers, pursuant to its authority to
    scrutinize all ANDA agreements under the provisions of the Medicare
    Modernization Act of 2003.

    The FTC Report contains
    information from the 2012 fiscal year (from October 1, 2011 through September
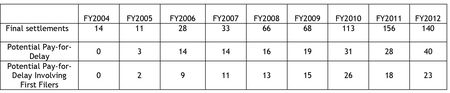
    30, 2012).  There were 40 reverse payment
    agreements among 140 "final resolutions" of ANDA litigation.  According to the Report, these agreements
    involved 31 different branded drugs having total combined sales of $8.3
    billion.  Nineteen of the 40 agreements
    contained provisions wherein the branded drug maker agreed not to bring an "authorized
    generic" version of the branded drug to market (thus removing a strong
    competitor for the generic drug), at least for some period of time.  The Report also notes that seven branded
    drugs entered into "multiple" reverse payment settlements, i.e., with different generic
    challengers.  The Report compares this
    trend with historical data from 2004, noting that in one case a branded drug
    maker had entered into reverse payment settlement agreements with ten generic
    challengers, with 14 branded drugs being involved in payments to three or more
    generic drug makers.

    Of the other 100
    settlement agreements, 81 contained provisions without express payments but
    with restrictions on time for generic entry, which the final 19 agreements
    contained no restrictions of the generic challenger.

    Forty-three of these
    140 settlements were between the branded drug maker and the first ANDA filer.  Of these, 23 settlement agreements had provisions both for payments and delay
    or other restrictions on generic drug entry with another 16 agreements
    providing for generic delay but no express compensation.

    The Report draws
    conclusions from this data, the most significant of which is that there were a "record"
    number of reverse payment settlement agreements in fiscal 2012.  The 40 agreements containing reverse payment
    provisions increased from the 28 such agreements in fiscal 2011, and the 19
    agreements providing for no authorized generic entry (but without express
    payments) were also a record (up from 11 in fiscal 2012, perhaps reflecting the
    effects of the FTC's campaign against agreements containing payment provisions
    to generic drugmakers).

    In the end, however,
    the FTC was forced to admit that "[d]espite the record number of potential
    pay-for delay settlements in FY 2012, the vast majority of patent settlements
    (greater than 70%) continued to be resolved without compensation to the generic
    manufacturer," at least suggesting that the Commission itself has created
    this particular tempest in its teapot.  The Report concludes with a table comparing rates of ANDA settlements
    and the frequency of reverse payment settlement agreements since FY 2004.

    Table
    Looking at these data dispassionately,
    the number of settlements of ANDA litigation has decreased, while the number of
    settlements having some economic incentive for generic drug makers has
    increased.  However, the data also show
    that this economic incentive in the form of direct payments has actually
    decreased, and the Commission's statistics show an increase because it has
    included agreements with "no authorized generic" provisions as "potential
    pay-for-delay" agreements.

    In a press release
    accompany the Report, the Commission highlighted these increases,
    characterizing them as having "significantly increased" and
    comprising a "record number" of such agreements.  (As is its wont, the Commission characterizes
    these agreements as "pay for delay," even for agreements with no
    express payments from the branded to the generic drugmaker.)  But the Commission considers even these
    agreement to be "potentially anticompetitive," saying that "[s]uch 'no-AG' [authorized generic] promises are valuable to generic firms,
    as they significantly reduce the level of competition the new generic entrant
    will face, allowing the generic firm to secure greater market share and extract
    higher prices from consumers."  The
    latter statement is, of course, at least unlikely to be untrue:  the value of a "no-AG"
    promise is that the generic drug company will make more money, of course, but
    not by charging higher prices but by recouping more of the total generic market
    as a consequence of not having to split the market with the authorized generic
    product.

    FTC Chairman Jon Leibowitz added his own
    ideological opinion on these results, saying:

    Sadly, this year's report makes it clear that
    the problem of pay-for-delay is getting worse, not better.  . . .  More
    and more brand and generic drug companies are engaging in these sweetheart
    deals, and consumers continue to pay the price.  Until this issue is
    resolved, we will all suffer the consequences of delayed generic entry — higher
    prices for consumers, businesses, and the U.S. taxpayer.

    This sentiment is
    continued in the remainder of the press release, which correctly notes that the
    average "delay" for generic drug entry is 17 months, but ignores the
    reality that this is a delay from a putative successful ANDA challenge.  It does not take into effect the acceleration
    of generic drug entry that results from settlement of ANDA litigation rather
    than that proportion of these cases where the generic drug challenger loses and
    is barred from the marketplace until the challenged Orange Book listed patents
    expire.  While this emphasis on delay is
    understandable in view of the Commission's ideological biases (and its
    unsupported and unrealistic belief that only holders of "bad" patents
    settle ANDA litigation), it does not give the public a clear picture of the
    actual circumstances behind the data the Commission presents.

    This may be the last
    Report from the Commission on reverse payment settlement agreement frequency,
    should the FTC prevail in its challenge to these agreements before the Supreme
    Court (FTC v. Watson Pharmaceuticals).  That would be the culmination of a steady
    stream of Commission, district court, and appellate court challenges to these
    agreements, as well as several stalled legislative efforts.  The Commission's position has been rejected
    often by several appellate courts (including the 2nd, 9th,
    11th and Federal Circuit courts of appeal) based on an analysis of
    the business and legal realities not overburdened with the Commission's
    ideological bias (i.e., that any delay in generic drug entry is anticompetitive
    per se).  The Supreme Court should have the last word
    on this issue.